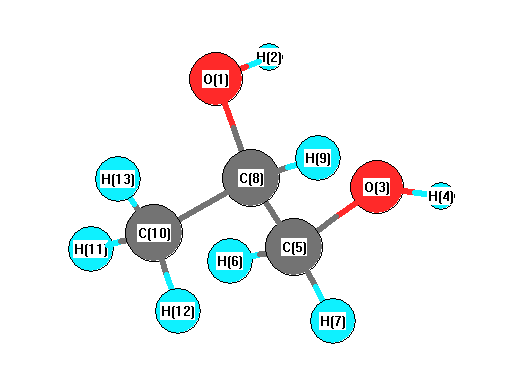
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3656 |
3563 |
8.83 |
|
|
|
| 2 |
A |
3616 |
3524 |
16.52 |
|
|
|
| 3 |
A |
3107 |
3028 |
54.49 |
|
|
|
| 4 |
A |
3101 |
3022 |
24.88 |
|
|
|
| 5 |
A |
3083 |
3005 |
31.04 |
|
|
|
| 6 |
A |
3003 |
2926 |
71.17 |
|
|
|
| 7 |
A |
2995 |
2919 |
9.27 |
|
|
|
| 8 |
A |
2933 |
2858 |
81.65 |
|
|
|
| 9 |
A |
1522 |
1483 |
5.29 |
|
|
|
| 10 |
A |
1504 |
1466 |
7.55 |
|
|
|
| 11 |
A |
1496 |
1458 |
1.79 |
|
|
|
| 12 |
A |
1435 |
1399 |
15.83 |
|
|
|
| 13 |
A |
1426 |
1390 |
17.31 |
|
|
|
| 14 |
A |
1380 |
1345 |
7.87 |
|
|
|
| 15 |
A |
1372 |
1337 |
21.82 |
|
|
|
| 16 |
A |
1335 |
1301 |
26.94 |
|
|
|
| 17 |
A |
1273 |
1240 |
50.51 |
|
|
|
| 18 |
A |
1208 |
1177 |
14.84 |
|
|
|
| 19 |
A |
1158 |
1129 |
6.81 |
|
|
|
| 20 |
A |
1078 |
1050 |
24.02 |
|
|
|
| 21 |
A |
1062 |
1035 |
54.10 |
|
|
|
| 22 |
A |
996 |
970 |
95.48 |
|
|
|
| 23 |
A |
943 |
919 |
9.85 |
|
|
|
| 24 |
A |
907 |
884 |
24.40 |
|
|
|
| 25 |
A |
829 |
808 |
18.96 |
|
|
|
| 26 |
A |
523 |
509 |
196.03 |
|
|
|
| 27 |
A |
495 |
483 |
34.69 |
|
|
|
| 28 |
A |
467 |
455 |
6.05 |
|
|
|
| 29 |
A |
349 |
340 |
10.98 |
|
|
|
| 30 |
A |
285 |
278 |
110.92 |
|
|
|
| 31 |
A |
238 |
232 |
5.36 |
|
|
|
| 32 |
A |
206 |
201 |
0.76 |
|
|
|
| 33 |
A |
136 |
133 |
7.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24559.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 23932.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.480 |
|
|
|
| 2 |
H |
0.347 |
|
|
|
| 3 |
O |
-0.467 |
|
|
|
| 4 |
H |
0.338 |
|
|
|
| 5 |
C |
-0.258 |
|
|
|
| 6 |
H |
0.170 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
C |
0.154 |
|
|
|
| 9 |
H |
0.103 |
|
|
|
| 10 |
C |
-0.493 |
|
|
|
| 11 |
H |
0.154 |
|
|
|
| 12 |
H |
0.132 |
|
|
|
| 13 |
H |
0.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.485 |
-2.397 |
1.836 |
3.058 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.798 |
2.125 |
3.433 |
| y |
2.125 |
-33.155 |
0.189 |
| z |
3.433 |
0.189 |
-30.214 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.113 |
2.125 |
3.433 |
| y |
2.125 |
-1.149 |
0.189 |
| z |
3.433 |
0.189 |
3.262 |
|
| Polar |
|---|
| 3z2-r2 | 6.524 |
|---|
| x2-y2 | -0.643 |
|---|
| xy | 2.125 |
|---|
| xz | 3.433 |
|---|
| yz | 0.189 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.166 |
0.301 |
0.198 |
| y |
0.301 |
6.455 |
-0.178 |
| z |
0.198 |
-0.178 |
6.110 |
<r2> (average value of r
2) Å
2
| <r2> |
114.857 |
| (<r2>)1/2 |
10.717 |