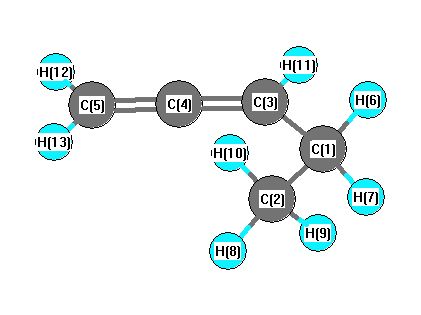
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3169 |
3088 |
14.70 |
|
|
|
| 2 |
A |
3088 |
3009 |
19.25 |
|
|
|
| 3 |
A |
3083 |
3004 |
18.00 |
|
|
|
| 4 |
A |
3081 |
3002 |
56.04 |
|
|
|
| 5 |
A |
3077 |
2999 |
55.39 |
|
|
|
| 6 |
A |
3003 |
2927 |
28.59 |
|
|
|
| 7 |
A |
2992 |
2915 |
29.11 |
|
|
|
| 8 |
A |
2970 |
2894 |
57.92 |
|
|
|
| 9 |
A |
2008 |
1957 |
34.62 |
|
|
|
| 10 |
A |
1525 |
1486 |
7.00 |
|
|
|
| 11 |
A |
1518 |
1479 |
9.07 |
|
|
|
| 12 |
A |
1512 |
1474 |
0.64 |
|
|
|
| 13 |
A |
1480 |
1443 |
5.45 |
|
|
|
| 14 |
A |
1435 |
1398 |
6.50 |
|
|
|
| 15 |
A |
1385 |
1350 |
14.01 |
|
|
|
| 16 |
A |
1339 |
1305 |
1.74 |
|
|
|
| 17 |
A |
1285 |
1253 |
0.27 |
|
|
|
| 18 |
A |
1153 |
1124 |
1.54 |
|
|
|
| 19 |
A |
1114 |
1086 |
2.90 |
|
|
|
| 20 |
A |
1093 |
1065 |
2.86 |
|
|
|
| 21 |
A |
1044 |
1017 |
0.01 |
|
|
|
| 22 |
A |
997 |
971 |
1.26 |
|
|
|
| 23 |
A |
911 |
888 |
17.86 |
|
|
|
| 24 |
A |
900 |
877 |
72.43 |
|
|
|
| 25 |
A |
836 |
815 |
2.35 |
|
|
|
| 26 |
A |
798 |
778 |
4.53 |
|
|
|
| 27 |
A |
636 |
620 |
4.43 |
|
|
|
| 28 |
A |
532 |
519 |
3.95 |
|
|
|
| 29 |
A |
361 |
352 |
4.50 |
|
|
|
| 30 |
A |
350 |
341 |
9.86 |
|
|
|
| 31 |
A |
236 |
230 |
0.06 |
|
|
|
| 32 |
A |
150 |
146 |
0.55 |
|
|
|
| 33 |
A |
88 |
85 |
0.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24574.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 23948.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.248 |
|
|
|
| 2 |
C |
-0.472 |
|
|
|
| 3 |
C |
-0.276 |
|
|
|
| 4 |
C |
0.399 |
|
|
|
| 5 |
C |
-0.578 |
|
|
|
| 6 |
H |
0.143 |
|
|
|
| 7 |
H |
0.143 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.141 |
|
|
|
| 10 |
H |
0.151 |
|
|
|
| 11 |
H |
0.158 |
|
|
|
| 12 |
H |
0.146 |
|
|
|
| 13 |
H |
0.145 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.509 |
0.071 |
-0.001 |
0.514 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.759 |
-1.867 |
0.000 |
| y |
-1.867 |
-31.872 |
-0.001 |
| z |
0.000 |
-0.001 |
-32.309 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.332 |
-1.867 |
0.000 |
| y |
-1.867 |
-0.338 |
-0.001 |
| z |
0.000 |
-0.001 |
-0.993 |
|
| Polar |
|---|
| 3z2-r2 | -1.986 |
|---|
| x2-y2 | 1.113 |
|---|
| xy | -1.867 |
|---|
| xz | 0.000 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.135 |
-2.499 |
-0.004 |
| y |
-2.499 |
8.246 |
-0.003 |
| z |
-0.004 |
-0.003 |
6.139 |
<r2> (average value of r
2) Å
2
| <r2> |
122.374 |
| (<r2>)1/2 |
11.062 |