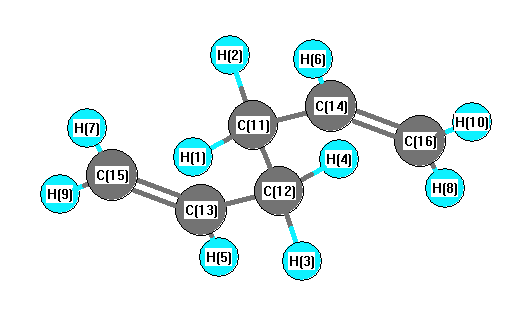
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3197 |
3115 |
19.23 |
|
|
|
| 2 |
A |
3101 |
3022 |
1.67 |
|
|
|
| 3 |
A |
3086 |
3008 |
69.42 |
|
|
|
| 4 |
A |
3050 |
2973 |
30.35 |
|
|
|
| 5 |
A |
2967 |
2892 |
14.17 |
|
|
|
| 6 |
A |
1671 |
1628 |
6.86 |
|
|
|
| 7 |
A |
1506 |
1467 |
1.88 |
|
|
|
| 8 |
A |
1458 |
1421 |
3.45 |
|
|
|
| 9 |
A |
1363 |
1328 |
0.86 |
|
|
|
| 10 |
A |
1322 |
1288 |
0.64 |
|
|
|
| 11 |
A |
1243 |
1211 |
0.15 |
|
|
|
| 12 |
A |
1091 |
1063 |
0.14 |
|
|
|
| 13 |
A |
1028 |
1002 |
5.10 |
|
|
|
| 14 |
A |
1026 |
1000 |
6.34 |
|
|
|
| 15 |
A |
961 |
936 |
25.57 |
|
|
|
| 16 |
A |
945 |
920 |
3.61 |
|
|
|
| 17 |
A |
792 |
772 |
0.39 |
|
|
|
| 18 |
A |
629 |
613 |
8.61 |
|
|
|
| 19 |
A |
411 |
401 |
0.00 |
|
|
|
| 20 |
A |
344 |
335 |
0.18 |
|
|
|
| 21 |
A |
95 |
92 |
0.04 |
|
|
|
| 22 |
A |
70 |
68 |
0.04 |
|
|
|
| 23 |
B |
3197 |
3115 |
59.86 |
|
|
|
| 24 |
B |
3100 |
3021 |
18.00 |
|
|
|
| 25 |
B |
3085 |
3006 |
20.15 |
|
|
|
| 26 |
B |
3029 |
2952 |
9.05 |
|
|
|
| 27 |
B |
2969 |
2894 |
66.25 |
|
|
|
| 28 |
B |
1674 |
1631 |
23.26 |
|
|
|
| 29 |
B |
1520 |
1481 |
11.50 |
|
|
|
| 30 |
B |
1460 |
1423 |
0.56 |
|
|
|
| 31 |
B |
1328 |
1294 |
0.47 |
|
|
|
| 32 |
B |
1309 |
1275 |
0.53 |
|
|
|
| 33 |
B |
1288 |
1255 |
1.19 |
|
|
|
| 34 |
B |
1182 |
1152 |
2.53 |
|
|
|
| 35 |
B |
1031 |
1005 |
26.58 |
|
|
|
| 36 |
B |
969 |
945 |
7.55 |
|
|
|
| 37 |
B |
958 |
934 |
91.68 |
|
|
|
| 38 |
B |
925 |
902 |
0.02 |
|
|
|
| 39 |
B |
661 |
644 |
25.00 |
|
|
|
| 40 |
B |
435 |
424 |
1.61 |
|
|
|
| 41 |
B |
214 |
209 |
0.14 |
|
|
|
| 42 |
B |
109 |
106 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30898.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30110.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.382 |
|
|
|
| 2 |
C |
-0.382 |
|
|
|
| 3 |
C |
0.143 |
|
|
|
| 4 |
C |
0.143 |
|
|
|
| 5 |
C |
-0.411 |
|
|
|
| 6 |
C |
-0.411 |
|
|
|
| 7 |
H |
0.137 |
|
|
|
| 8 |
H |
0.140 |
|
|
|
| 9 |
H |
0.137 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
H |
0.121 |
|
|
|
| 12 |
H |
0.121 |
|
|
|
| 13 |
H |
0.118 |
|
|
|
| 14 |
H |
0.133 |
|
|
|
| 15 |
H |
0.118 |
|
|
|
| 16 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.317 |
0.317 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.857 |
1.677 |
0.000 |
| y |
1.677 |
-40.271 |
0.000 |
| z |
0.000 |
0.000 |
-37.564 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.939 |
1.677 |
0.000 |
| y |
1.677 |
-1.560 |
0.000 |
| z |
0.000 |
0.000 |
2.500 |
|
| Polar |
|---|
| 3z2-r2 | 4.999 |
|---|
| x2-y2 | 0.414 |
|---|
| xy | 1.677 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.861 |
1.436 |
0.000 |
| y |
1.436 |
13.796 |
0.000 |
| z |
0.000 |
0.000 |
9.882 |
<r2> (average value of r
2) Å
2
| <r2> |
207.528 |
| (<r2>)1/2 |
14.406 |