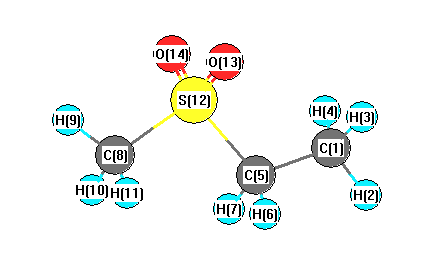
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3190 |
3109 |
1.64 |
|
|
|
| 2 |
A |
3187 |
3106 |
0.56 |
|
|
|
| 3 |
A |
3147 |
3067 |
5.94 |
|
|
|
| 4 |
A |
3108 |
3029 |
4.57 |
|
|
|
| 5 |
A |
3082 |
3004 |
21.38 |
|
|
|
| 6 |
A |
3058 |
2980 |
5.13 |
|
|
|
| 7 |
A |
3039 |
2961 |
2.80 |
|
|
|
| 8 |
A |
3001 |
2925 |
20.73 |
|
|
|
| 9 |
A |
1524 |
1485 |
8.80 |
|
|
|
| 10 |
A |
1510 |
1471 |
15.19 |
|
|
|
| 11 |
A |
1471 |
1434 |
7.58 |
|
|
|
| 12 |
A |
1462 |
1425 |
11.18 |
|
|
|
| 13 |
A |
1446 |
1409 |
7.97 |
|
|
|
| 14 |
A |
1439 |
1402 |
15.11 |
|
|
|
| 15 |
A |
1342 |
1308 |
1.08 |
|
|
|
| 16 |
A |
1290 |
1257 |
17.02 |
|
|
|
| 17 |
A |
1255 |
1223 |
1.94 |
|
|
|
| 18 |
A |
1089 |
1062 |
18.01 |
|
|
|
| 19 |
A |
1079 |
1051 |
5.40 |
|
|
|
| 20 |
A |
1005 |
979 |
24.00 |
|
|
|
| 21 |
A |
994 |
968 |
5.98 |
|
|
|
| 22 |
A |
975 |
950 |
22.13 |
|
|
|
| 23 |
A |
842 |
821 |
43.62 |
|
|
|
| 24 |
A |
789 |
769 |
21.64 |
|
|
|
| 25 |
A |
740 |
721 |
29.02 |
|
|
|
| 26 |
A |
623 |
607 |
16.97 |
|
|
|
| 27 |
A |
520 |
507 |
2.46 |
|
|
|
| 28 |
A |
372 |
362 |
6.91 |
|
|
|
| 29 |
A |
303 |
296 |
25.64 |
|
|
|
| 30 |
A |
280 |
273 |
32.77 |
|
|
|
| 31 |
A |
237 |
231 |
2.58 |
|
|
|
| 32 |
A |
220 |
214 |
1.34 |
|
|
|
| 33 |
A |
176 |
172 |
3.03 |
|
|
|
| 34 |
A |
161 |
157 |
1.58 |
|
|
|
| 35 |
A |
127 |
124 |
0.56 |
|
|
|
| 36 |
A |
64 |
62 |
3.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24073.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 23459.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.459 |
|
|
|
| 2 |
H |
0.214 |
|
|
|
| 3 |
H |
0.177 |
|
|
|
| 4 |
H |
0.154 |
|
|
|
| 5 |
C |
-0.401 |
|
|
|
| 6 |
H |
0.233 |
|
|
|
| 7 |
H |
0.194 |
|
|
|
| 8 |
C |
-0.567 |
|
|
|
| 9 |
H |
0.200 |
|
|
|
| 10 |
H |
0.239 |
|
|
|
| 11 |
H |
0.199 |
|
|
|
| 12 |
S |
0.874 |
|
|
|
| 13 |
O |
-0.533 |
|
|
|
| 14 |
O |
-0.523 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.051 |
3.534 |
-3.900 |
6.642 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.997 |
-8.360 |
-0.245 |
| y |
-8.360 |
-43.947 |
-1.992 |
| z |
-0.245 |
-1.992 |
-50.967 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.460 |
-8.360 |
-0.245 |
| y |
-8.360 |
4.035 |
-1.992 |
| z |
-0.245 |
-1.992 |
-6.495 |
|
| Polar |
|---|
| 3z2-r2 | -12.990 |
|---|
| x2-y2 | -1.050 |
|---|
| xy | -8.360 |
|---|
| xz | -0.245 |
|---|
| yz | -1.992 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.191 |
-0.150 |
0.237 |
| y |
-0.150 |
9.748 |
0.144 |
| z |
0.237 |
0.144 |
9.450 |
<r2> (average value of r
2) Å
2
| <r2> |
174.755 |
| (<r2>)1/2 |
13.219 |