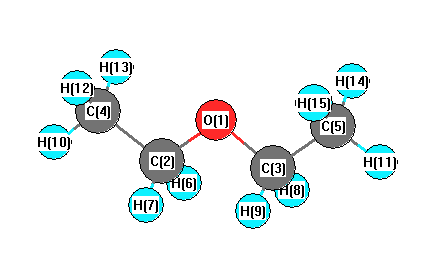
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3085 |
3006 |
65.88 |
|
|
|
| 2 |
A1 |
2999 |
2922 |
4.25 |
|
|
|
| 3 |
A1 |
2940 |
2865 |
144.07 |
|
|
|
| 4 |
A1 |
1546 |
1507 |
2.98 |
|
|
|
| 5 |
A1 |
1521 |
1483 |
5.15 |
|
|
|
| 6 |
A1 |
1446 |
1409 |
6.52 |
|
|
|
| 7 |
A1 |
1423 |
1387 |
1.19 |
|
|
|
| 8 |
A1 |
1164 |
1134 |
9.87 |
|
|
|
| 9 |
A1 |
1043 |
1016 |
10.40 |
|
|
|
| 10 |
A1 |
823 |
802 |
8.35 |
|
|
|
| 11 |
A1 |
418 |
408 |
0.09 |
|
|
|
| 12 |
A1 |
184 |
179 |
0.96 |
|
|
|
| 13 |
A2 |
3099 |
3020 |
0.00 |
|
|
|
| 14 |
A2 |
2979 |
2903 |
0.00 |
|
|
|
| 15 |
A2 |
1503 |
1464 |
0.00 |
|
|
|
| 16 |
A2 |
1279 |
1246 |
0.00 |
|
|
|
| 17 |
A2 |
1160 |
1130 |
0.00 |
|
|
|
| 18 |
A2 |
821 |
800 |
0.00 |
|
|
|
| 19 |
A2 |
237 |
231 |
0.00 |
|
|
|
| 20 |
A2 |
93 |
90 |
0.00 |
|
|
|
| 21 |
B1 |
3099 |
3020 |
103.63 |
|
|
|
| 22 |
B1 |
2982 |
2906 |
150.74 |
|
|
|
| 23 |
B1 |
1503 |
1465 |
14.61 |
|
|
|
| 24 |
B1 |
1287 |
1254 |
1.28 |
|
|
|
| 25 |
B1 |
1186 |
1156 |
14.24 |
|
|
|
| 26 |
B1 |
835 |
814 |
1.55 |
|
|
|
| 27 |
B1 |
241 |
235 |
2.15 |
|
|
|
| 28 |
B1 |
100 |
97 |
7.12 |
|
|
|
| 29 |
B2 |
3085 |
3006 |
31.66 |
|
|
|
| 30 |
B2 |
2998 |
2922 |
57.63 |
|
|
|
| 31 |
B2 |
2928 |
2853 |
8.82 |
|
|
|
| 32 |
B2 |
1532 |
1493 |
0.64 |
|
|
|
| 33 |
B2 |
1516 |
1478 |
8.50 |
|
|
|
| 34 |
B2 |
1433 |
1397 |
20.41 |
|
|
|
| 35 |
B2 |
1375 |
1340 |
53.32 |
|
|
|
| 36 |
B2 |
1117 |
1088 |
93.92 |
|
|
|
| 37 |
B2 |
1092 |
1064 |
95.76 |
|
|
|
| 38 |
B2 |
940 |
916 |
15.01 |
|
|
|
| 39 |
B2 |
417 |
407 |
4.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29712.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 28955.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.279 |
|
|
|
| 2 |
C |
-0.094 |
|
|
|
| 3 |
C |
-0.094 |
|
|
|
| 4 |
C |
-0.447 |
|
|
|
| 5 |
C |
-0.447 |
|
|
|
| 6 |
H |
0.115 |
|
|
|
| 7 |
H |
0.115 |
|
|
|
| 8 |
H |
0.115 |
|
|
|
| 9 |
H |
0.115 |
|
|
|
| 10 |
H |
0.135 |
|
|
|
| 11 |
H |
0.135 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
| 14 |
H |
0.157 |
|
|
|
| 15 |
H |
0.157 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.507 |
1.507 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.084 |
0.000 |
0.000 |
| y |
0.000 |
9.579 |
0.000 |
| z |
0.000 |
0.000 |
7.398 |
<r2> (average value of r
2) Å
2
| <r2> |
151.778 |
| (<r2>)1/2 |
12.320 |