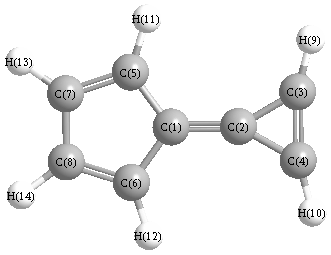
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3278 |
3195 |
4.12 |
|
|
|
| 2 |
A1 |
3212 |
3131 |
28.56 |
|
|
|
| 3 |
A1 |
3182 |
3101 |
11.95 |
|
|
|
| 4 |
A1 |
1776 |
1730 |
553.62 |
|
|
|
| 5 |
A1 |
1515 |
1476 |
73.74 |
|
|
|
| 6 |
A1 |
1472 |
1435 |
61.38 |
|
|
|
| 7 |
A1 |
1393 |
1358 |
110.45 |
|
|
|
| 8 |
A1 |
1188 |
1158 |
31.85 |
|
|
|
| 9 |
A1 |
1089 |
1061 |
11.41 |
|
|
|
| 10 |
A1 |
1014 |
988 |
6.26 |
|
|
|
| 11 |
A1 |
951 |
927 |
6.66 |
|
|
|
| 12 |
A1 |
875 |
853 |
50.48 |
|
|
|
| 13 |
A1 |
474 |
462 |
0.36 |
|
|
|
| 14 |
A2 |
947 |
922 |
0.00 |
|
|
|
| 15 |
A2 |
913 |
890 |
0.00 |
|
|
|
| 16 |
A2 |
761 |
742 |
0.00 |
|
|
|
| 17 |
A2 |
591 |
576 |
0.00 |
|
|
|
| 18 |
A2 |
211 |
206 |
0.00 |
|
|
|
| 19 |
B1 |
931 |
907 |
2.75 |
|
|
|
| 20 |
B1 |
799 |
779 |
119.96 |
|
|
|
| 21 |
B1 |
722 |
704 |
81.80 |
|
|
|
| 22 |
B1 |
633 |
617 |
7.08 |
|
|
|
| 23 |
B1 |
426 |
415 |
12.72 |
|
|
|
| 24 |
B1 |
146 |
143 |
6.05 |
|
|
|
| 25 |
B2 |
3231 |
3149 |
2.10 |
|
|
|
| 26 |
B2 |
3200 |
3118 |
47.12 |
|
|
|
| 27 |
B2 |
3171 |
3090 |
1.79 |
|
|
|
| 28 |
B2 |
1549 |
1510 |
1.06 |
|
|
|
| 29 |
B2 |
1333 |
1299 |
0.16 |
|
|
|
| 30 |
B2 |
1227 |
1196 |
1.84 |
|
|
|
| 31 |
B2 |
1100 |
1072 |
25.04 |
|
|
|
| 32 |
B2 |
1091 |
1063 |
6.34 |
|
|
|
| 33 |
B2 |
898 |
875 |
5.19 |
|
|
|
| 34 |
B2 |
833 |
812 |
1.62 |
|
|
|
| 35 |
B2 |
485 |
472 |
0.95 |
|
|
|
| 36 |
B2 |
147 |
143 |
1.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23382.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 22786.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.724 |
|
|
|
| 2 |
C |
-0.697 |
|
|
|
| 3 |
C |
-0.194 |
|
|
|
| 4 |
C |
-0.194 |
|
|
|
| 5 |
C |
-0.180 |
|
|
|
| 6 |
C |
-0.180 |
|
|
|
| 7 |
C |
-0.215 |
|
|
|
| 8 |
C |
-0.215 |
|
|
|
| 9 |
H |
0.250 |
|
|
|
| 10 |
H |
0.250 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
| 12 |
H |
0.175 |
|
|
|
| 13 |
H |
0.151 |
|
|
|
| 14 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
4.412 |
4.412 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.427 |
0.000 |
0.000 |
| y |
0.000 |
-41.793 |
0.000 |
| z |
0.000 |
0.000 |
-35.714 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.674 |
0.000 |
0.000 |
| y |
0.000 |
1.778 |
0.000 |
| z |
0.000 |
0.000 |
10.896 |
|
| Polar |
|---|
| 3z2-r2 | 21.792 |
|---|
| x2-y2 | -9.634 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.625 |
0.000 |
0.000 |
| y |
0.000 |
12.712 |
0.000 |
| z |
0.000 |
0.000 |
22.643 |
<r2> (average value of r
2) Å
2
| <r2> |
214.220 |
| (<r2>)1/2 |
14.636 |