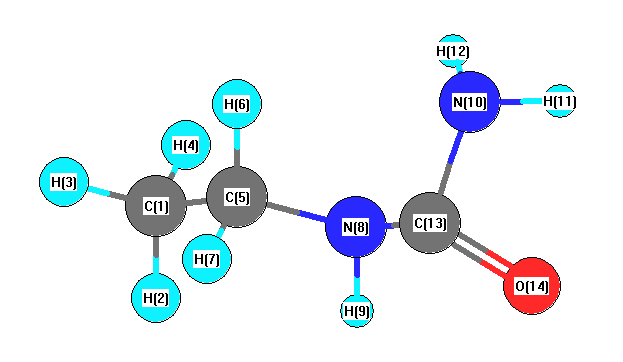
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3745 |
3650 |
23.64 |
|
|
|
| 2 |
A |
3613 |
3521 |
19.30 |
|
|
|
| 3 |
A |
3590 |
3498 |
22.85 |
|
|
|
| 4 |
A |
3093 |
3014 |
42.99 |
|
|
|
| 5 |
A |
3083 |
3005 |
53.59 |
|
|
|
| 6 |
A |
2996 |
2919 |
28.59 |
|
|
|
| 7 |
A |
2992 |
2916 |
47.18 |
|
|
|
| 8 |
A |
2956 |
2880 |
59.75 |
|
|
|
| 9 |
A |
1681 |
1638 |
375.66 |
|
|
|
| 10 |
A |
1631 |
1589 |
269.28 |
|
|
|
| 11 |
A |
1552 |
1513 |
6.93 |
|
|
|
| 12 |
A |
1525 |
1486 |
15.05 |
|
|
|
| 13 |
A |
1518 |
1479 |
9.44 |
|
|
|
| 14 |
A |
1461 |
1424 |
51.67 |
|
|
|
| 15 |
A |
1439 |
1402 |
7.64 |
|
|
|
| 16 |
A |
1423 |
1386 |
314.33 |
|
|
|
| 17 |
A |
1371 |
1336 |
25.25 |
|
|
|
| 18 |
A |
1303 |
1269 |
0.02 |
|
|
|
| 19 |
A |
1194 |
1163 |
16.22 |
|
|
|
| 20 |
A |
1177 |
1147 |
1.68 |
|
|
|
| 21 |
A |
1105 |
1077 |
17.61 |
|
|
|
| 22 |
A |
1056 |
1029 |
15.35 |
|
|
|
| 23 |
A |
985 |
960 |
0.48 |
|
|
|
| 24 |
A |
887 |
864 |
2.56 |
|
|
|
| 25 |
A |
837 |
816 |
0.83 |
|
|
|
| 26 |
A |
733 |
714 |
66.81 |
|
|
|
| 27 |
A |
551 |
537 |
7.60 |
|
|
|
| 28 |
A |
545 |
531 |
166.30 |
|
|
|
| 29 |
A |
521 |
508 |
17.18 |
|
|
|
| 30 |
A |
479 |
467 |
0.03 |
|
|
|
| 31 |
A |
377 |
368 |
228.49 |
|
|
|
| 32 |
A |
354 |
345 |
0.04 |
|
|
|
| 33 |
A |
260 |
254 |
0.00 |
|
|
|
| 34 |
A |
193 |
188 |
6.35 |
|
|
|
| 35 |
A |
100 |
98 |
1.11 |
|
|
|
| 36 |
A |
60 |
59 |
1.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26192.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 25524.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.537 |
|
|
|
| 2 |
H |
0.165 |
|
|
|
| 3 |
H |
0.161 |
|
|
|
| 4 |
H |
0.165 |
|
|
|
| 5 |
C |
-0.178 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
| 7 |
H |
0.151 |
|
|
|
| 8 |
N |
-0.265 |
|
|
|
| 9 |
H |
0.326 |
|
|
|
| 10 |
N |
-0.518 |
|
|
|
| 11 |
H |
0.330 |
|
|
|
| 12 |
H |
0.314 |
|
|
|
| 13 |
C |
-0.033 |
|
|
|
| 14 |
O |
-0.233 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.886 |
3.472 |
0.001 |
5.211 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.174 |
-8.783 |
-0.001 |
| y |
-8.783 |
-35.116 |
0.002 |
| z |
-0.001 |
0.002 |
-39.036 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.098 |
-8.783 |
-0.001 |
| y |
-8.783 |
4.489 |
0.002 |
| z |
-0.001 |
0.002 |
-1.391 |
|
| Polar |
|---|
| 3z2-r2 | -2.782 |
|---|
| x2-y2 | -5.058 |
|---|
| xy | -8.783 |
|---|
| xz | -0.001 |
|---|
| yz | 0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.677 |
0.348 |
0.000 |
| y |
0.348 |
8.142 |
-0.002 |
| z |
0.000 |
-0.002 |
5.572 |
<r2> (average value of r
2) Å
2
| <r2> |
170.813 |
| (<r2>)1/2 |
13.070 |