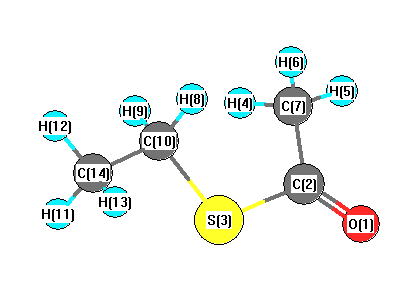
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3128 |
3048 |
14.95 |
|
|
|
| 2 |
A |
3114 |
3035 |
51.65 |
|
|
|
| 3 |
A |
3094 |
3015 |
8.57 |
|
|
|
| 4 |
A |
3086 |
3007 |
4.05 |
|
|
|
| 5 |
A |
3080 |
3001 |
36.52 |
|
|
|
| 6 |
A |
3030 |
2953 |
24.47 |
|
|
|
| 7 |
A |
3008 |
2931 |
4.08 |
|
|
|
| 8 |
A |
2996 |
2919 |
39.45 |
|
|
|
| 9 |
A |
1652 |
1610 |
298.60 |
|
|
|
| 10 |
A |
1525 |
1486 |
5.60 |
|
|
|
| 11 |
A |
1515 |
1477 |
12.91 |
|
|
|
| 12 |
A |
1498 |
1460 |
19.65 |
|
|
|
| 13 |
A |
1498 |
1460 |
1.25 |
|
|
|
| 14 |
A |
1484 |
1446 |
38.34 |
|
|
|
| 15 |
A |
1440 |
1403 |
9.54 |
|
|
|
| 16 |
A |
1415 |
1378 |
23.07 |
|
|
|
| 17 |
A |
1308 |
1274 |
36.75 |
|
|
|
| 18 |
A |
1274 |
1241 |
0.02 |
|
|
|
| 19 |
A |
1144 |
1115 |
134.63 |
|
|
|
| 20 |
A |
1082 |
1054 |
4.77 |
|
|
|
| 21 |
A |
1051 |
1024 |
0.34 |
|
|
|
| 22 |
A |
1043 |
1016 |
0.87 |
|
|
|
| 23 |
A |
996 |
970 |
9.08 |
|
|
|
| 24 |
A |
949 |
925 |
66.53 |
|
|
|
| 25 |
A |
802 |
781 |
6.64 |
|
|
|
| 26 |
A |
642 |
625 |
0.41 |
|
|
|
| 27 |
A |
601 |
586 |
78.75 |
|
|
|
| 28 |
A |
494 |
482 |
1.93 |
|
|
|
| 29 |
A |
431 |
420 |
3.93 |
|
|
|
| 30 |
A |
353 |
344 |
1.88 |
|
|
|
| 31 |
A |
290 |
282 |
0.99 |
|
|
|
| 32 |
A |
247 |
241 |
0.06 |
|
|
|
| 33 |
A |
148 |
144 |
1.13 |
|
|
|
| 34 |
A |
141 |
137 |
0.14 |
|
|
|
| 35 |
A |
71 |
69 |
0.62 |
|
|
|
| 36 |
A |
51 |
49 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24838.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 24205.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.135 |
|
|
|
| 2 |
C |
-0.120 |
|
|
|
| 3 |
S |
0.307 |
|
|
|
| 4 |
H |
0.172 |
|
|
|
| 5 |
H |
0.192 |
|
|
|
| 6 |
H |
0.173 |
|
|
|
| 7 |
C |
-0.491 |
|
|
|
| 8 |
H |
0.180 |
|
|
|
| 9 |
H |
0.180 |
|
|
|
| 10 |
C |
-0.509 |
|
|
|
| 11 |
H |
0.171 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.171 |
|
|
|
| 14 |
C |
-0.447 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.897 |
4.004 |
0.000 |
4.942 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.675 |
5.100 |
0.002 |
| y |
5.100 |
-46.358 |
0.000 |
| z |
0.002 |
0.000 |
-44.081 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.456 |
5.100 |
0.002 |
| y |
5.100 |
-0.480 |
0.000 |
| z |
0.002 |
0.000 |
2.936 |
|
| Polar |
|---|
| 3z2-r2 | 5.871 |
|---|
| x2-y2 | -1.318 |
|---|
| xy | 5.100 |
|---|
| xz | 0.002 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.462 |
-0.282 |
-0.003 |
| y |
-0.282 |
9.730 |
0.005 |
| z |
-0.003 |
0.005 |
6.963 |
<r2> (average value of r
2) Å
2
| <r2> |
208.047 |
| (<r2>)1/2 |
14.424 |