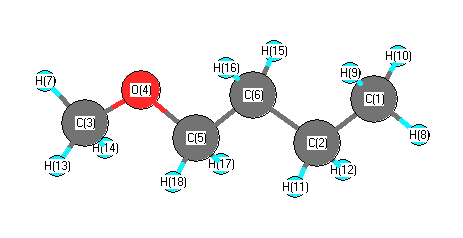
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3111 |
3031 |
41.21 |
|
|
|
| 2 |
A' |
3066 |
2988 |
68.08 |
|
|
|
| 3 |
A' |
2999 |
2923 |
50.72 |
|
|
|
| 4 |
A' |
2979 |
2903 |
28.37 |
|
|
|
| 5 |
A' |
2969 |
2893 |
65.30 |
|
|
|
| 6 |
A' |
2941 |
2866 |
101.41 |
|
|
|
| 7 |
A' |
2922 |
2848 |
41.68 |
|
|
|
| 8 |
A' |
1542 |
1503 |
6.33 |
|
|
|
| 9 |
A' |
1531 |
1492 |
3.69 |
|
|
|
| 10 |
A' |
1519 |
1481 |
2.49 |
|
|
|
| 11 |
A' |
1515 |
1476 |
0.07 |
|
|
|
| 12 |
A' |
1510 |
1471 |
11.93 |
|
|
|
| 13 |
A' |
1474 |
1436 |
0.13 |
|
|
|
| 14 |
A' |
1430 |
1394 |
2.33 |
|
|
|
| 15 |
A' |
1418 |
1382 |
26.91 |
|
|
|
| 16 |
A' |
1385 |
1349 |
0.05 |
|
|
|
| 17 |
A' |
1309 |
1275 |
0.43 |
|
|
|
| 18 |
A' |
1196 |
1166 |
16.63 |
|
|
|
| 19 |
A' |
1132 |
1103 |
13.46 |
|
|
|
| 20 |
A' |
1095 |
1067 |
127.90 |
|
|
|
| 21 |
A' |
1078 |
1051 |
9.25 |
|
|
|
| 22 |
A' |
1024 |
998 |
5.75 |
|
|
|
| 23 |
A' |
929 |
905 |
8.92 |
|
|
|
| 24 |
A' |
885 |
862 |
15.98 |
|
|
|
| 25 |
A' |
469 |
457 |
1.55 |
|
|
|
| 26 |
A' |
367 |
358 |
3.32 |
|
|
|
| 27 |
A' |
300 |
292 |
1.25 |
|
|
|
| 28 |
A' |
133 |
130 |
1.62 |
|
|
|
| 29 |
A" |
3068 |
2990 |
152.39 |
|
|
|
| 30 |
A" |
3052 |
2974 |
9.59 |
|
|
|
| 31 |
A" |
3010 |
2933 |
95.53 |
|
|
|
| 32 |
A" |
3005 |
2928 |
10.01 |
|
|
|
| 33 |
A" |
2970 |
2895 |
56.31 |
|
|
|
| 34 |
A" |
1521 |
1482 |
8.41 |
|
|
|
| 35 |
A" |
1495 |
1457 |
7.88 |
|
|
|
| 36 |
A" |
1320 |
1286 |
0.04 |
|
|
|
| 37 |
A" |
1307 |
1274 |
0.12 |
|
|
|
| 38 |
A" |
1245 |
1213 |
0.96 |
|
|
|
| 39 |
A" |
1187 |
1156 |
6.30 |
|
|
|
| 40 |
A" |
1144 |
1115 |
0.57 |
|
|
|
| 41 |
A" |
956 |
932 |
0.00 |
|
|
|
| 42 |
A" |
823 |
802 |
0.55 |
|
|
|
| 43 |
A" |
753 |
734 |
3.65 |
|
|
|
| 44 |
A" |
236 |
230 |
0.39 |
|
|
|
| 45 |
A" |
207 |
202 |
4.30 |
|
|
|
| 46 |
A" |
126 |
123 |
2.08 |
|
|
|
| 47 |
A" |
88 |
86 |
2.54 |
|
|
|
| 48 |
A" |
67 |
66 |
1.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35902.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 34986.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.473 |
|
|
|
| 2 |
C |
-0.204 |
|
|
|
| 3 |
C |
-0.297 |
|
|
|
| 4 |
O |
-0.265 |
|
|
|
| 5 |
C |
-0.159 |
|
|
|
| 6 |
C |
-0.200 |
|
|
|
| 7 |
H |
0.169 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.142 |
|
|
|
| 10 |
H |
0.142 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.122 |
|
|
|
| 13 |
H |
0.132 |
|
|
|
| 14 |
H |
0.132 |
|
|
|
| 15 |
H |
0.135 |
|
|
|
| 16 |
H |
0.135 |
|
|
|
| 17 |
H |
0.111 |
|
|
|
| 18 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.284 |
-0.946 |
0.000 |
1.595 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.583 |
1.396 |
0.000 |
| y |
1.396 |
-35.651 |
0.000 |
| z |
0.000 |
0.000 |
-40.056 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.729 |
1.396 |
0.000 |
| y |
1.396 |
5.669 |
0.000 |
| z |
0.000 |
0.000 |
-0.939 |
|
| Polar |
|---|
| 3z2-r2 | -1.879 |
|---|
| x2-y2 | -6.932 |
|---|
| xy | 1.396 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.604 |
1.360 |
0.000 |
| y |
1.360 |
10.998 |
0.000 |
| z |
0.000 |
0.000 |
8.489 |
<r2> (average value of r
2) Å
2
| <r2> |
247.600 |
| (<r2>)1/2 |
15.735 |