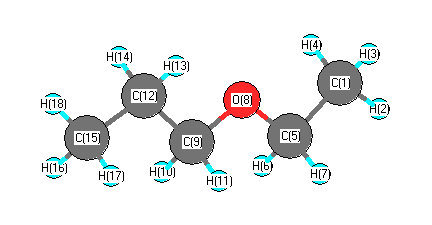
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3085 |
3006 |
49.18 |
|
|
|
| 2 |
A' |
3070 |
2992 |
61.80 |
|
|
|
| 3 |
A' |
3007 |
2930 |
29.45 |
|
|
|
| 4 |
A' |
2998 |
2922 |
30.31 |
|
|
|
| 5 |
A' |
2978 |
2902 |
55.66 |
|
|
|
| 6 |
A' |
2937 |
2862 |
117.79 |
|
|
|
| 7 |
A' |
2921 |
2847 |
23.20 |
|
|
|
| 8 |
A' |
1545 |
1506 |
4.73 |
|
|
|
| 9 |
A' |
1535 |
1496 |
4.76 |
|
|
|
| 10 |
A' |
1526 |
1487 |
4.08 |
|
|
|
| 11 |
A' |
1520 |
1481 |
1.87 |
|
|
|
| 12 |
A' |
1515 |
1477 |
7.51 |
|
|
|
| 13 |
A' |
1445 |
1408 |
3.00 |
|
|
|
| 14 |
A' |
1432 |
1396 |
15.64 |
|
|
|
| 15 |
A' |
1426 |
1390 |
2.60 |
|
|
|
| 16 |
A' |
1386 |
1351 |
33.06 |
|
|
|
| 17 |
A' |
1337 |
1303 |
13.14 |
|
|
|
| 18 |
A' |
1164 |
1135 |
3.85 |
|
|
|
| 19 |
A' |
1120 |
1091 |
23.95 |
|
|
|
| 20 |
A' |
1096 |
1068 |
180.64 |
|
|
|
| 21 |
A' |
1056 |
1029 |
1.99 |
|
|
|
| 22 |
A' |
1008 |
982 |
18.89 |
|
|
|
| 23 |
A' |
906 |
883 |
4.36 |
|
|
|
| 24 |
A' |
862 |
840 |
11.13 |
|
|
|
| 25 |
A' |
475 |
463 |
3.75 |
|
|
|
| 26 |
A' |
384 |
375 |
1.13 |
|
|
|
| 27 |
A' |
296 |
288 |
0.99 |
|
|
|
| 28 |
A' |
132 |
129 |
0.64 |
|
|
|
| 29 |
A" |
3099 |
3020 |
53.16 |
|
|
|
| 30 |
A" |
3073 |
2995 |
112.73 |
|
|
|
| 31 |
A" |
3043 |
2965 |
0.59 |
|
|
|
| 32 |
A" |
2980 |
2904 |
89.69 |
|
|
|
| 33 |
A" |
2970 |
2894 |
46.26 |
|
|
|
| 34 |
A" |
1524 |
1485 |
8.75 |
|
|
|
| 35 |
A" |
1502 |
1463 |
7.29 |
|
|
|
| 36 |
A" |
1311 |
1278 |
0.05 |
|
|
|
| 37 |
A" |
1284 |
1252 |
0.82 |
|
|
|
| 38 |
A" |
1258 |
1226 |
0.85 |
|
|
|
| 39 |
A" |
1188 |
1158 |
9.31 |
|
|
|
| 40 |
A" |
1164 |
1134 |
0.90 |
|
|
|
| 41 |
A" |
907 |
884 |
1.44 |
|
|
|
| 42 |
A" |
828 |
807 |
1.09 |
|
|
|
| 43 |
A" |
775 |
755 |
2.15 |
|
|
|
| 44 |
A" |
235 |
229 |
0.97 |
|
|
|
| 45 |
A" |
224 |
219 |
0.20 |
|
|
|
| 46 |
A" |
137 |
134 |
5.71 |
|
|
|
| 47 |
A" |
83 |
81 |
2.09 |
|
|
|
| 48 |
A" |
62 |
60 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35903.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 34988.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.447 |
|
|
|
| 2 |
H |
0.135 |
|
|
|
| 3 |
H |
0.157 |
|
|
|
| 4 |
H |
0.157 |
|
|
|
| 5 |
C |
-0.091 |
|
|
|
| 6 |
H |
0.115 |
|
|
|
| 7 |
H |
0.115 |
|
|
|
| 8 |
O |
-0.272 |
|
|
|
| 9 |
C |
-0.140 |
|
|
|
| 10 |
H |
0.109 |
|
|
|
| 11 |
H |
0.109 |
|
|
|
| 12 |
C |
-0.181 |
|
|
|
| 13 |
H |
0.142 |
|
|
|
| 14 |
H |
0.142 |
|
|
|
| 15 |
C |
-0.483 |
|
|
|
| 16 |
H |
0.140 |
|
|
|
| 17 |
H |
0.140 |
|
|
|
| 18 |
H |
0.154 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.247 |
-0.678 |
0.000 |
1.419 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.018 |
2.458 |
0.000 |
| y |
2.458 |
-38.950 |
0.000 |
| z |
0.000 |
0.000 |
-39.921 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.418 |
2.458 |
0.000 |
| y |
2.458 |
0.519 |
0.000 |
| z |
0.000 |
0.000 |
-0.937 |
|
| Polar |
|---|
| 3z2-r2 | -1.874 |
|---|
| x2-y2 | -0.067 |
|---|
| xy | 2.458 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.451 |
1.168 |
0.000 |
| y |
1.168 |
11.349 |
0.000 |
| z |
0.000 |
0.000 |
8.558 |
<r2> (average value of r
2) Å
2
| <r2> |
241.886 |
| (<r2>)1/2 |
15.553 |