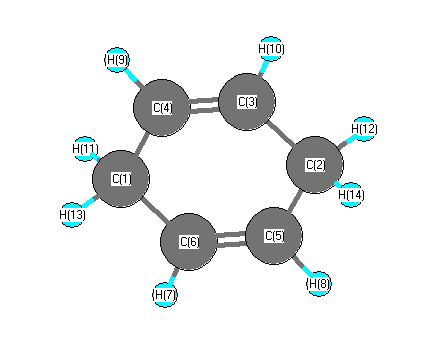
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3138 |
3058 |
0.00 |
|
|
|
| 2 |
Ag |
2948 |
2872 |
0.00 |
|
|
|
| 3 |
Ag |
1721 |
1677 |
0.00 |
|
|
|
| 4 |
Ag |
1501 |
1463 |
0.00 |
|
|
|
| 5 |
Ag |
1230 |
1198 |
0.00 |
|
|
|
| 6 |
Ag |
842 |
820 |
0.00 |
|
|
|
| 7 |
Ag |
539 |
525 |
0.00 |
|
|
|
| 8 |
Au |
1214 |
1183 |
0.00 |
|
|
|
| 9 |
Au |
1004 |
978 |
0.00 |
|
|
|
| 10 |
Au |
375 |
365 |
0.00 |
|
|
|
| 11 |
B1g |
3104 |
3025 |
0.00 |
|
|
|
| 12 |
B1g |
1396 |
1361 |
0.00 |
|
|
|
| 13 |
B1g |
1373 |
1338 |
0.00 |
|
|
|
| 14 |
B1g |
1060 |
1033 |
0.00 |
|
|
|
| 15 |
B1g |
574 |
559 |
0.00 |
|
|
|
| 16 |
B1u |
2966 |
2891 |
85.63 |
|
|
|
| 17 |
B1u |
1002 |
976 |
29.72 |
|
|
|
| 18 |
B1u |
648 |
631 |
86.36 |
|
|
|
| 19 |
B1u |
113 |
110 |
1.46 |
|
|
|
| 20 |
B2g |
2967 |
2892 |
0.00 |
|
|
|
| 21 |
B2g |
1031 |
1005 |
0.00 |
|
|
|
| 22 |
B2g |
975 |
950 |
0.00 |
|
|
|
| 23 |
B2g |
402 |
392 |
0.00 |
|
|
|
| 24 |
B2u |
3133 |
3053 |
136.44 |
|
|
|
| 25 |
B2u |
1678 |
1636 |
8.13 |
|
|
|
| 26 |
B2u |
1383 |
1348 |
0.39 |
|
|
|
| 27 |
B2u |
1186 |
1156 |
0.05 |
|
|
|
| 28 |
B2u |
941 |
917 |
1.06 |
|
|
|
| 29 |
B3g |
1220 |
1189 |
0.00 |
|
|
|
| 30 |
B3g |
729 |
711 |
0.00 |
|
|
|
| 31 |
B3u |
3105 |
3026 |
30.83 |
|
|
|
| 32 |
B3u |
2949 |
2873 |
134.24 |
|
|
|
| 33 |
B3u |
1506 |
1467 |
8.08 |
|
|
|
| 34 |
B3u |
1433 |
1397 |
0.04 |
|
|
|
| 35 |
B3u |
967 |
943 |
0.66 |
|
|
|
| 36 |
B3u |
885 |
863 |
12.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26618.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 25939.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.450 |
|
|
|
| 2 |
C |
-0.450 |
|
|
|
| 3 |
C |
-0.049 |
|
|
|
| 4 |
C |
-0.049 |
|
|
|
| 5 |
C |
-0.049 |
|
|
|
| 6 |
C |
-0.049 |
|
|
|
| 7 |
H |
0.127 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
H |
0.127 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
| 12 |
H |
0.147 |
|
|
|
| 13 |
H |
0.147 |
|
|
|
| 14 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.964 |
0.000 |
0.000 |
| y |
0.000 |
-33.561 |
0.000 |
| z |
0.000 |
0.000 |
-39.821 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.726 |
0.000 |
0.000 |
| y |
0.000 |
4.332 |
0.000 |
| z |
0.000 |
0.000 |
-5.059 |
|
| Polar |
|---|
| 3z2-r2 | -10.117 |
|---|
| x2-y2 | -2.404 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.199 |
0.000 |
0.000 |
| y |
0.000 |
13.048 |
0.000 |
| z |
0.000 |
0.000 |
6.014 |
<r2> (average value of r
2) Å
2
| <r2> |
122.364 |
| (<r2>)1/2 |
11.062 |