Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -167.465828 |
| Energy at 298.15K | |
| HF Energy | -167.465828 |
| Nuclear repulsion energy | 294.770123 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1779 |
1734 |
0.00 |
|
|
|
| 2 |
Ag |
1354 |
1320 |
0.00 |
|
|
|
| 3 |
Ag |
1225 |
1194 |
0.00 |
|
|
|
| 4 |
Ag |
1102 |
1074 |
0.00 |
|
|
|
| 5 |
Ag |
658 |
641 |
0.00 |
|
|
|
| 6 |
Ag |
543 |
529 |
0.00 |
|
|
|
| 7 |
Ag |
357 |
348 |
0.00 |
|
|
|
| 8 |
Ag |
325 |
317 |
0.00 |
|
|
|
| 9 |
Ag |
209 |
204 |
0.00 |
|
|
|
| 10 |
Au |
492 |
479 |
1.17 |
|
|
|
| 11 |
Au |
309 |
301 |
9.42 |
|
|
|
| 12 |
Au |
107 |
104 |
0.35 |
|
|
|
| 13 |
Au |
32i |
31i |
0.10 |
|
|
|
| 14 |
Bg |
575 |
560 |
0.00 |
|
|
|
| 15 |
Bg |
411 |
401 |
0.00 |
|
|
|
| 16 |
Bg |
162 |
158 |
0.00 |
|
|
|
| 17 |
Bu |
1725 |
1681 |
305.55 |
|
|
|
| 18 |
Bu |
1238 |
1206 |
359.31 |
|
|
|
| 19 |
Bu |
1123 |
1095 |
190.74 |
|
|
|
| 20 |
Bu |
886 |
863 |
245.48 |
|
|
|
| 21 |
Bu |
572 |
557 |
3.63 |
|
|
|
| 22 |
Bu |
447 |
436 |
7.51 |
|
|
|
| 23 |
Bu |
277 |
270 |
6.86 |
|
|
|
| 24 |
Bu |
129 |
126 |
1.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7986.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 7782.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.155 |
1.889 |
0.000 |
| C2 |
0.461 |
0.569 |
0.000 |
| C3 |
-0.461 |
-0.569 |
0.000 |
| C4 |
-0.155 |
-1.889 |
0.000 |
| F5 |
1.110 |
2.871 |
0.000 |
| F6 |
-1.110 |
2.409 |
0.000 |
| F7 |
1.821 |
0.234 |
0.000 |
| F8 |
-1.821 |
-0.234 |
0.000 |
| F9 |
1.110 |
-2.409 |
0.000 |
| F10 |
-1.110 |
-2.871 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3545 | 2.5345 | 3.7906 | 1.3703 | 1.3684 | 2.3485 | 2.9005 | 4.4030 | 4.9257 |
C2 | 1.3545 | | 1.4658 | 2.5345 | 2.3917 | 2.4197 | 1.4008 | 2.4199 | 3.0486 | 3.7829 | C3 | 2.5345 | 1.4658 | | 1.3545 | 3.7829 | 3.0486 | 2.4199 | 1.4008 | 2.4197 | 2.3917 | C4 | 3.7906 | 2.5345 | 1.3545 | | 4.9257 | 4.4030 | 2.9005 | 2.3485 | 1.3684 | 1.3703 | F5 | 1.3703 | 2.3917 | 3.7829 | 4.9257 | | 2.2684 | 2.7318 | 4.2706 | 5.2807 | 6.1573 | F6 | 1.3684 | 2.4197 | 3.0486 | 4.4030 | 2.2684 | | 3.6507 | 2.7370 | 5.3056 | 5.2807 | F7 | 2.3485 | 1.4008 | 2.4199 | 2.9005 | 2.7318 | 3.6507 | | 3.6726 | 2.7370 | 4.2706 | F8 | 2.9005 | 2.4199 | 1.4008 | 2.3485 | 4.2706 | 2.7370 | 3.6726 | | 3.6507 | 2.7318 | F9 | 4.4030 | 3.0486 | 2.4197 | 1.3684 | 5.2807 | 5.3056 | 2.7370 | 3.6507 | | 2.2684 | F10 | 4.9257 | 3.7829 | 2.3917 | 1.3703 | 6.1573 | 5.2807 | 4.2706 | 2.7318 | 2.2684 | |
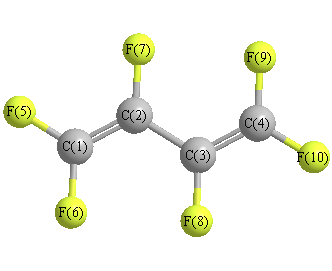 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.926 |
|
C1 |
C2 |
F7 |
116.925 |
| C2 |
C1 |
F5 |
122.748 |
|
C2 |
C1 |
F6 |
125.408 |
| C2 |
C3 |
C4 |
127.926 |
|
C2 |
C3 |
F8 |
115.149 |
| C3 |
C2 |
F7 |
115.149 |
|
C3 |
C4 |
F9 |
125.408 |
| C3 |
C4 |
F10 |
122.748 |
|
C4 |
C3 |
F8 |
116.925 |
| F5 |
C1 |
F6 |
111.843 |
|
F9 |
C4 |
F10 |
111.843 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.122 |
|
|
|
| 2 |
C |
0.271 |
|
|
|
| 3 |
C |
0.271 |
|
|
|
| 4 |
C |
0.122 |
|
|
|
| 5 |
F |
-0.120 |
|
|
|
| 6 |
F |
-0.114 |
|
|
|
| 7 |
F |
-0.159 |
|
|
|
| 8 |
F |
-0.159 |
|
|
|
| 9 |
F |
-0.114 |
|
|
|
| 10 |
F |
-0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-60.230 |
-0.336 |
0.000 |
| y |
-0.336 |
-58.067 |
0.000 |
| z |
0.000 |
0.000 |
-49.910 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.242 |
-0.336 |
0.000 |
| y |
-0.336 |
-2.997 |
0.000 |
| z |
0.000 |
0.000 |
9.239 |
|
| Polar |
|---|
| 3z2-r2 | 18.478 |
|---|
| x2-y2 | -2.163 |
|---|
| xy | -0.336 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.809 |
-0.207 |
0.000 |
| y |
-0.207 |
13.048 |
0.000 |
| z |
0.000 |
0.000 |
3.520 |
<r2> (average value of r
2) Å
2
| <r2> |
346.478 |
| (<r2>)1/2 |
18.614 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -167.467689 |
| Energy at 298.15K | -167.468387 |
| HF Energy | -167.467689 |
| Nuclear repulsion energy | 296.377455 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
1786 |
1740 |
68.37 |
|
|
|
| 2 |
A |
1325 |
1291 |
23.25 |
|
|
|
| 3 |
A |
1236 |
1205 |
122.07 |
|
|
|
| 4 |
A |
1057 |
1030 |
232.81 |
|
|
|
| 5 |
A |
656 |
640 |
0.05 |
|
|
|
| 6 |
A |
614 |
599 |
1.05 |
|
|
|
| 7 |
A |
476 |
463 |
0.03 |
|
|
|
| 8 |
A |
425 |
414 |
0.83 |
|
|
|
| 9 |
A |
354 |
345 |
2.03 |
|
|
|
| 10 |
A |
246 |
240 |
0.14 |
|
|
|
| 11 |
A |
170 |
166 |
0.54 |
|
|
|
| 12 |
A |
90 |
88 |
0.63 |
|
|
|
| 13 |
A |
44 |
43 |
0.00 |
|
|
|
| 14 |
B |
1744 |
1700 |
223.18 |
|
|
|
| 15 |
B |
1232 |
1201 |
226.51 |
|
|
|
| 16 |
B |
1114 |
1085 |
87.61 |
|
|
|
| 17 |
B |
902 |
879 |
169.04 |
|
|
|
| 18 |
B |
576 |
561 |
4.76 |
|
|
|
| 19 |
B |
562 |
548 |
1.09 |
|
|
|
| 20 |
B |
502 |
490 |
6.74 |
|
|
|
| 21 |
B |
379 |
369 |
7.14 |
|
|
|
| 22 |
B |
275 |
268 |
6.20 |
|
|
|
| 23 |
B |
194 |
189 |
5.06 |
|
|
|
| 24 |
B |
102 |
99 |
0.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8029.8 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 7825.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.189 |
1.598 |
-0.376 |
| C2 |
0.140 |
0.718 |
0.594 |
| C3 |
-0.140 |
-0.718 |
0.594 |
| C4 |
0.189 |
-1.598 |
-0.376 |
| F5 |
0.140 |
2.923 |
-0.363 |
| F6 |
-0.883 |
1.254 |
-1.509 |
| F7 |
0.808 |
1.210 |
1.726 |
| F8 |
-0.808 |
-1.210 |
1.726 |
| F9 |
0.883 |
-1.254 |
-1.509 |
| F10 |
-0.140 |
-2.923 |
-0.363 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3501 | 2.5113 | 3.2181 | 1.3653 | 1.3720 | 2.3588 | 3.5619 | 3.2509 | 4.5210 |
C2 | 1.3501 | | 1.4632 | 2.5113 | 2.4032 | 2.3989 | 1.4040 | 2.4289 | 2.9773 | 3.7749 | C3 | 2.5113 | 1.4632 | | 1.3501 | 3.7749 | 2.9773 | 2.4289 | 1.4040 | 2.3989 | 2.4032 | C4 | 3.2181 | 2.5113 | 1.3501 | | 4.5210 | 3.2509 | 3.5619 | 2.3588 | 1.3720 | 1.3653 | F5 | 1.3653 | 2.4032 | 3.7749 | 4.5210 | | 2.2679 | 2.7825 | 4.7270 | 4.3950 | 5.8524 | F6 | 1.3720 | 2.3989 | 2.9773 | 3.2509 | 2.2679 | | 3.6506 | 4.0677 | 3.0676 | 4.3950 | F7 | 2.3588 | 1.4040 | 2.4289 | 3.5619 | 2.7825 | 3.6506 | | 2.9107 | 4.0677 | 4.7270 | F8 | 3.5619 | 2.4289 | 1.4040 | 2.3588 | 4.7270 | 4.0677 | 2.9107 | | 3.6506 | 2.7825 | F9 | 3.2509 | 2.9773 | 2.3989 | 1.3720 | 4.3950 | 3.0676 | 4.0677 | 3.6506 | | 2.2679 | F10 | 4.5210 | 3.7749 | 2.4032 | 1.3653 | 5.8524 | 4.3950 | 4.7270 | 2.7825 | 2.2679 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.371 |
|
C1 |
C2 |
F7 |
117.838 |
| C2 |
C1 |
F5 |
124.510 |
|
C2 |
C1 |
F6 |
123.592 |
| C2 |
C3 |
C4 |
126.371 |
|
C2 |
C3 |
F8 |
115.789 |
| C3 |
C2 |
F7 |
115.789 |
|
C3 |
C4 |
F9 |
123.592 |
| C3 |
C4 |
F10 |
124.510 |
|
C4 |
C3 |
F8 |
117.838 |
| F5 |
C1 |
F6 |
111.892 |
|
F9 |
C4 |
F10 |
111.892 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.253 |
|
|
|
| 2 |
C |
0.156 |
|
|
|
| 3 |
C |
0.156 |
|
|
|
| 4 |
C |
0.253 |
|
|
|
| 5 |
F |
-0.116 |
|
|
|
| 6 |
F |
-0.127 |
|
|
|
| 7 |
F |
-0.166 |
|
|
|
| 8 |
F |
-0.166 |
|
|
|
| 9 |
F |
-0.127 |
|
|
|
| 10 |
F |
-0.116 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.782 |
0.782 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-52.398 |
-1.932 |
0.000 |
| y |
-1.932 |
-57.619 |
0.000 |
| z |
0.000 |
0.000 |
-58.355 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.590 |
-1.932 |
0.000 |
| y |
-1.932 |
-2.243 |
0.000 |
| z |
0.000 |
0.000 |
-3.347 |
|
| Polar |
|---|
| 3z2-r2 | -6.694 |
|---|
| x2-y2 | 5.222 |
|---|
| xy | -1.932 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.514 |
-0.085 |
0.000 |
| y |
-0.085 |
9.860 |
0.000 |
| z |
0.000 |
0.000 |
7.450 |
<r2> (average value of r
2) Å
2
| <r2> |
321.896 |
| (<r2>)1/2 |
17.941 |
