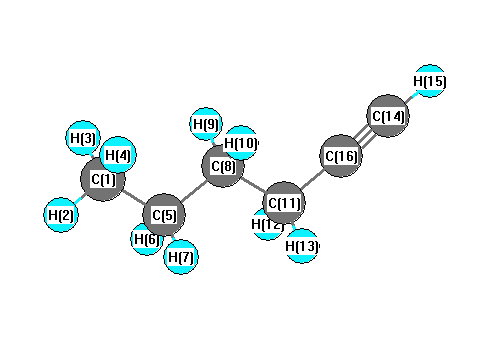
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3450 |
3362 |
58.12 |
|
|
|
| 2 |
A' |
3067 |
2989 |
66.84 |
|
|
|
| 3 |
A' |
2995 |
2919 |
78.41 |
|
|
|
| 4 |
A' |
2983 |
2907 |
21.83 |
|
|
|
| 5 |
A' |
2979 |
2903 |
27.11 |
|
|
|
| 6 |
A' |
2969 |
2893 |
37.11 |
|
|
|
| 7 |
A' |
2162 |
2107 |
7.62 |
|
|
|
| 8 |
A' |
1534 |
1495 |
10.94 |
|
|
|
| 9 |
A' |
1521 |
1483 |
1.57 |
|
|
|
| 10 |
A' |
1512 |
1474 |
0.80 |
|
|
|
| 11 |
A' |
1507 |
1468 |
1.52 |
|
|
|
| 12 |
A' |
1430 |
1394 |
4.65 |
|
|
|
| 13 |
A' |
1391 |
1356 |
1.06 |
|
|
|
| 14 |
A' |
1362 |
1327 |
3.15 |
|
|
|
| 15 |
A' |
1291 |
1258 |
1.99 |
|
|
|
| 16 |
A' |
1125 |
1096 |
4.95 |
|
|
|
| 17 |
A' |
1062 |
1035 |
0.09 |
|
|
|
| 18 |
A' |
1020 |
994 |
0.45 |
|
|
|
| 19 |
A' |
939 |
915 |
0.68 |
|
|
|
| 20 |
A' |
897 |
874 |
5.05 |
|
|
|
| 21 |
A' |
796 |
775 |
79.47 |
|
|
|
| 22 |
A' |
547 |
533 |
21.07 |
|
|
|
| 23 |
A' |
376 |
367 |
0.11 |
|
|
|
| 24 |
A' |
286 |
279 |
1.46 |
|
|
|
| 25 |
A' |
125 |
122 |
0.05 |
|
|
|
| 26 |
A" |
3068 |
2990 |
143.94 |
|
|
|
| 27 |
A" |
3051 |
2973 |
16.21 |
|
|
|
| 28 |
A" |
3018 |
2941 |
1.61 |
|
|
|
| 29 |
A" |
3005 |
2928 |
9.51 |
|
|
|
| 30 |
A" |
1520 |
1481 |
8.63 |
|
|
|
| 31 |
A" |
1324 |
1291 |
0.04 |
|
|
|
| 32 |
A" |
1313 |
1279 |
0.97 |
|
|
|
| 33 |
A" |
1245 |
1213 |
0.19 |
|
|
|
| 34 |
A" |
1153 |
1123 |
0.13 |
|
|
|
| 35 |
A" |
944 |
920 |
1.59 |
|
|
|
| 36 |
A" |
809 |
788 |
12.55 |
|
|
|
| 37 |
A" |
785 |
765 |
74.80 |
|
|
|
| 38 |
A" |
740 |
722 |
2.23 |
|
|
|
| 39 |
A" |
424 |
413 |
15.51 |
|
|
|
| 40 |
A" |
224 |
218 |
0.02 |
|
|
|
| 41 |
A" |
107 |
104 |
0.00 |
|
|
|
| 42 |
A" |
83 |
81 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31067.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30275.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.473 |
|
|
|
| 2 |
H |
0.148 |
|
|
|
| 3 |
H |
0.143 |
|
|
|
| 4 |
H |
0.143 |
|
|
|
| 5 |
C |
-0.175 |
|
|
|
| 6 |
H |
0.124 |
|
|
|
| 7 |
H |
0.124 |
|
|
|
| 8 |
C |
-0.274 |
|
|
|
| 9 |
H |
0.134 |
|
|
|
| 10 |
H |
0.134 |
|
|
|
| 11 |
C |
-0.180 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
| 14 |
C |
-2.034 |
|
|
|
| 15 |
H |
0.307 |
|
|
|
| 16 |
C |
1.566 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.838 |
0.452 |
0.000 |
0.952 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.772 |
0.212 |
0.000 |
| y |
0.212 |
-40.538 |
0.000 |
| z |
0.000 |
0.000 |
-39.312 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.153 |
0.212 |
0.000 |
| y |
0.212 |
-2.496 |
0.000 |
| z |
0.000 |
0.000 |
-0.657 |
|
| Polar |
|---|
| 3z2-r2 | -1.314 |
|---|
| x2-y2 | 3.766 |
|---|
| xy | 0.212 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.261 |
2.367 |
0.000 |
| y |
2.367 |
9.683 |
0.000 |
| z |
0.000 |
0.000 |
7.570 |
<r2> (average value of r
2) Å
2
| <r2> |
215.385 |
| (<r2>)1/2 |
14.676 |