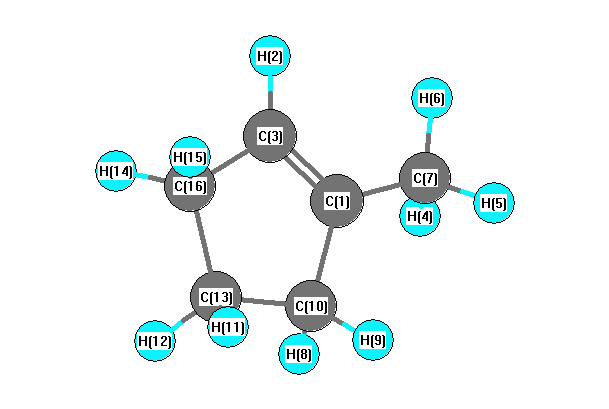
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3148 |
3068 |
30.90 |
|
|
|
| 2 |
A |
3075 |
2997 |
38.70 |
|
|
|
| 3 |
A |
3066 |
2988 |
81.49 |
|
|
|
| 4 |
A |
3029 |
2951 |
48.74 |
|
|
|
| 5 |
A |
3023 |
2946 |
78.11 |
|
|
|
| 6 |
A |
3013 |
2936 |
40.89 |
|
|
|
| 7 |
A |
2998 |
2921 |
21.28 |
|
|
|
| 8 |
A |
2968 |
2893 |
73.75 |
|
|
|
| 9 |
A |
2963 |
2887 |
81.82 |
|
|
|
| 10 |
A |
2959 |
2883 |
50.49 |
|
|
|
| 11 |
A |
1687 |
1644 |
2.99 |
|
|
|
| 12 |
A |
1529 |
1490 |
2.09 |
|
|
|
| 13 |
A |
1515 |
1477 |
6.50 |
|
|
|
| 14 |
A |
1511 |
1472 |
3.99 |
|
|
|
| 15 |
A |
1508 |
1469 |
1.02 |
|
|
|
| 16 |
A |
1504 |
1466 |
11.00 |
|
|
|
| 17 |
A |
1434 |
1397 |
3.47 |
|
|
|
| 18 |
A |
1360 |
1326 |
1.99 |
|
|
|
| 19 |
A |
1327 |
1294 |
2.00 |
|
|
|
| 20 |
A |
1317 |
1283 |
1.86 |
|
|
|
| 21 |
A |
1295 |
1262 |
0.85 |
|
|
|
| 22 |
A |
1245 |
1213 |
1.58 |
|
|
|
| 23 |
A |
1229 |
1198 |
1.76 |
|
|
|
| 24 |
A |
1173 |
1143 |
1.06 |
|
|
|
| 25 |
A |
1156 |
1126 |
0.82 |
|
|
|
| 26 |
A |
1101 |
1073 |
0.94 |
|
|
|
| 27 |
A |
1047 |
1021 |
2.50 |
|
|
|
| 28 |
A |
1033 |
1007 |
7.42 |
|
|
|
| 29 |
A |
1032 |
1006 |
4.65 |
|
|
|
| 30 |
A |
936 |
912 |
3.09 |
|
|
|
| 31 |
A |
913 |
890 |
0.67 |
|
|
|
| 32 |
A |
900 |
877 |
1.88 |
|
|
|
| 33 |
A |
864 |
842 |
0.17 |
|
|
|
| 34 |
A |
837 |
815 |
11.80 |
|
|
|
| 35 |
A |
805 |
784 |
8.33 |
|
|
|
| 36 |
A |
659 |
642 |
2.02 |
|
|
|
| 37 |
A |
573 |
559 |
0.07 |
|
|
|
| 38 |
A |
434 |
423 |
4.63 |
|
|
|
| 39 |
A |
316 |
307 |
0.60 |
|
|
|
| 40 |
A |
238 |
232 |
3.12 |
|
|
|
| 41 |
A |
172 |
168 |
0.70 |
|
|
|
| 42 |
A |
126 |
123 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31507.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30704.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.374 |
|
|
|
| 2 |
H |
0.156 |
|
|
|
| 3 |
C |
-0.201 |
|
|
|
| 4 |
H |
0.152 |
|
|
|
| 5 |
H |
0.153 |
|
|
|
| 6 |
H |
0.163 |
|
|
|
| 7 |
C |
-0.633 |
|
|
|
| 8 |
H |
0.136 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
| 10 |
C |
-0.373 |
|
|
|
| 11 |
H |
0.142 |
|
|
|
| 12 |
H |
0.128 |
|
|
|
| 13 |
C |
-0.276 |
|
|
|
| 14 |
H |
0.137 |
|
|
|
| 15 |
H |
0.134 |
|
|
|
| 16 |
C |
-0.326 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.131 |
-0.236 |
0.072 |
0.279 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.666 |
0.162 |
0.069 |
| y |
0.162 |
-38.047 |
-0.043 |
| z |
0.069 |
-0.043 |
-39.410 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.062 |
0.162 |
0.069 |
| y |
0.162 |
0.491 |
-0.043 |
| z |
0.069 |
-0.043 |
-1.554 |
|
| Polar |
|---|
| 3z2-r2 | -3.107 |
|---|
| x2-y2 | 0.381 |
|---|
| xy | 0.162 |
|---|
| xz | 0.069 |
|---|
| yz | -0.043 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.591 |
1.017 |
0.083 |
| y |
1.017 |
10.812 |
-0.114 |
| z |
0.083 |
-0.114 |
7.705 |
<r2> (average value of r
2) Å
2
| <r2> |
145.604 |
| (<r2>)1/2 |
12.067 |