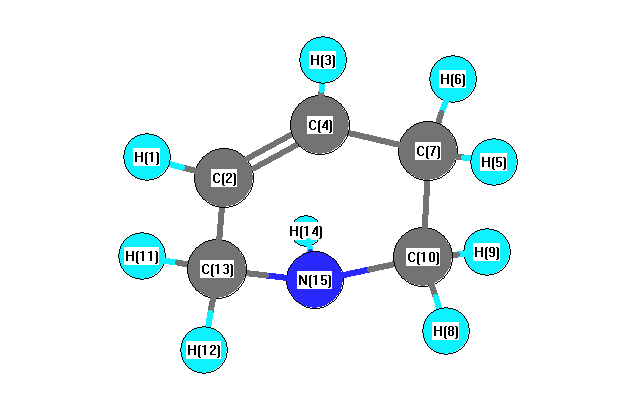
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3481 |
3392 |
4.28 |
|
|
|
| 2 |
A |
3131 |
3052 |
65.30 |
|
|
|
| 3 |
A |
3100 |
3021 |
17.69 |
|
|
|
| 4 |
A |
3059 |
2981 |
61.90 |
|
|
|
| 5 |
A |
3025 |
2948 |
41.21 |
|
|
|
| 6 |
A |
3009 |
2932 |
100.49 |
|
|
|
| 7 |
A |
2990 |
2914 |
19.46 |
|
|
|
| 8 |
A |
2958 |
2882 |
75.26 |
|
|
|
| 9 |
A |
2955 |
2880 |
53.15 |
|
|
|
| 10 |
A |
1682 |
1639 |
2.76 |
|
|
|
| 11 |
A |
1510 |
1472 |
3.79 |
|
|
|
| 12 |
A |
1505 |
1467 |
4.97 |
|
|
|
| 13 |
A |
1501 |
1463 |
5.08 |
|
|
|
| 14 |
A |
1487 |
1450 |
7.99 |
|
|
|
| 15 |
A |
1413 |
1377 |
0.75 |
|
|
|
| 16 |
A |
1385 |
1349 |
3.00 |
|
|
|
| 17 |
A |
1361 |
1327 |
1.44 |
|
|
|
| 18 |
A |
1361 |
1326 |
4.85 |
|
|
|
| 19 |
A |
1321 |
1288 |
0.10 |
|
|
|
| 20 |
A |
1251 |
1219 |
3.57 |
|
|
|
| 21 |
A |
1219 |
1188 |
11.43 |
|
|
|
| 22 |
A |
1195 |
1165 |
7.03 |
|
|
|
| 23 |
A |
1127 |
1098 |
21.42 |
|
|
|
| 24 |
A |
1105 |
1076 |
1.33 |
|
|
|
| 25 |
A |
1043 |
1016 |
3.81 |
|
|
|
| 26 |
A |
1005 |
980 |
3.29 |
|
|
|
| 27 |
A |
1004 |
978 |
0.11 |
|
|
|
| 28 |
A |
975 |
950 |
7.49 |
|
|
|
| 29 |
A |
901 |
878 |
3.62 |
|
|
|
| 30 |
A |
879 |
857 |
4.59 |
|
|
|
| 31 |
A |
837 |
816 |
10.60 |
|
|
|
| 32 |
A |
769 |
749 |
14.45 |
|
|
|
| 33 |
A |
682 |
664 |
110.60 |
|
|
|
| 34 |
A |
633 |
616 |
102.88 |
|
|
|
| 35 |
A |
520 |
507 |
4.54 |
|
|
|
| 36 |
A |
480 |
467 |
0.65 |
|
|
|
| 37 |
A |
386 |
376 |
1.99 |
|
|
|
| 38 |
A |
275 |
268 |
15.13 |
|
|
|
| 39 |
A |
151 |
147 |
3.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29334.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 28586.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.128 |
|
|
|
| 2 |
C |
-0.106 |
|
|
|
| 3 |
H |
0.132 |
|
|
|
| 4 |
C |
-0.032 |
|
|
|
| 5 |
H |
0.133 |
|
|
|
| 6 |
H |
0.132 |
|
|
|
| 7 |
C |
-0.330 |
|
|
|
| 8 |
H |
0.149 |
|
|
|
| 9 |
H |
0.149 |
|
|
|
| 10 |
C |
-0.273 |
|
|
|
| 11 |
H |
0.148 |
|
|
|
| 12 |
H |
0.155 |
|
|
|
| 13 |
C |
-0.348 |
|
|
|
| 14 |
H |
0.246 |
|
|
|
| 15 |
N |
-0.282 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.490 |
0.783 |
-0.662 |
1.136 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.542 |
-1.861 |
0.395 |
| y |
-1.861 |
-40.980 |
1.507 |
| z |
0.395 |
1.507 |
-37.312 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.604 |
-1.861 |
0.395 |
| y |
-1.861 |
-4.553 |
1.507 |
| z |
0.395 |
1.507 |
0.949 |
|
| Polar |
|---|
| 3z2-r2 | 1.897 |
|---|
| x2-y2 | 5.438 |
|---|
| xy | -1.861 |
|---|
| xz | 0.395 |
|---|
| yz | 1.507 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.389 |
0.049 |
-0.231 |
| y |
0.049 |
8.958 |
0.093 |
| z |
-0.231 |
0.093 |
7.237 |
<r2> (average value of r
2) Å
2
| <r2> |
125.678 |
| (<r2>)1/2 |
11.211 |