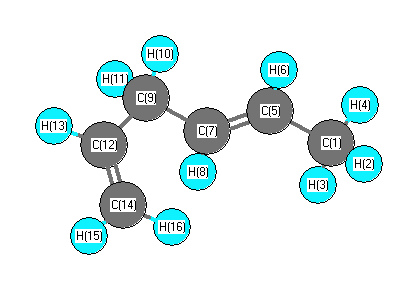
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3206 |
3124 |
39.41 |
|
|
|
| 2 |
A |
3112 |
3033 |
17.69 |
|
|
|
| 3 |
A |
3097 |
3018 |
81.65 |
|
|
|
| 4 |
A |
3092 |
3013 |
40.61 |
|
|
|
| 5 |
A |
3089 |
3010 |
7.36 |
|
|
|
| 6 |
A |
3068 |
2990 |
9.74 |
|
|
|
| 7 |
A |
3036 |
2959 |
40.54 |
|
|
|
| 8 |
A |
2996 |
2920 |
26.98 |
|
|
|
| 9 |
A |
2972 |
2896 |
52.85 |
|
|
|
| 10 |
A |
2956 |
2881 |
48.12 |
|
|
|
| 11 |
A |
1706 |
1663 |
1.16 |
|
|
|
| 12 |
A |
1672 |
1629 |
16.52 |
|
|
|
| 13 |
A |
1517 |
1478 |
11.72 |
|
|
|
| 14 |
A |
1506 |
1468 |
8.32 |
|
|
|
| 15 |
A |
1501 |
1463 |
14.39 |
|
|
|
| 16 |
A |
1452 |
1415 |
2.46 |
|
|
|
| 17 |
A |
1436 |
1399 |
4.76 |
|
|
|
| 18 |
A |
1369 |
1334 |
0.85 |
|
|
|
| 19 |
A |
1335 |
1301 |
0.91 |
|
|
|
| 20 |
A |
1327 |
1293 |
2.63 |
|
|
|
| 21 |
A |
1319 |
1286 |
0.68 |
|
|
|
| 22 |
A |
1227 |
1196 |
1.47 |
|
|
|
| 23 |
A |
1141 |
1112 |
5.27 |
|
|
|
| 24 |
A |
1114 |
1086 |
5.46 |
|
|
|
| 25 |
A |
1079 |
1052 |
0.45 |
|
|
|
| 26 |
A |
1049 |
1023 |
6.19 |
|
|
|
| 27 |
A |
1031 |
1005 |
11.94 |
|
|
|
| 28 |
A |
1008 |
983 |
42.25 |
|
|
|
| 29 |
A |
974 |
949 |
21.08 |
|
|
|
| 30 |
A |
954 |
930 |
40.81 |
|
|
|
| 31 |
A |
912 |
889 |
2.19 |
|
|
|
| 32 |
A |
894 |
871 |
2.51 |
|
|
|
| 33 |
A |
783 |
763 |
0.39 |
|
|
|
| 34 |
A |
602 |
587 |
6.98 |
|
|
|
| 35 |
A |
545 |
532 |
8.31 |
|
|
|
| 36 |
A |
460 |
448 |
0.33 |
|
|
|
| 37 |
A |
312 |
304 |
0.40 |
|
|
|
| 38 |
A |
275 |
268 |
2.22 |
|
|
|
| 39 |
A |
204 |
199 |
1.30 |
|
|
|
| 40 |
A |
155 |
151 |
0.58 |
|
|
|
| 41 |
A |
123 |
120 |
0.03 |
|
|
|
| 42 |
A |
74 |
72 |
0.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30840.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30054.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.595 |
|
|
|
| 2 |
H |
0.155 |
|
|
|
| 3 |
H |
0.160 |
|
|
|
| 4 |
H |
0.155 |
|
|
|
| 5 |
C |
-0.028 |
|
|
|
| 6 |
H |
0.133 |
|
|
|
| 7 |
C |
-0.002 |
|
|
|
| 8 |
H |
0.142 |
|
|
|
| 9 |
C |
-0.473 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
| 11 |
H |
0.140 |
|
|
|
| 12 |
C |
0.047 |
|
|
|
| 13 |
H |
0.122 |
|
|
|
| 14 |
C |
-0.356 |
|
|
|
| 15 |
H |
0.112 |
|
|
|
| 16 |
H |
0.132 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.335 |
-0.320 |
-0.049 |
0.466 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.852 |
1.544 |
-0.490 |
| y |
1.544 |
-38.215 |
0.724 |
| z |
-0.490 |
0.724 |
-39.778 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.145 |
1.544 |
-0.490 |
| y |
1.544 |
0.100 |
0.724 |
| z |
-0.490 |
0.724 |
-2.244 |
|
| Polar |
|---|
| 3z2-r2 | -4.489 |
|---|
| x2-y2 | 1.363 |
|---|
| xy | 1.544 |
|---|
| xz | -0.490 |
|---|
| yz | 0.724 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.017 |
0.220 |
-0.969 |
| y |
0.220 |
10.389 |
0.136 |
| z |
-0.969 |
0.136 |
7.894 |
<r2> (average value of r
2) Å
2
| <r2> |
194.587 |
| (<r2>)1/2 |
13.949 |