Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -14.902196 |
| Energy at 298.15K | -14.908100 |
| HF Energy | -14.902196 |
| Nuclear repulsion energy | 26.334605 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
2986 |
2910 |
0.00 |
282.61 |
0.01 |
0.02 |
| 2 |
A1g |
1438 |
1401 |
0.00 |
3.91 |
0.75 |
0.86 |
| 3 |
A1g |
999 |
973 |
0.00 |
16.77 |
0.26 |
0.42 |
| 4 |
A1u |
293 |
286 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
2979 |
2903 |
90.79 |
0.00 |
0.00 |
0.00 |
| 6 |
A2u |
1431 |
1394 |
4.69 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3049 |
2972 |
0.00 |
157.70 |
0.75 |
0.86 |
| 7 |
Eg |
3049 |
2972 |
0.00 |
157.71 |
0.75 |
0.86 |
| 8 |
Eg |
1525 |
1487 |
0.00 |
37.33 |
0.75 |
0.86 |
| 8 |
Eg |
1525 |
1487 |
0.00 |
37.32 |
0.75 |
0.86 |
| 9 |
Eg |
1226 |
1195 |
0.00 |
2.52 |
0.75 |
0.86 |
| 9 |
Eg |
1226 |
1195 |
0.00 |
2.51 |
0.75 |
0.86 |
| 10 |
Eu |
3079 |
3001 |
118.17 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3079 |
3001 |
118.24 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1531 |
1492 |
14.13 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1531 |
1492 |
14.13 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
832 |
811 |
5.77 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
832 |
811 |
5.77 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16305.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 15889.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.775 |
| C2 |
0.000 |
0.000 |
-0.775 |
| H3 |
0.000 |
1.024 |
1.172 |
| H4 |
-0.887 |
-0.512 |
1.172 |
| H5 |
0.887 |
-0.512 |
1.172 |
| H6 |
0.000 |
-1.024 |
-1.172 |
| H7 |
-0.887 |
0.512 |
-1.172 |
| H8 |
0.887 |
0.512 |
-1.172 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5497 | 1.0988 | 1.0988 | 1.0988 | 2.2003 | 2.2003 | 2.2003 |
C2 | 1.5497 | | 2.2003 | 2.2003 | 2.2003 | 1.0988 | 1.0988 | 1.0988 | H3 | 1.0988 | 2.2003 | | 1.7742 | 1.7742 | 3.1137 | 2.5588 | 2.5588 | H4 | 1.0988 | 2.2003 | 1.7742 | | 1.7742 | 2.5588 | 2.5588 | 3.1137 | H5 | 1.0988 | 2.2003 | 1.7742 | 1.7742 | | 2.5588 | 3.1137 | 2.5588 | H6 | 2.2003 | 1.0988 | 3.1137 | 2.5588 | 2.5588 | | 1.7742 | 1.7742 | H7 | 2.2003 | 1.0988 | 2.5588 | 2.5588 | 3.1137 | 1.7742 | | 1.7742 | H8 | 2.2003 | 1.0988 | 2.5588 | 3.1137 | 2.5588 | 1.7742 | 1.7742 | |
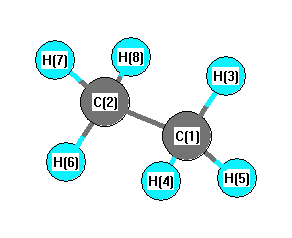 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.214 |
|
C1 |
C2 |
H7 |
111.214 |
| C1 |
C2 |
H8 |
111.214 |
|
C2 |
C1 |
H3 |
111.214 |
| C2 |
C1 |
H4 |
111.214 |
|
C2 |
C1 |
H5 |
111.214 |
| H3 |
C1 |
H4 |
107.673 |
|
H3 |
C1 |
H5 |
107.673 |
| H4 |
C1 |
H5 |
107.673 |
|
H6 |
C2 |
H7 |
107.673 |
| H6 |
C2 |
H8 |
107.673 |
|
H7 |
C2 |
H8 |
107.673 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.432 |
|
|
|
| 2 |
C |
-0.432 |
|
|
|
| 3 |
H |
0.144 |
|
|
|
| 4 |
H |
0.144 |
|
|
|
| 5 |
H |
0.144 |
|
|
|
| 6 |
H |
0.144 |
|
|
|
| 7 |
H |
0.144 |
|
|
|
| 8 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.029 |
0.000 |
0.000 |
| y |
0.000 |
-15.029 |
0.000 |
| z |
0.000 |
0.000 |
-15.623 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.297 |
0.000 |
0.000 |
| y |
0.000 |
0.297 |
0.000 |
| z |
0.000 |
0.000 |
-0.595 |
|
| Polar |
|---|
| 3z2-r2 | -1.189 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.704 |
0.000 |
0.000 |
| y |
0.000 |
3.704 |
0.000 |
| z |
0.000 |
0.000 |
3.942 |
<r2> (average value of r
2) Å
2
| <r2> |
28.857 |
| (<r2>)1/2 |
5.372 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -14.898256 |
| Energy at 298.15K | |
| HF Energy | -14.898256 |
| Nuclear repulsion energy | 26.258804 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3000 |
2924 |
0.00 |
|
|
|
| 2 |
A1' |
1461 |
1424 |
0.00 |
|
|
|
| 3 |
A1' |
995 |
969 |
0.00 |
|
|
|
| 4 |
A1" |
285i |
277i |
0.00 |
|
|
|
| 5 |
A2" |
2985 |
2909 |
103.08 |
|
|
|
| 6 |
A2" |
1429 |
1393 |
5.03 |
|
|
|
| 7 |
E' |
3084 |
3005 |
103.33 |
|
|
|
| 7 |
E' |
3084 |
3005 |
103.37 |
|
|
|
| 8 |
E' |
1538 |
1499 |
16.21 |
|
|
|
| 8 |
E' |
1538 |
1499 |
16.21 |
|
|
|
| 9 |
E' |
906 |
883 |
6.57 |
|
|
|
| 9 |
E' |
906 |
883 |
6.57 |
|
|
|
| 10 |
E" |
3057 |
2979 |
0.00 |
|
|
|
| 10 |
E" |
3057 |
2979 |
0.00 |
|
|
|
| 11 |
E" |
1526 |
1487 |
0.00 |
|
|
|
| 11 |
E" |
1526 |
1487 |
0.00 |
|
|
|
| 12 |
E" |
1182 |
1152 |
0.00 |
|
|
|
| 12 |
E" |
1182 |
1152 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16085.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 15675.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.781 |
| C2 |
0.000 |
0.000 |
-0.781 |
| H3 |
0.000 |
1.020 |
1.186 |
| H4 |
-0.884 |
-0.510 |
1.186 |
| H5 |
0.884 |
-0.510 |
1.186 |
| H6 |
0.000 |
1.020 |
-1.186 |
| H7 |
0.884 |
-0.510 |
-1.186 |
| H8 |
-0.884 |
-0.510 |
-1.186 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5622 | 1.0978 | 1.0978 | 1.0978 | 2.2160 | 2.2160 | 2.2160 |
C2 | 1.5622 | | 2.2160 | 2.2160 | 2.2160 | 1.0978 | 1.0978 | 1.0978 | H3 | 1.0978 | 2.2160 | | 1.7674 | 1.7674 | 2.3720 | 2.9580 | 2.9580 | H4 | 1.0978 | 2.2160 | 1.7674 | | 1.7674 | 2.9580 | 2.9580 | 2.3720 | H5 | 1.0978 | 2.2160 | 1.7674 | 1.7674 | | 2.9580 | 2.3720 | 2.9580 | H6 | 2.2160 | 1.0978 | 2.3720 | 2.9580 | 2.9580 | | 1.7674 | 1.7674 | H7 | 2.2160 | 1.0978 | 2.9580 | 2.9580 | 2.3720 | 1.7674 | | 1.7674 | H8 | 2.2160 | 1.0978 | 2.9580 | 2.3720 | 2.9580 | 1.7674 | 1.7674 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.643 |
|
C1 |
C2 |
H7 |
111.643 |
| C1 |
C2 |
H8 |
111.643 |
|
C2 |
C1 |
H3 |
111.643 |
| C2 |
C1 |
H4 |
111.643 |
|
C2 |
C1 |
H5 |
111.643 |
| H3 |
C1 |
H4 |
107.215 |
|
H3 |
C1 |
H5 |
107.215 |
| H4 |
C1 |
H5 |
107.215 |
|
H6 |
C2 |
H7 |
107.215 |
| H6 |
C2 |
H8 |
107.215 |
|
H7 |
C2 |
H8 |
107.215 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.446 |
|
|
|
| 2 |
C |
-0.446 |
|
|
|
| 3 |
H |
0.149 |
|
|
|
| 4 |
H |
0.149 |
|
|
|
| 5 |
H |
0.149 |
|
|
|
| 6 |
H |
0.149 |
|
|
|
| 7 |
H |
0.149 |
|
|
|
| 8 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.014 |
0.000 |
0.000 |
| y |
0.000 |
-15.014 |
0.000 |
| z |
0.000 |
0.000 |
-15.584 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.285 |
0.000 |
0.000 |
| y |
0.000 |
0.285 |
0.000 |
| z |
0.000 |
0.000 |
-0.570 |
|
| Polar |
|---|
| 3z2-r2 | -1.139 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.639 |
0.000 |
0.000 |
| y |
0.000 |
3.639 |
0.000 |
| z |
0.000 |
0.000 |
3.769 |
<r2> (average value of r
2) Å
2
| <r2> |
29.064 |
| (<r2>)1/2 |
5.391 |
