Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -25.382978 |
| Energy at 298.15K | -25.390974 |
| Nuclear repulsion energy | 49.395942 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3483 |
3394 |
3.92 |
|
|
|
| 2 |
A' |
3055 |
2977 |
84.03 |
|
|
|
| 3 |
A' |
2998 |
2921 |
44.44 |
|
|
|
| 4 |
A' |
2967 |
2892 |
43.87 |
|
|
|
| 5 |
A' |
1698 |
1654 |
33.50 |
|
|
|
| 6 |
A' |
1523 |
1484 |
5.05 |
|
|
|
| 7 |
A' |
1505 |
1467 |
0.25 |
|
|
|
| 8 |
A' |
1421 |
1385 |
7.48 |
|
|
|
| 9 |
A' |
1386 |
1350 |
16.84 |
|
|
|
| 10 |
A' |
1139 |
1110 |
19.39 |
|
|
|
| 11 |
A' |
1059 |
1032 |
28.05 |
|
|
|
| 12 |
A' |
879 |
856 |
3.15 |
|
|
|
| 13 |
A' |
638 |
621 |
312.33 |
|
|
|
| 14 |
A' |
390 |
380 |
17.34 |
|
|
|
| 15 |
A" |
3609 |
3517 |
0.84 |
|
|
|
| 16 |
A" |
3069 |
2991 |
107.83 |
|
|
|
| 17 |
A" |
3036 |
2958 |
1.61 |
|
|
|
| 18 |
A" |
1513 |
1474 |
8.27 |
|
|
|
| 19 |
A" |
1373 |
1338 |
0.20 |
|
|
|
| 20 |
A" |
1263 |
1231 |
0.06 |
|
|
|
| 21 |
A" |
993 |
967 |
0.09 |
|
|
|
| 22 |
A" |
778 |
759 |
1.81 |
|
|
|
| 23 |
A" |
261 |
254 |
27.78 |
|
|
|
| 24 |
A" |
235 |
229 |
27.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20134.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 19621.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.320 |
-0.082 |
0.000 |
| C2 |
0.000 |
0.591 |
0.000 |
| C3 |
1.234 |
-0.356 |
0.000 |
| H4 |
2.174 |
0.215 |
0.000 |
| H5 |
1.228 |
-1.004 |
0.889 |
| H6 |
1.228 |
-1.004 |
-0.889 |
| H7 |
0.042 |
1.248 |
-0.880 |
| H8 |
0.042 |
1.248 |
0.880 |
| H9 |
-1.524 |
-0.615 |
0.843 |
| H10 |
-1.524 |
-0.615 |
-0.843 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4819 | 2.5688 | 3.5073 | 2.8524 | 2.8524 | 2.0975 | 2.0975 | 1.0173 | 1.0173 |
C2 | 1.4819 | | 1.5553 | 2.2065 | 2.2003 | 2.2003 | 1.0992 | 1.0992 | 2.1178 | 2.1178 | C3 | 2.5688 | 1.5553 | | 1.1006 | 1.0998 | 1.0998 | 2.1841 | 2.1841 | 2.8950 | 2.8950 | H4 | 3.5073 | 2.2065 | 1.1006 | | 1.7806 | 1.7806 | 2.5278 | 2.5278 | 3.8826 | 3.8826 | H5 | 2.8524 | 2.2003 | 1.0998 | 1.7806 | | 1.7785 | 3.0998 | 2.5456 | 2.7800 | 3.2749 | H6 | 2.8524 | 2.2003 | 1.0998 | 1.7806 | 1.7785 | | 2.5456 | 3.0998 | 3.2749 | 2.7800 | H7 | 2.0975 | 1.0992 | 2.1841 | 2.5278 | 3.0998 | 2.5456 | | 1.7593 | 2.9813 | 2.4339 | H8 | 2.0975 | 1.0992 | 2.1841 | 2.5278 | 2.5456 | 3.0998 | 1.7593 | | 2.4339 | 2.9813 | H9 | 1.0173 | 2.1178 | 2.8950 | 3.8826 | 2.7800 | 3.2749 | 2.9813 | 2.4339 | | 1.6850 | H10 | 1.0173 | 2.1178 | 2.8950 | 3.8826 | 3.2749 | 2.7800 | 2.4339 | 2.9813 | 1.6850 | |
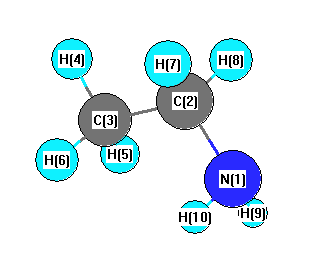 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.489 |
|
N1 |
C2 |
H7 |
107.787 |
| N1 |
C2 |
H8 |
107.787 |
|
C2 |
N1 |
H9 |
114.577 |
| C2 |
N1 |
H10 |
114.577 |
|
C2 |
C3 |
H4 |
111.210 |
| C2 |
C3 |
H5 |
110.761 |
|
C2 |
C3 |
H6 |
110.761 |
| C3 |
C2 |
H7 |
109.531 |
|
C3 |
C2 |
H8 |
109.531 |
| H4 |
C3 |
H5 |
108.038 |
|
H4 |
C3 |
H6 |
108.038 |
| H5 |
C3 |
H6 |
107.908 |
|
H7 |
C2 |
H8 |
106.310 |
| H9 |
N1 |
H10 |
111.822 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.558 |
|
|
|
| 2 |
C |
-0.202 |
|
|
|
| 3 |
C |
-0.437 |
|
|
|
| 4 |
H |
0.135 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.129 |
|
|
|
| 7 |
H |
0.143 |
|
|
|
| 8 |
H |
0.143 |
|
|
|
| 9 |
H |
0.258 |
|
|
|
| 10 |
H |
0.258 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.749 |
-1.005 |
0.000 |
1.253 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.392 |
-0.064 |
-0.093 |
| y |
-0.064 |
4.860 |
-0.085 |
| z |
-0.093 |
-0.085 |
4.445 |
<r2> (average value of r
2) Å
2
| <r2> |
52.887 |
| (<r2>)1/2 |
7.272 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -25.382603 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
