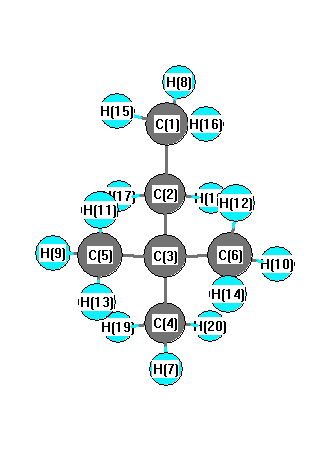
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3072 |
2993 |
90.91 |
|
|
|
| 2 |
A' |
3063 |
2985 |
180.78 |
|
|
|
| 3 |
A' |
3060 |
2982 |
29.86 |
|
|
|
| 4 |
A' |
3050 |
2972 |
0.57 |
|
|
|
| 5 |
A' |
2989 |
2913 |
21.13 |
|
|
|
| 6 |
A' |
2979 |
2903 |
27.48 |
|
|
|
| 7 |
A' |
2966 |
2891 |
65.07 |
|
|
|
| 8 |
A' |
2960 |
2884 |
35.98 |
|
|
|
| 9 |
A' |
1540 |
1501 |
14.55 |
|
|
|
| 10 |
A' |
1533 |
1494 |
9.73 |
|
|
|
| 11 |
A' |
1530 |
1491 |
7.69 |
|
|
|
| 12 |
A' |
1513 |
1475 |
0.05 |
|
|
|
| 13 |
A' |
1504 |
1466 |
0.15 |
|
|
|
| 14 |
A' |
1444 |
1407 |
4.08 |
|
|
|
| 15 |
A' |
1431 |
1394 |
13.26 |
|
|
|
| 16 |
A' |
1416 |
1379 |
8.53 |
|
|
|
| 17 |
A' |
1376 |
1341 |
0.14 |
|
|
|
| 18 |
A' |
1279 |
1246 |
4.51 |
|
|
|
| 19 |
A' |
1241 |
1209 |
9.10 |
|
|
|
| 20 |
A' |
1097 |
1069 |
2.75 |
|
|
|
| 21 |
A' |
1029 |
1003 |
3.95 |
|
|
|
| 22 |
A' |
1018 |
992 |
2.61 |
|
|
|
| 23 |
A' |
933 |
909 |
0.78 |
|
|
|
| 24 |
A' |
869 |
847 |
0.19 |
|
|
|
| 25 |
A' |
694 |
676 |
0.32 |
|
|
|
| 26 |
A' |
481 |
468 |
0.39 |
|
|
|
| 27 |
A' |
403 |
393 |
0.03 |
|
|
|
| 28 |
A' |
349 |
340 |
0.02 |
|
|
|
| 29 |
A' |
269 |
262 |
0.04 |
|
|
|
| 30 |
A' |
252 |
245 |
0.04 |
|
|
|
| 31 |
A" |
3078 |
2999 |
64.33 |
|
|
|
| 32 |
A" |
3063 |
2985 |
68.85 |
|
|
|
| 33 |
A" |
3054 |
2976 |
31.34 |
|
|
|
| 34 |
A" |
3050 |
2972 |
0.18 |
|
|
|
| 35 |
A" |
3004 |
2927 |
21.82 |
|
|
|
| 36 |
A" |
2970 |
2895 |
57.72 |
|
|
|
| 37 |
A" |
1536 |
1497 |
0.10 |
|
|
|
| 38 |
A" |
1525 |
1486 |
13.83 |
|
|
|
| 39 |
A" |
1511 |
1472 |
0.12 |
|
|
|
| 40 |
A" |
1505 |
1466 |
0.13 |
|
|
|
| 41 |
A" |
1416 |
1380 |
12.86 |
|
|
|
| 42 |
A" |
1336 |
1302 |
1.60 |
|
|
|
| 43 |
A" |
1249 |
1218 |
2.64 |
|
|
|
| 44 |
A" |
1107 |
1079 |
0.38 |
|
|
|
| 45 |
A" |
1006 |
980 |
0.91 |
|
|
|
| 46 |
A" |
967 |
943 |
0.01 |
|
|
|
| 47 |
A" |
936 |
912 |
0.03 |
|
|
|
| 48 |
A" |
792 |
772 |
2.86 |
|
|
|
| 49 |
A" |
415 |
404 |
0.02 |
|
|
|
| 50 |
A" |
329 |
321 |
0.01 |
|
|
|
| 51 |
A" |
278 |
271 |
0.00 |
|
|
|
| 52 |
A" |
241 |
235 |
0.00 |
|
|
|
| 53 |
A" |
207 |
201 |
0.00 |
|
|
|
| 54 |
A" |
92 |
90 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41002.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 39956.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.511 |
|
|
|
| 2 |
C |
-0.228 |
|
|
|
| 3 |
C |
0.371 |
|
|
|
| 4 |
C |
-0.512 |
|
|
|
| 5 |
C |
-0.535 |
|
|
|
| 6 |
C |
-0.535 |
|
|
|
| 7 |
H |
0.140 |
|
|
|
| 8 |
H |
0.143 |
|
|
|
| 9 |
H |
0.140 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
H |
0.146 |
|
|
|
| 12 |
H |
0.146 |
|
|
|
| 13 |
H |
0.138 |
|
|
|
| 14 |
H |
0.138 |
|
|
|
| 15 |
H |
0.148 |
|
|
|
| 16 |
H |
0.148 |
|
|
|
| 17 |
H |
0.123 |
|
|
|
| 18 |
H |
0.123 |
|
|
|
| 19 |
H |
0.140 |
|
|
|
| 20 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.018 |
-0.055 |
0.000 |
0.058 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.599 |
-0.290 |
0.000 |
| y |
-0.290 |
-42.661 |
0.000 |
| z |
0.000 |
0.000 |
-42.467 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.035 |
-0.290 |
0.000 |
| y |
-0.290 |
-0.128 |
0.000 |
| z |
0.000 |
0.000 |
0.162 |
|
| Polar |
|---|
| 3z2-r2 | 0.325 |
|---|
| x2-y2 | 0.062 |
|---|
| xy | -0.290 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.365 |
0.032 |
0.002 |
| y |
0.032 |
10.401 |
-0.000 |
| z |
0.002 |
-0.000 |
10.395 |
<r2> (average value of r
2) Å
2
| <r2> |
170.135 |
| (<r2>)1/2 |
13.044 |