Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -52.775310 |
| Energy at 298.15K | -52.780072 |
| HF Energy | -52.775310 |
| Nuclear repulsion energy | 66.122225 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3089 |
3011 |
17.12 |
|
|
|
| 2 |
A' |
3059 |
2981 |
20.22 |
|
|
|
| 3 |
A' |
1522 |
1483 |
3.73 |
|
|
|
| 4 |
A' |
1501 |
1463 |
4.08 |
|
|
|
| 5 |
A' |
1407 |
1371 |
1.55 |
|
|
|
| 6 |
A' |
1290 |
1257 |
16.34 |
|
|
|
| 7 |
A' |
1078 |
1051 |
9.86 |
|
|
|
| 8 |
A' |
953 |
929 |
92.98 |
|
|
|
| 9 |
A' |
701 |
683 |
53.18 |
|
|
|
| 10 |
A' |
352 |
343 |
3.00 |
|
|
|
| 11 |
A' |
227 |
221 |
18.87 |
|
|
|
| 12 |
A" |
3182 |
3101 |
18.49 |
|
|
|
| 13 |
A" |
3137 |
3057 |
14.69 |
|
|
|
| 14 |
A" |
1289 |
1256 |
0.00 |
|
|
|
| 15 |
A" |
1184 |
1154 |
0.84 |
|
|
|
| 16 |
A" |
1047 |
1020 |
2.20 |
|
|
|
| 17 |
A" |
807 |
787 |
1.19 |
|
|
|
| 18 |
A" |
116 |
113 |
15.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12970.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12640.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.636 |
0.000 |
| C2 |
1.031 |
-0.497 |
0.000 |
| Cl3 |
-1.728 |
-0.138 |
0.000 |
| F4 |
2.349 |
0.138 |
0.000 |
| H5 |
0.062 |
1.249 |
0.899 |
| H6 |
0.062 |
1.249 |
-0.899 |
| H7 |
0.969 |
-1.113 |
0.901 |
| H8 |
0.969 |
-1.113 |
-0.901 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5314 | 1.8937 | 2.4008 | 1.0904 | 1.0904 | 2.1932 | 2.1932 |
C2 | 1.5314 | | 2.7824 | 1.4624 | 2.1902 | 2.1902 | 1.0938 | 1.0938 | Cl3 | 1.8937 | 2.7824 | | 4.0862 | 2.4367 | 2.4367 | 3.0063 | 3.0063 | F4 | 2.4008 | 1.4624 | 4.0862 | | 2.6973 | 2.6973 | 2.0690 | 2.0690 | H5 | 1.0904 | 2.1902 | 2.4367 | 2.6973 | | 1.7986 | 2.5309 | 3.1060 | H6 | 1.0904 | 2.1902 | 2.4367 | 2.6973 | 1.7986 | | 3.1060 | 2.5309 | H7 | 2.1932 | 1.0938 | 3.0063 | 2.0690 | 2.5309 | 3.1060 | | 1.8025 | H8 | 2.1932 | 1.0938 | 3.0063 | 2.0690 | 3.1060 | 2.5309 | 1.8025 | |
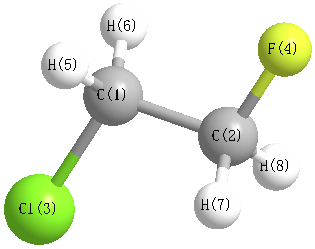 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
106.608 |
|
C1 |
C2 |
H7 |
112.253 |
| C1 |
C2 |
H8 |
112.253 |
|
C2 |
C1 |
Cl3 |
108.187 |
| C2 |
C1 |
H5 |
112.219 |
|
C2 |
C1 |
H6 |
112.219 |
| Cl3 |
C1 |
H5 |
106.347 |
|
Cl3 |
C1 |
H6 |
106.347 |
| F4 |
C2 |
H7 |
107.197 |
|
F4 |
C2 |
H8 |
107.197 |
| H5 |
C1 |
H6 |
111.124 |
|
H7 |
C2 |
H8 |
110.971 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.465 |
|
|
|
| 2 |
C |
0.008 |
|
|
|
| 3 |
Cl |
-0.031 |
|
|
|
| 4 |
F |
-0.274 |
|
|
|
| 5 |
H |
0.218 |
|
|
|
| 6 |
H |
0.218 |
|
|
|
| 7 |
H |
0.164 |
|
|
|
| 8 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.068 |
-0.017 |
0.000 |
0.071 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.244 |
-1.747 |
0.000 |
| y |
-1.747 |
-28.262 |
0.000 |
| z |
0.000 |
0.000 |
-28.957 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.635 |
-1.747 |
0.000 |
| y |
-1.747 |
6.338 |
0.000 |
| z |
0.000 |
0.000 |
5.296 |
|
| Polar |
|---|
| 3z2-r2 | 10.593 |
|---|
| x2-y2 | -11.982 |
|---|
| xy | -1.747 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.803 |
0.734 |
0.000 |
| y |
0.734 |
4.336 |
0.000 |
| z |
0.000 |
0.000 |
3.761 |
<r2> (average value of r
2) Å
2
| <r2> |
97.659 |
| (<r2>)1/2 |
9.882 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -52.773565 |
| Energy at 298.15K | -52.778442 |
| HF Energy | -52.773565 |
| Nuclear repulsion energy | 67.166658 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3158 |
3077 |
11.09 |
|
|
|
| 2 |
A |
3112 |
3032 |
20.67 |
|
|
|
| 3 |
A |
3070 |
2992 |
18.59 |
|
|
|
| 4 |
A |
3022 |
2945 |
26.17 |
|
|
|
| 5 |
A |
1499 |
1461 |
4.68 |
|
|
|
| 6 |
A |
1471 |
1434 |
10.97 |
|
|
|
| 7 |
A |
1413 |
1377 |
10.12 |
|
|
|
| 8 |
A |
1333 |
1299 |
34.70 |
|
|
|
| 9 |
A |
1243 |
1211 |
0.66 |
|
|
|
| 10 |
A |
1191 |
1160 |
0.70 |
|
|
|
| 11 |
A |
1084 |
1057 |
16.99 |
|
|
|
| 12 |
A |
1018 |
992 |
40.33 |
|
|
|
| 13 |
A |
964 |
939 |
19.02 |
|
|
|
| 14 |
A |
831 |
810 |
19.17 |
|
|
|
| 15 |
A |
617 |
602 |
33.54 |
|
|
|
| 16 |
A |
438 |
427 |
18.28 |
|
|
|
| 17 |
A |
275 |
268 |
1.75 |
|
|
|
| 18 |
A |
118 |
115 |
3.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12928.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12598.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.050 |
0.860 |
-0.324 |
| C2 |
1.221 |
0.432 |
0.398 |
| Cl3 |
-1.502 |
-0.286 |
0.066 |
| F4 |
1.752 |
-0.775 |
-0.211 |
| H5 |
-0.366 |
1.855 |
0.002 |
| H6 |
0.074 |
0.828 |
-1.408 |
| H7 |
1.992 |
1.206 |
0.284 |
| H8 |
1.048 |
0.213 |
1.457 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5234 | 1.8903 | 2.4360 | 1.0935 | 1.0913 | 2.1591 | 2.1900 |
C2 | 1.5234 | | 2.8352 | 1.4525 | 2.1674 | 2.1761 | 1.0990 | 1.0944 | Cl3 | 1.8903 | 2.8352 | | 3.3018 | 2.4248 | 2.4284 | 3.8057 | 2.9473 | F4 | 2.4360 | 1.4525 | 3.3018 | | 3.3830 | 2.6115 | 2.0561 | 2.0622 | H5 | 1.0935 | 2.1674 | 2.4248 | 3.3830 | | 1.7988 | 2.4616 | 2.6092 | H6 | 1.0913 | 2.1761 | 2.4284 | 2.6115 | 1.7988 | | 2.5858 | 3.0877 | H7 | 2.1591 | 1.0990 | 3.8057 | 2.0561 | 2.4616 | 2.5858 | | 1.8033 | H8 | 2.1900 | 1.0944 | 2.9473 | 2.0622 | 2.6092 | 3.0877 | 1.8033 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.857 |
|
C1 |
C2 |
H7 |
109.782 |
| C1 |
C2 |
H8 |
112.535 |
|
C2 |
C1 |
Cl3 |
111.863 |
| C2 |
C1 |
H5 |
110.767 |
|
C2 |
C1 |
H6 |
111.596 |
| Cl3 |
C1 |
H5 |
105.587 |
|
Cl3 |
C1 |
H6 |
105.939 |
| F4 |
C2 |
H7 |
106.558 |
|
F4 |
C2 |
H8 |
107.299 |
| H5 |
C1 |
H6 |
110.835 |
|
H7 |
C2 |
H8 |
110.606 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.461 |
|
|
|
| 2 |
C |
0.002 |
|
|
|
| 3 |
Cl |
-0.022 |
|
|
|
| 4 |
F |
-0.265 |
|
|
|
| 5 |
H |
0.198 |
|
|
|
| 6 |
H |
0.221 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
H |
0.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.932 |
3.415 |
0.470 |
3.571 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.278 |
3.126 |
1.548 |
| y |
3.126 |
-29.198 |
-0.634 |
| z |
1.548 |
-0.634 |
-28.751 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.303 |
3.126 |
1.548 |
| y |
3.126 |
2.316 |
-0.634 |
| z |
1.548 |
-0.634 |
2.987 |
|
| Polar |
|---|
| 3z2-r2 | 5.974 |
|---|
| x2-y2 | -5.079 |
|---|
| xy | 3.126 |
|---|
| xz | 1.548 |
|---|
| yz | -0.634 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.048 |
1.088 |
-0.210 |
| y |
1.088 |
4.873 |
-0.181 |
| z |
-0.210 |
-0.181 |
3.973 |
<r2> (average value of r
2) Å
2
| <r2> |
87.338 |
| (<r2>)1/2 |
9.345 |
