Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/CEP-121G
Geometric Data calculated at B3LYP/CEP-121G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -41.060332 |
| Energy at 298.15K | -41.072554 |
| HF Energy | -41.060332 |
| Nuclear repulsion energy | 128.171230 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3114 |
3034 |
100.34 |
|
|
|
| 2 |
A |
3090 |
3011 |
17.41 |
|
|
|
| 3 |
A |
3077 |
2998 |
32.60 |
|
|
|
| 4 |
A |
3064 |
2986 |
70.01 |
|
|
|
| 5 |
A |
3063 |
2985 |
94.48 |
|
|
|
| 6 |
A |
3040 |
2962 |
10.11 |
|
|
|
| 7 |
A |
3035 |
2958 |
40.20 |
|
|
|
| 8 |
A |
3016 |
2939 |
7.90 |
|
|
|
| 9 |
A |
2980 |
2904 |
19.43 |
|
|
|
| 10 |
A |
2976 |
2900 |
50.62 |
|
|
|
| 11 |
A |
2975 |
2899 |
82.08 |
|
|
|
| 12 |
A |
2962 |
2887 |
27.64 |
|
|
|
| 13 |
A |
1695 |
1652 |
3.39 |
|
|
|
| 14 |
A |
1533 |
1493 |
10.71 |
|
|
|
| 15 |
A |
1521 |
1483 |
8.50 |
|
|
|
| 16 |
A |
1520 |
1481 |
5.90 |
|
|
|
| 17 |
A |
1516 |
1477 |
0.77 |
|
|
|
| 18 |
A |
1511 |
1472 |
8.05 |
|
|
|
| 19 |
A |
1507 |
1469 |
1.73 |
|
|
|
| 20 |
A |
1440 |
1403 |
9.92 |
|
|
|
| 21 |
A |
1429 |
1393 |
2.30 |
|
|
|
| 22 |
A |
1424 |
1388 |
6.60 |
|
|
|
| 23 |
A |
1377 |
1341 |
0.63 |
|
|
|
| 24 |
A |
1321 |
1287 |
0.07 |
|
|
|
| 25 |
A |
1311 |
1278 |
2.21 |
|
|
|
| 26 |
A |
1296 |
1263 |
1.13 |
|
|
|
| 27 |
A |
1258 |
1226 |
0.24 |
|
|
|
| 28 |
A |
1175 |
1145 |
1.17 |
|
|
|
| 29 |
A |
1115 |
1086 |
1.25 |
|
|
|
| 30 |
A |
1075 |
1047 |
2.01 |
|
|
|
| 31 |
A |
1067 |
1040 |
1.56 |
|
|
|
| 32 |
A |
1054 |
1027 |
2.54 |
|
|
|
| 33 |
A |
1012 |
987 |
0.53 |
|
|
|
| 34 |
A |
999 |
973 |
6.16 |
|
|
|
| 35 |
A |
912 |
889 |
3.34 |
|
|
|
| 36 |
A |
895 |
872 |
8.35 |
|
|
|
| 37 |
A |
873 |
851 |
5.96 |
|
|
|
| 38 |
A |
756 |
737 |
2.45 |
|
|
|
| 39 |
A |
718 |
700 |
42.33 |
|
|
|
| 40 |
A |
569 |
555 |
7.68 |
|
|
|
| 41 |
A |
463 |
451 |
0.96 |
|
|
|
| 42 |
A |
360 |
351 |
0.01 |
|
|
|
| 43 |
A |
281 |
274 |
0.03 |
|
|
|
| 44 |
A |
232 |
226 |
0.01 |
|
|
|
| 45 |
A |
196 |
191 |
0.03 |
|
|
|
| 46 |
A |
127 |
124 |
0.43 |
|
|
|
| 47 |
A |
87 |
85 |
0.02 |
|
|
|
| 48 |
A |
48 |
47 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36032.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 35113.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.524 |
0.956 |
-0.081 |
| C2 |
2.029 |
-0.482 |
-0.139 |
| C3 |
0.781 |
-0.943 |
0.141 |
| C4 |
-0.441 |
-0.136 |
0.558 |
| C5 |
-1.618 |
-0.260 |
-0.458 |
| C6 |
-2.884 |
0.514 |
-0.007 |
| H7 |
1.738 |
1.664 |
0.201 |
| H8 |
3.344 |
1.056 |
0.646 |
| H9 |
2.926 |
1.267 |
-1.058 |
| H10 |
2.783 |
-1.217 |
-0.435 |
| H11 |
0.610 |
-2.020 |
0.045 |
| H12 |
-0.188 |
0.925 |
0.688 |
| H13 |
-0.793 |
-0.494 |
1.541 |
| H14 |
-1.284 |
0.110 |
-1.439 |
| H15 |
-1.866 |
-1.324 |
-0.596 |
| H16 |
-3.696 |
0.410 |
-0.739 |
| H17 |
-3.252 |
0.142 |
0.960 |
| H18 |
-2.671 |
1.587 |
0.107 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5219 | 2.5868 | 3.2236 | 4.3336 | 5.4262 | 1.0953 | 1.1008 | 1.1006 | 2.2168 | 3.5403 | 2.8194 | 3.9670 | 4.1301 | 4.9739 | 6.2790 | 5.9251 | 5.2369 |
C2 | 1.5219 | | 1.3592 | 2.5899 | 3.6684 | 5.0147 | 2.1926 | 2.1703 | 2.1703 | 1.0935 | 2.1005 | 2.7535 | 3.2840 | 3.6080 | 4.0117 | 5.8258 | 5.4301 | 5.1418 | C3 | 2.5868 | 1.3592 | | 1.5232 | 2.5659 | 3.9468 | 2.7776 | 3.2893 | 3.3046 | 2.1005 | 1.0944 | 2.1750 | 2.1541 | 2.8050 | 2.7744 | 4.7595 | 4.2561 | 4.2805 | C4 | 3.2236 | 2.5899 | 1.5232 | | 1.5604 | 2.5900 | 2.8484 | 3.9693 | 3.9893 | 3.5424 | 2.2173 | 1.0985 | 1.1038 | 2.1817 | 2.1854 | 3.5464 | 2.8529 | 2.8540 | C5 | 4.3336 | 3.6684 | 2.5659 | 1.5604 | | 1.5504 | 3.9241 | 5.2515 | 4.8314 | 4.5042 | 2.8837 | 2.1827 | 2.1756 | 1.1005 | 1.1017 | 2.2012 | 2.2003 | 2.2000 | C6 | 5.4262 | 5.0147 | 3.9468 | 2.5900 | 1.5504 | | 4.7667 | 6.2854 | 5.9515 | 5.9405 | 4.3160 | 2.8137 | 2.7902 | 2.1852 | 2.1823 | 1.0987 | 1.0998 | 1.0997 | H7 | 1.0953 | 2.1926 | 2.7776 | 2.8484 | 3.9241 | 4.7667 | | 1.7747 | 1.7755 | 3.1303 | 3.8557 | 2.1194 | 3.5859 | 3.7727 | 4.7491 | 5.6556 | 5.2713 | 4.4105 | H8 | 1.1008 | 2.1703 | 3.2893 | 3.9693 | 5.2515 | 6.2854 | 1.7747 | | 1.7668 | 2.5788 | 4.1589 | 3.5352 | 4.5078 | 5.1634 | 5.8616 | 7.2047 | 6.6664 | 6.0629 | H9 | 1.1006 | 2.1703 | 3.3046 | 3.9893 | 4.8314 | 5.9515 | 1.7755 | 1.7668 | | 2.5655 | 4.1696 | 3.5862 | 4.8667 | 4.3824 | 5.4677 | 6.6853 | 6.5954 | 5.7259 | H10 | 2.2168 | 1.0935 | 2.1005 | 3.5424 | 4.5042 | 5.9405 | 3.1303 | 2.5788 | 2.5655 | | 2.3657 | 3.8313 | 4.1487 | 4.3937 | 4.6531 | 6.6873 | 6.3410 | 6.1567 | H11 | 3.5403 | 2.1005 | 1.0944 | 2.2173 | 2.8837 | 4.3160 | 3.8557 | 4.1589 | 4.1696 | 2.3657 | | 3.1182 | 2.5554 | 3.2130 | 2.6506 | 5.0060 | 4.5189 | 4.8763 | H12 | 2.8194 | 2.7535 | 2.1750 | 1.0985 | 2.1827 | 2.8137 | 2.1194 | 3.5352 | 3.5862 | 3.8313 | 3.1182 | | 1.7633 | 2.5278 | 3.0864 | 3.8222 | 3.1738 | 2.6344 | H13 | 3.9670 | 3.2840 | 2.1541 | 1.1038 | 2.1756 | 2.7902 | 3.5859 | 4.5078 | 4.8667 | 4.1487 | 2.5554 | 1.7633 | | 3.0802 | 2.5311 | 3.8006 | 2.6054 | 3.1491 | H14 | 4.1301 | 3.6080 | 2.8050 | 2.1817 | 1.1005 | 2.1852 | 3.7727 | 5.1634 | 4.3824 | 4.3937 | 3.2130 | 2.5278 | 3.0802 | | 1.7630 | 2.5304 | 3.1035 | 2.5495 | H15 | 4.9739 | 4.0117 | 2.7744 | 2.1854 | 1.1017 | 2.1823 | 4.7491 | 5.8616 | 5.4677 | 4.6531 | 2.6506 | 3.0864 | 2.5311 | 1.7630 | | 2.5252 | 2.5477 | 3.1016 | H16 | 6.2790 | 5.8258 | 4.7595 | 3.5464 | 2.2012 | 1.0987 | 5.6556 | 7.2047 | 6.6853 | 6.6873 | 5.0060 | 3.8222 | 3.8006 | 2.5304 | 2.5252 | | 1.7764 | 1.7759 | H17 | 5.9251 | 5.4301 | 4.2561 | 2.8529 | 2.2003 | 1.0998 | 5.2713 | 6.6664 | 6.5954 | 6.3410 | 4.5189 | 3.1738 | 2.6054 | 3.1035 | 2.5477 | 1.7764 | | 1.7756 | H18 | 5.2369 | 5.1418 | 4.2805 | 2.8540 | 2.2000 | 1.0997 | 4.4105 | 6.0629 | 5.7259 | 6.1567 | 4.8763 | 2.6344 | 3.1491 | 2.5495 | 3.1016 | 1.7759 | 1.7756 | |
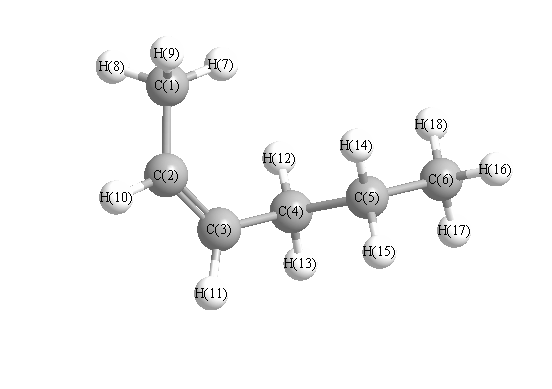 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.664 |
|
C1 |
C2 |
H10 |
114.912 |
| C2 |
C1 |
H7 |
112.787 |
|
C2 |
C1 |
H8 |
110.663 |
| C2 |
C1 |
H9 |
110.672 |
|
C2 |
C3 |
C4 |
127.838 |
| C2 |
C3 |
H11 |
117.352 |
|
C3 |
C2 |
H10 |
117.422 |
| C3 |
C4 |
C5 |
112.630 |
|
C3 |
C4 |
H12 |
111.083 |
| C3 |
C4 |
H13 |
109.124 |
|
C4 |
C3 |
H11 |
114.804 |
| C4 |
C5 |
C6 |
112.735 |
|
C4 |
C5 |
H14 |
108.924 |
| C4 |
C5 |
H15 |
109.146 |
|
C5 |
C4 |
H12 |
109.114 |
| C5 |
C4 |
H13 |
108.273 |
|
C5 |
C6 |
H16 |
111.244 |
| C5 |
C6 |
H17 |
111.106 |
|
C5 |
C6 |
H18 |
111.090 |
| C6 |
C5 |
H14 |
109.878 |
|
C6 |
C5 |
H15 |
109.585 |
| H7 |
C1 |
H8 |
107.824 |
|
H7 |
C1 |
H9 |
107.900 |
| H8 |
C1 |
H9 |
106.752 |
|
H12 |
C4 |
H13 |
106.386 |
| H14 |
C5 |
H16 |
94.103 |
|
H16 |
C6 |
H17 |
107.803 |
| H16 |
C6 |
H18 |
107.772 |
|
H17 |
C6 |
H18 |
107.660 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.564 |
|
|
|
| 2 |
C |
-0.042 |
|
|
|
| 3 |
C |
0.008 |
|
|
|
| 4 |
C |
-0.344 |
|
|
|
| 5 |
C |
-0.206 |
|
|
|
| 6 |
C |
-0.473 |
|
|
|
| 7 |
H |
0.150 |
|
|
|
| 8 |
H |
0.155 |
|
|
|
| 9 |
H |
0.154 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
H |
0.120 |
|
|
|
| 13 |
H |
0.126 |
|
|
|
| 14 |
H |
0.131 |
|
|
|
| 15 |
H |
0.127 |
|
|
|
| 16 |
H |
0.147 |
|
|
|
| 17 |
H |
0.139 |
|
|
|
| 18 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.054 |
0.221 |
-0.020 |
0.228 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.288 |
0.894 |
-0.641 |
| y |
0.894 |
-39.589 |
0.003 |
| z |
-0.641 |
0.003 |
-41.742 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.377 |
0.894 |
-0.641 |
| y |
0.894 |
0.926 |
0.003 |
| z |
-0.641 |
0.003 |
-2.303 |
|
| Polar |
|---|
| 3z2-r2 | -4.607 |
|---|
| x2-y2 | 0.301 |
|---|
| xy | 0.894 |
|---|
| xz | -0.641 |
|---|
| yz | 0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.917 |
0.743 |
-0.654 |
| y |
0.743 |
9.843 |
-0.014 |
| z |
-0.654 |
-0.014 |
8.435 |
<r2> (average value of r
2) Å
2
| <r2> |
219.030 |
| (<r2>)1/2 |
14.800 |
