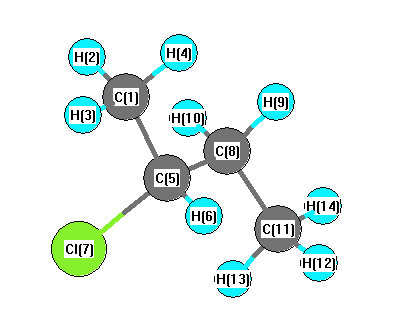
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3111 |
3032 |
39.85 |
|
|
|
| 2 |
A |
3097 |
3018 |
40.32 |
|
|
|
| 3 |
A |
3083 |
3004 |
52.04 |
|
|
|
| 4 |
A |
3073 |
2994 |
42.78 |
|
|
|
| 5 |
A |
3066 |
2988 |
4.62 |
|
|
|
| 6 |
A |
3034 |
2957 |
5.51 |
|
|
|
| 7 |
A |
2992 |
2915 |
21.76 |
|
|
|
| 8 |
A |
2989 |
2913 |
50.88 |
|
|
|
| 9 |
A |
2971 |
2895 |
29.92 |
|
|
|
| 10 |
A |
1529 |
1490 |
9.19 |
|
|
|
| 11 |
A |
1520 |
1481 |
13.40 |
|
|
|
| 12 |
A |
1518 |
1480 |
1.57 |
|
|
|
| 13 |
A |
1505 |
1467 |
10.67 |
|
|
|
| 14 |
A |
1494 |
1456 |
1.51 |
|
|
|
| 15 |
A |
1435 |
1398 |
17.83 |
|
|
|
| 16 |
A |
1433 |
1396 |
1.19 |
|
|
|
| 17 |
A |
1395 |
1359 |
1.31 |
|
|
|
| 18 |
A |
1327 |
1294 |
6.24 |
|
|
|
| 19 |
A |
1315 |
1281 |
17.69 |
|
|
|
| 20 |
A |
1255 |
1223 |
18.85 |
|
|
|
| 21 |
A |
1184 |
1154 |
10.75 |
|
|
|
| 22 |
A |
1134 |
1105 |
2.63 |
|
|
|
| 23 |
A |
1095 |
1067 |
3.24 |
|
|
|
| 24 |
A |
1035 |
1008 |
0.69 |
|
|
|
| 25 |
A |
1016 |
990 |
11.36 |
|
|
|
| 26 |
A |
978 |
953 |
9.38 |
|
|
|
| 27 |
A |
843 |
821 |
8.24 |
|
|
|
| 28 |
A |
797 |
776 |
14.90 |
|
|
|
| 29 |
A |
549 |
535 |
34.30 |
|
|
|
| 30 |
A |
449 |
437 |
2.34 |
|
|
|
| 31 |
A |
366 |
357 |
5.19 |
|
|
|
| 32 |
A |
312 |
304 |
2.24 |
|
|
|
| 33 |
A |
236 |
230 |
0.38 |
|
|
|
| 34 |
A |
224 |
219 |
0.13 |
|
|
|
| 35 |
A |
204 |
199 |
1.59 |
|
|
|
| 36 |
A |
105 |
102 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26832.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 26147.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.433 |
|
|
|
| 2 |
H |
0.165 |
|
|
|
| 3 |
H |
0.175 |
|
|
|
| 4 |
H |
0.149 |
|
|
|
| 5 |
C |
-0.261 |
|
|
|
| 6 |
H |
0.185 |
|
|
|
| 7 |
Cl |
-0.081 |
|
|
|
| 8 |
C |
-0.182 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
| 10 |
H |
0.154 |
|
|
|
| 11 |
C |
-0.460 |
|
|
|
| 12 |
H |
0.139 |
|
|
|
| 13 |
H |
0.165 |
|
|
|
| 14 |
H |
0.150 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.413 |
-2.765 |
0.530 |
2.845 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.905 |
-1.890 |
0.581 |
| y |
-1.890 |
-42.871 |
0.058 |
| z |
0.581 |
0.058 |
-38.874 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.967 |
-1.890 |
0.581 |
| y |
-1.890 |
-3.982 |
0.058 |
| z |
0.581 |
0.058 |
2.015 |
|
| Polar |
|---|
| 3z2-r2 | 4.030 |
|---|
| x2-y2 | 3.966 |
|---|
| xy | -1.890 |
|---|
| xz | 0.581 |
|---|
| yz | 0.058 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.831 |
0.219 |
-0.010 |
| y |
0.219 |
10.049 |
-0.391 |
| z |
-0.010 |
-0.391 |
7.146 |
<r2> (average value of r
2) Å
2
| <r2> |
139.472 |
| (<r2>)1/2 |
11.810 |