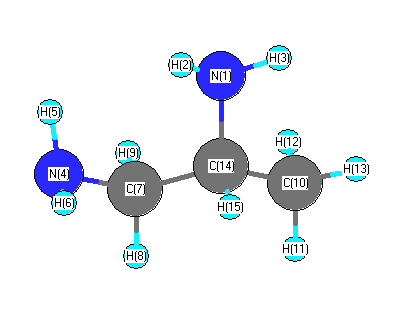
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3663 |
3570 |
0.21 |
|
|
|
| 2 |
A |
3600 |
3508 |
0.56 |
|
|
|
| 3 |
A |
3524 |
3435 |
1.24 |
|
|
|
| 4 |
A |
3477 |
3388 |
5.14 |
|
|
|
| 5 |
A |
3080 |
3001 |
65.42 |
|
|
|
| 6 |
A |
3060 |
2982 |
56.63 |
|
|
|
| 7 |
A |
3056 |
2978 |
29.14 |
|
|
|
| 8 |
A |
2990 |
2914 |
74.28 |
|
|
|
| 9 |
A |
2979 |
2903 |
25.78 |
|
|
|
| 10 |
A |
2854 |
2782 |
124.50 |
|
|
|
| 11 |
A |
1686 |
1643 |
56.74 |
|
|
|
| 12 |
A |
1680 |
1637 |
31.22 |
|
|
|
| 13 |
A |
1521 |
1483 |
4.85 |
|
|
|
| 14 |
A |
1510 |
1472 |
6.56 |
|
|
|
| 15 |
A |
1491 |
1453 |
1.43 |
|
|
|
| 16 |
A |
1423 |
1387 |
13.19 |
|
|
|
| 17 |
A |
1405 |
1369 |
0.58 |
|
|
|
| 18 |
A |
1387 |
1352 |
18.58 |
|
|
|
| 19 |
A |
1361 |
1326 |
6.77 |
|
|
|
| 20 |
A |
1344 |
1310 |
10.45 |
|
|
|
| 21 |
A |
1247 |
1215 |
1.48 |
|
|
|
| 22 |
A |
1189 |
1158 |
2.15 |
|
|
|
| 23 |
A |
1168 |
1139 |
5.83 |
|
|
|
| 24 |
A |
1104 |
1076 |
41.27 |
|
|
|
| 25 |
A |
1032 |
1006 |
7.69 |
|
|
|
| 26 |
A |
1007 |
981 |
1.99 |
|
|
|
| 27 |
A |
944 |
920 |
0.91 |
|
|
|
| 28 |
A |
872 |
850 |
2.70 |
|
|
|
| 29 |
A |
825 |
804 |
17.84 |
|
|
|
| 30 |
A |
673 |
656 |
185.07 |
|
|
|
| 31 |
A |
509 |
496 |
96.46 |
|
|
|
| 32 |
A |
465 |
453 |
36.24 |
|
|
|
| 33 |
A |
424 |
413 |
165.87 |
|
|
|
| 34 |
A |
360 |
351 |
6.75 |
|
|
|
| 35 |
A |
325 |
317 |
106.61 |
|
|
|
| 36 |
A |
251 |
244 |
10.51 |
|
|
|
| 37 |
A |
217 |
212 |
2.64 |
|
|
|
| 38 |
A |
183 |
179 |
93.86 |
|
|
|
| 39 |
A |
120 |
117 |
6.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30002.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29237.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.614 |
|
|
|
| 2 |
H |
0.243 |
|
|
|
| 3 |
H |
0.262 |
|
|
|
| 4 |
N |
-0.574 |
|
|
|
| 5 |
H |
0.271 |
|
|
|
| 6 |
H |
0.261 |
|
|
|
| 7 |
C |
-0.238 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
| 10 |
C |
-0.521 |
|
|
|
| 11 |
H |
0.138 |
|
|
|
| 12 |
H |
0.154 |
|
|
|
| 13 |
H |
0.138 |
|
|
|
| 14 |
C |
0.132 |
|
|
|
| 15 |
H |
0.089 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.494 |
1.105 |
1.566 |
1.979 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.461 |
-0.191 |
-0.052 |
| y |
-0.191 |
7.844 |
-0.144 |
| z |
-0.052 |
-0.144 |
6.987 |
<r2> (average value of r
2) Å
2
| <r2> |
124.480 |
| (<r2>)1/2 |
11.157 |