Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/CEP-121G
Geometric Data calculated at B3LYP/CEP-121G
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -43.338911 |
| Energy at 298.15K | -43.347124 |
| Nuclear repulsion energy | 97.315783 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3125 |
3045 |
19.45 |
|
|
|
| 2 |
A |
3101 |
3022 |
39.58 |
|
|
|
| 3 |
A |
3088 |
3009 |
41.71 |
|
|
|
| 4 |
A |
3069 |
2991 |
22.92 |
|
|
|
| 5 |
A |
3013 |
2937 |
18.08 |
|
|
|
| 6 |
A |
3003 |
2927 |
29.83 |
|
|
|
| 7 |
A |
2993 |
2916 |
15.59 |
|
|
|
| 8 |
A |
2974 |
2898 |
26.40 |
|
|
|
| 9 |
A |
1665 |
1622 |
105.90 |
|
|
|
| 10 |
A |
1521 |
1482 |
10.97 |
|
|
|
| 11 |
A |
1512 |
1474 |
9.13 |
|
|
|
| 12 |
A |
1499 |
1461 |
13.77 |
|
|
|
| 13 |
A |
1488 |
1450 |
37.68 |
|
|
|
| 14 |
A |
1485 |
1447 |
3.93 |
|
|
|
| 15 |
A |
1436 |
1400 |
11.35 |
|
|
|
| 16 |
A |
1415 |
1379 |
35.68 |
|
|
|
| 17 |
A |
1376 |
1341 |
22.14 |
|
|
|
| 18 |
A |
1282 |
1250 |
0.15 |
|
|
|
| 19 |
A |
1201 |
1171 |
70.13 |
|
|
|
| 20 |
A |
1141 |
1112 |
0.33 |
|
|
|
| 21 |
A |
1108 |
1080 |
3.23 |
|
|
|
| 22 |
A |
1008 |
983 |
4.26 |
|
|
|
| 23 |
A |
975 |
950 |
3.61 |
|
|
|
| 24 |
A |
960 |
936 |
9.69 |
|
|
|
| 25 |
A |
771 |
752 |
3.73 |
|
|
|
| 26 |
A |
756 |
737 |
2.25 |
|
|
|
| 27 |
A |
577 |
562 |
8.93 |
|
|
|
| 28 |
A |
472 |
460 |
0.48 |
|
|
|
| 29 |
A |
392 |
382 |
3.96 |
|
|
|
| 30 |
A |
243 |
237 |
5.73 |
|
|
|
| 31 |
A |
183 |
178 |
0.25 |
|
|
|
| 32 |
A |
103 |
101 |
0.11 |
|
|
|
| 33 |
A |
47 |
46 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24490.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 23866.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.925 |
-0.475 |
0.041 |
| C2 |
0.542 |
0.181 |
-0.024 |
| C3 |
-0.678 |
-0.756 |
-0.073 |
| C4 |
-2.034 |
-0.027 |
0.059 |
| O5 |
0.416 |
1.432 |
-0.031 |
| H6 |
2.701 |
0.275 |
-0.132 |
| H7 |
2.078 |
-0.922 |
1.035 |
| H8 |
2.015 |
-1.285 |
-0.696 |
| H9 |
-2.864 |
-0.741 |
-0.018 |
| H10 |
-2.109 |
0.491 |
1.022 |
| H11 |
-2.148 |
0.730 |
-0.726 |
| H12 |
-0.627 |
-1.312 |
-1.025 |
| H13 |
-0.559 |
-1.517 |
0.715 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5323 | 2.6210 | 3.9844 | 2.4330 | 1.0933 | 1.1002 | 1.0981 | 4.7965 | 4.2623 | 4.3161 | 2.8897 | 2.7762 |
C2 | 1.5323 | | 1.5390 | 2.5854 | 1.2573 | 2.1639 | 2.1673 | 2.1839 | 3.5280 | 2.8661 | 2.8334 | 2.1439 | 2.1538 | C3 | 2.6210 | 1.5390 | | 1.5451 | 2.4465 | 3.5336 | 2.9756 | 2.8147 | 2.1861 | 2.1913 | 2.1892 | 1.1035 | 1.1020 | C4 | 3.9844 | 2.5854 | 1.5451 | | 2.8525 | 4.7483 | 4.3204 | 4.3068 | 1.0977 | 1.0966 | 1.0956 | 2.1922 | 2.1977 | O5 | 2.4330 | 1.2573 | 2.4465 | 2.8525 | | 2.5628 | 3.0723 | 3.2213 | 3.9341 | 2.8926 | 2.7473 | 3.0986 | 3.1942 | H6 | 1.0933 | 2.1639 | 3.5336 | 4.7483 | 2.5628 | | 1.7839 | 1.7947 | 5.6578 | 4.9506 | 4.9059 | 3.7937 | 3.8149 | H7 | 1.1002 | 2.1673 | 2.9756 | 4.3204 | 3.0723 | 1.7839 | | 1.7694 | 5.0561 | 4.4189 | 4.8670 | 3.4228 | 2.7221 | H8 | 1.0981 | 2.1839 | 2.8147 | 4.3068 | 3.2213 | 1.7947 | 1.7694 | | 4.9558 | 4.8074 | 4.6249 | 2.6630 | 2.9440 | H9 | 4.7965 | 3.5280 | 2.1861 | 1.0977 | 3.9341 | 5.6578 | 5.0561 | 4.9558 | | 1.7807 | 1.7826 | 2.5185 | 2.5402 | H10 | 4.2623 | 2.8661 | 2.1913 | 1.0966 | 2.8926 | 4.9506 | 4.4189 | 4.8074 | 1.7807 | | 1.7648 | 3.1045 | 2.5559 | H11 | 4.3161 | 2.8334 | 2.1892 | 1.0956 | 2.7473 | 4.9059 | 4.8670 | 4.6249 | 1.7826 | 1.7648 | | 2.5631 | 3.1065 | H12 | 2.8897 | 2.1439 | 1.1035 | 2.1922 | 3.0986 | 3.7937 | 3.4228 | 2.6630 | 2.5185 | 3.1045 | 2.5631 | | 1.7532 | H13 | 2.7762 | 2.1538 | 1.1020 | 2.1977 | 3.1942 | 3.8149 | 2.7221 | 2.9440 | 2.5402 | 2.5559 | 3.1065 | 1.7532 | |
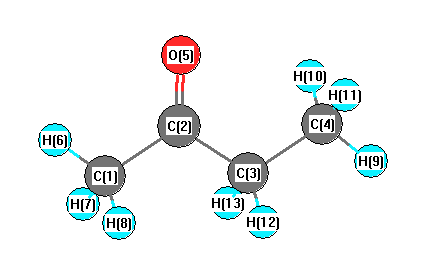 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
117.161 |
|
C1 |
C2 |
O5 |
121.100 |
| C2 |
C1 |
H6 |
109.872 |
|
C2 |
C1 |
H7 |
109.739 |
| C2 |
C1 |
H8 |
111.176 |
|
C2 |
C3 |
C4 |
113.922 |
| C2 |
C3 |
H12 |
107.300 |
|
C2 |
C3 |
H13 |
108.136 |
| C3 |
C2 |
O5 |
121.738 |
|
C3 |
C4 |
H9 |
110.481 |
| C3 |
C4 |
H10 |
110.952 |
|
C3 |
C4 |
H11 |
110.853 |
| C4 |
C3 |
H12 |
110.614 |
|
C4 |
C3 |
H13 |
111.141 |
| H6 |
C1 |
H7 |
108.833 |
|
H6 |
C1 |
H8 |
109.961 |
| H7 |
C1 |
H8 |
107.196 |
|
H9 |
C4 |
H10 |
108.490 |
| H9 |
C4 |
H11 |
108.736 |
|
H10 |
C4 |
H11 |
107.224 |
| H12 |
C3 |
H13 |
105.290 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.561 |
|
|
|
| 2 |
C |
0.161 |
|
|
|
| 3 |
C |
-0.280 |
|
|
|
| 4 |
C |
-0.464 |
|
|
|
| 5 |
O |
-0.159 |
|
|
|
| 6 |
H |
0.188 |
|
|
|
| 7 |
H |
0.173 |
|
|
|
| 8 |
H |
0.163 |
|
|
|
| 9 |
H |
0.144 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
| 11 |
H |
0.164 |
|
|
|
| 12 |
H |
0.160 |
|
|
|
| 13 |
H |
0.154 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.247 |
-3.248 |
0.039 |
3.258 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.998 |
-1.350 |
0.196 |
| y |
-1.350 |
-36.489 |
0.085 |
| z |
0.196 |
0.085 |
-30.874 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.684 |
-1.350 |
0.196 |
| y |
-1.350 |
-6.053 |
0.085 |
| z |
0.196 |
0.085 |
2.369 |
|
| Polar |
|---|
| 3z2-r2 | 4.739 |
|---|
| x2-y2 | 6.491 |
|---|
| xy | -1.350 |
|---|
| xz | 0.196 |
|---|
| yz | 0.085 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.011 |
-0.192 |
0.055 |
| y |
-0.192 |
7.795 |
0.009 |
| z |
0.055 |
0.009 |
5.916 |
<r2> (average value of r
2) Å
2
| <r2> |
117.634 |
| (<r2>)1/2 |
10.846 |
