Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -58.231715 |
| Energy at 298.15K | -58.234083 |
| HF Energy | -58.231715 |
| Nuclear repulsion energy | 82.020829 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3163 |
3082 |
0.91 |
|
|
|
| 2 |
A' |
3023 |
2946 |
36.40 |
|
|
|
| 3 |
A' |
1673 |
1631 |
123.21 |
|
|
|
| 4 |
A' |
1396 |
1360 |
6.33 |
|
|
|
| 5 |
A' |
1215 |
1184 |
15.30 |
|
|
|
| 6 |
A' |
1062 |
1035 |
21.30 |
|
|
|
| 7 |
A' |
736 |
717 |
30.28 |
|
|
|
| 8 |
A' |
410 |
399 |
6.21 |
|
|
|
| 9 |
A' |
308 |
300 |
31.63 |
|
|
|
| 10 |
A' |
236 |
230 |
2.16 |
|
|
|
| 11 |
A" |
1232 |
1200 |
22.22 |
|
|
|
| 12 |
A" |
1007 |
981 |
28.86 |
|
|
|
| 13 |
A" |
650 |
634 |
139.30 |
|
|
|
| 14 |
A" |
270 |
263 |
4.29 |
|
|
|
| 15 |
A" |
86 |
84 |
11.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8232.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 8022.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.404 |
0.011 |
0.000 |
| C2 |
-0.179 |
1.429 |
0.000 |
| H3 |
1.493 |
0.004 |
0.000 |
| Cl4 |
-0.179 |
-0.870 |
1.546 |
| Cl5 |
-0.179 |
-0.870 |
-1.546 |
| O6 |
0.563 |
2.428 |
0.000 |
| H7 |
-1.279 |
1.495 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5337 | 1.0886 | 1.8723 | 1.8723 | 2.4228 | 2.2440 |
C2 | 1.5337 | | 2.1968 | 2.7706 | 2.7706 | 1.2439 | 1.1021 | H3 | 1.0886 | 2.1968 | | 2.4387 | 2.4387 | 2.5968 | 3.1473 | Cl4 | 1.8723 | 2.7706 | 2.4387 | | 3.0926 | 3.7171 | 3.0319 | Cl5 | 1.8723 | 2.7706 | 2.4387 | 3.0926 | | 3.7171 | 3.0319 | O6 | 2.4228 | 1.2439 | 2.5968 | 3.7171 | 3.7171 | | 2.0644 | H7 | 2.2440 | 1.1021 | 3.1473 | 3.0319 | 3.0319 | 2.0644 | |
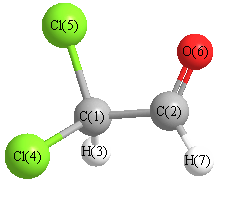 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
121.094 |
|
C1 |
C2 |
H7 |
115.747 |
| C2 |
C1 |
H3 |
112.695 |
|
C2 |
C1 |
Cl4 |
108.458 |
| C2 |
C1 |
Cl5 |
108.458 |
|
H3 |
C1 |
Cl4 |
107.956 |
| H3 |
C1 |
Cl5 |
107.956 |
|
Cl4 |
C1 |
Cl5 |
111.355 |
| O6 |
C2 |
H7 |
123.159 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.553 |
|
|
|
| 2 |
C |
0.084 |
|
|
|
| 3 |
H |
0.272 |
|
|
|
| 4 |
Cl |
0.082 |
|
|
|
| 5 |
Cl |
0.082 |
|
|
|
| 6 |
O |
-0.115 |
|
|
|
| 7 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.340 |
-0.619 |
0.000 |
0.706 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.852 |
-3.750 |
0.000 |
| y |
-3.750 |
-49.197 |
0.000 |
| z |
0.000 |
0.000 |
-43.001 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.247 |
-3.750 |
0.000 |
| y |
-3.750 |
-7.771 |
0.000 |
| z |
0.000 |
0.000 |
1.524 |
|
| Polar |
|---|
| 3z2-r2 | 3.047 |
|---|
| x2-y2 | 9.345 |
|---|
| xy | -3.750 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.961 |
1.013 |
0.000 |
| y |
1.013 |
7.375 |
0.000 |
| z |
0.000 |
0.000 |
8.101 |
<r2> (average value of r
2) Å
2
| <r2> |
124.331 |
| (<r2>)1/2 |
11.150 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -58.227644 |
| Energy at 298.15K | |
| HF Energy | -58.227644 |
| Nuclear repulsion energy | 82.668245 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3111 |
3032 |
4.76 |
90.35 |
0.16 |
0.28 |
| 2 |
A |
2983 |
2907 |
41.61 |
149.14 |
0.32 |
0.48 |
| 3 |
A |
1687 |
1644 |
116.64 |
23.84 |
0.35 |
0.52 |
| 4 |
A |
1396 |
1361 |
13.39 |
5.36 |
0.66 |
0.80 |
| 5 |
A |
1249 |
1217 |
12.20 |
8.15 |
0.70 |
0.82 |
| 6 |
A |
1209 |
1178 |
11.28 |
14.20 |
0.69 |
0.82 |
| 7 |
A |
1017 |
991 |
14.70 |
1.70 |
0.70 |
0.82 |
| 8 |
A |
924 |
901 |
3.91 |
8.33 |
0.54 |
0.70 |
| 9 |
A |
760 |
740 |
81.21 |
8.60 |
0.71 |
0.83 |
| 10 |
A |
597 |
582 |
37.49 |
17.00 |
0.08 |
0.15 |
| 11 |
A |
571 |
556 |
56.97 |
13.37 |
0.52 |
0.69 |
| 12 |
A |
325 |
316 |
2.69 |
6.53 |
0.24 |
0.38 |
| 13 |
A |
252 |
246 |
4.59 |
8.70 |
0.62 |
0.77 |
| 14 |
A |
209 |
204 |
2.32 |
2.96 |
0.73 |
0.85 |
| 15 |
A |
80 |
78 |
14.35 |
2.53 |
0.70 |
0.83 |
Unscaled Zero Point Vibrational Energy (zpe) 8185.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 7976.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.082 |
-0.040 |
0.543 |
| C2 |
-0.652 |
-1.352 |
0.201 |
| H3 |
0.222 |
0.071 |
1.621 |
| Cl4 |
1.825 |
-0.181 |
-0.171 |
| Cl5 |
-0.798 |
1.467 |
-0.061 |
| O6 |
-1.771 |
-1.415 |
-0.330 |
| H7 |
-0.083 |
-2.256 |
0.487 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5425 | 1.0924 | 1.8883 | 1.8463 | 2.4668 | 2.2228 |
C2 | 1.5425 | | 2.1923 | 2.7651 | 2.8351 | 1.2393 | 1.1054 | H3 | 1.0924 | 2.1923 | | 2.4168 | 2.4116 | 3.1594 | 2.6060 | Cl4 | 1.8883 | 2.7651 | 2.4168 | | 3.0995 | 3.8045 | 2.8942 | Cl5 | 1.8463 | 2.8351 | 2.4116 | 3.0995 | | 3.0534 | 3.8299 | O6 | 2.4668 | 1.2393 | 3.1594 | 3.8045 | 3.0534 | | 2.0546 | H7 | 2.2228 | 1.1054 | 2.6060 | 2.8942 | 3.8299 | 2.0546 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.582 |
|
C1 |
C2 |
H7 |
113.127 |
| C2 |
C1 |
H3 |
111.462 |
|
C2 |
C1 |
Cl4 |
106.972 |
| C2 |
C1 |
Cl5 |
113.260 |
|
H3 |
C1 |
Cl4 |
105.203 |
| H3 |
C1 |
Cl5 |
107.510 |
|
Cl4 |
C1 |
Cl5 |
112.183 |
| O6 |
C2 |
H7 |
122.289 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.572 |
|
|
|
| 2 |
C |
0.069 |
|
|
|
| 3 |
H |
0.269 |
|
|
|
| 4 |
Cl |
0.066 |
|
|
|
| 5 |
Cl |
0.113 |
|
|
|
| 6 |
O |
-0.091 |
|
|
|
| 7 |
H |
0.146 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.380 |
-0.940 |
2.382 |
2.909 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.440 |
-2.623 |
-1.328 |
| y |
-2.623 |
-41.229 |
-1.804 |
| z |
-1.328 |
-1.804 |
-39.275 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.188 |
-2.623 |
-1.328 |
| y |
-2.623 |
2.628 |
-1.804 |
| z |
-1.328 |
-1.804 |
5.560 |
|
| Polar |
|---|
| 3z2-r2 | 11.120 |
|---|
| x2-y2 | -7.211 |
|---|
| xy | -2.623 |
|---|
| xz | -1.328 |
|---|
| yz | -1.804 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.779 |
-0.938 |
0.186 |
| y |
-0.938 |
6.928 |
-0.406 |
| z |
0.186 |
-0.406 |
4.291 |
<r2> (average value of r
2) Å
2
| <r2> |
120.013 |
| (<r2>)1/2 |
10.955 |
