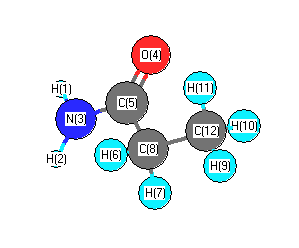
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3718 |
3623 |
16.03 |
|
|
|
| 2 |
A |
3558 |
3468 |
19.36 |
|
|
|
| 3 |
A |
3111 |
3031 |
28.49 |
|
|
|
| 4 |
A |
3083 |
3005 |
53.45 |
|
|
|
| 5 |
A |
3040 |
2962 |
27.02 |
|
|
|
| 6 |
A |
3001 |
2925 |
26.07 |
|
|
|
| 7 |
A |
2992 |
2915 |
43.06 |
|
|
|
| 8 |
A |
1670 |
1628 |
274.04 |
|
|
|
| 9 |
A |
1635 |
1593 |
82.60 |
|
|
|
| 10 |
A |
1530 |
1491 |
12.62 |
|
|
|
| 11 |
A |
1516 |
1477 |
9.87 |
|
|
|
| 12 |
A |
1496 |
1458 |
9.62 |
|
|
|
| 13 |
A |
1431 |
1394 |
14.10 |
|
|
|
| 14 |
A |
1398 |
1362 |
72.18 |
|
|
|
| 15 |
A |
1311 |
1278 |
67.19 |
|
|
|
| 16 |
A |
1280 |
1247 |
50.21 |
|
|
|
| 17 |
A |
1127 |
1098 |
4.95 |
|
|
|
| 18 |
A |
1105 |
1077 |
3.33 |
|
|
|
| 19 |
A |
1059 |
1032 |
7.99 |
|
|
|
| 20 |
A |
1014 |
988 |
3.11 |
|
|
|
| 21 |
A |
815 |
794 |
11.98 |
|
|
|
| 22 |
A |
790 |
769 |
1.89 |
|
|
|
| 23 |
A |
679 |
662 |
54.50 |
|
|
|
| 24 |
A |
598 |
583 |
71.10 |
|
|
|
| 25 |
A |
519 |
506 |
103.58 |
|
|
|
| 26 |
A |
491 |
478 |
95.46 |
|
|
|
| 27 |
A |
423 |
412 |
3.83 |
|
|
|
| 28 |
A |
254 |
247 |
8.81 |
|
|
|
| 29 |
A |
206 |
201 |
1.13 |
|
|
|
| 30 |
A |
47 |
46 |
4.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22447.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 21874.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.325 |
|
|
|
| 2 |
H |
0.314 |
|
|
|
| 3 |
N |
-0.499 |
|
|
|
| 4 |
O |
-0.205 |
|
|
|
| 5 |
C |
0.121 |
|
|
|
| 6 |
H |
0.171 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
C |
-0.351 |
|
|
|
| 9 |
H |
0.145 |
|
|
|
| 10 |
H |
0.148 |
|
|
|
| 11 |
H |
0.185 |
|
|
|
| 12 |
C |
-0.483 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.651 |
-4.123 |
-0.201 |
4.179 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.574 |
3.419 |
1.398 |
| y |
3.419 |
-34.299 |
-0.186 |
| z |
1.398 |
-0.186 |
-31.708 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.430 |
3.419 |
1.398 |
| y |
3.419 |
-5.658 |
-0.186 |
| z |
1.398 |
-0.186 |
-1.772 |
|
| Polar |
|---|
| 3z2-r2 | -3.544 |
|---|
| x2-y2 | 8.726 |
|---|
| xy | 3.419 |
|---|
| xz | 1.398 |
|---|
| yz | -0.186 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.579 |
0.355 |
0.286 |
| y |
0.355 |
7.185 |
0.068 |
| z |
0.286 |
0.068 |
5.098 |
<r2> (average value of r
2) Å
2
| <r2> |
109.394 |
| (<r2>)1/2 |
10.459 |