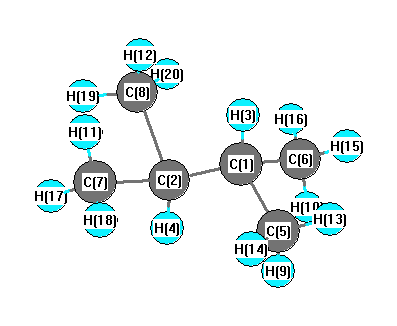
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3085 |
3006 |
0.00 |
|
|
|
| 2 |
Ag |
3057 |
2979 |
0.00 |
|
|
|
| 3 |
Ag |
2980 |
2904 |
0.00 |
|
|
|
| 4 |
Ag |
2937 |
2862 |
0.00 |
|
|
|
| 5 |
Ag |
1545 |
1506 |
0.00 |
|
|
|
| 6 |
Ag |
1523 |
1484 |
0.00 |
|
|
|
| 7 |
Ag |
1438 |
1401 |
0.00 |
|
|
|
| 8 |
Ag |
1372 |
1337 |
0.00 |
|
|
|
| 9 |
Ag |
1216 |
1185 |
0.00 |
|
|
|
| 10 |
Ag |
1197 |
1167 |
0.00 |
|
|
|
| 11 |
Ag |
945 |
921 |
0.00 |
|
|
|
| 12 |
Ag |
747 |
728 |
0.00 |
|
|
|
| 13 |
Ag |
491 |
479 |
0.00 |
|
|
|
| 14 |
Ag |
375 |
366 |
0.00 |
|
|
|
| 15 |
Ag |
233 |
227 |
0.00 |
|
|
|
| 16 |
Au |
3083 |
3004 |
96.18 |
|
|
|
| 17 |
Au |
3048 |
2970 |
1.12 |
|
|
|
| 18 |
Au |
2973 |
2897 |
69.14 |
|
|
|
| 19 |
Au |
1528 |
1489 |
5.46 |
|
|
|
| 20 |
Au |
1505 |
1467 |
2.69 |
|
|
|
| 21 |
Au |
1420 |
1384 |
16.02 |
|
|
|
| 22 |
Au |
1336 |
1302 |
4.35 |
|
|
|
| 23 |
Au |
1083 |
1055 |
4.85 |
|
|
|
| 24 |
Au |
976 |
951 |
0.14 |
|
|
|
| 25 |
Au |
934 |
910 |
2.59 |
|
|
|
| 26 |
Au |
297 |
289 |
0.00 |
|
|
|
| 27 |
Au |
191 |
187 |
0.05 |
|
|
|
| 28 |
Au |
58 |
57 |
0.00 |
|
|
|
| 29 |
Bg |
3067 |
2989 |
0.00 |
|
|
|
| 30 |
Bg |
3048 |
2971 |
0.00 |
|
|
|
| 31 |
Bg |
2971 |
2896 |
0.00 |
|
|
|
| 32 |
Bg |
1514 |
1475 |
0.00 |
|
|
|
| 33 |
Bg |
1510 |
1472 |
0.00 |
|
|
|
| 34 |
Bg |
1411 |
1375 |
0.00 |
|
|
|
| 35 |
Bg |
1355 |
1320 |
0.00 |
|
|
|
| 36 |
Bg |
1171 |
1141 |
0.00 |
|
|
|
| 37 |
Bg |
962 |
938 |
0.00 |
|
|
|
| 38 |
Bg |
927 |
903 |
0.00 |
|
|
|
| 39 |
Bg |
420 |
410 |
0.00 |
|
|
|
| 40 |
Bg |
191 |
186 |
0.00 |
|
|
|
| 41 |
Bu |
3073 |
2995 |
130.45 |
|
|
|
| 42 |
Bu |
3061 |
2983 |
246.35 |
|
|
|
| 43 |
Bu |
2976 |
2900 |
101.61 |
|
|
|
| 44 |
Bu |
2953 |
2878 |
41.95 |
|
|
|
| 45 |
Bu |
1531 |
1492 |
25.08 |
|
|
|
| 46 |
Bu |
1523 |
1484 |
17.73 |
|
|
|
| 47 |
Bu |
1430 |
1394 |
20.67 |
|
|
|
| 48 |
Bu |
1315 |
1281 |
5.95 |
|
|
|
| 49 |
Bu |
1177 |
1147 |
10.96 |
|
|
|
| 50 |
Bu |
1008 |
982 |
7.78 |
|
|
|
| 51 |
Bu |
862 |
840 |
1.98 |
|
|
|
| 52 |
Bu |
399 |
389 |
0.11 |
|
|
|
| 53 |
Bu |
348 |
339 |
0.18 |
|
|
|
| 54 |
Bu |
212 |
206 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40993.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 39947.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.089 |
|
|
|
| 2 |
C |
0.089 |
|
|
|
| 3 |
H |
0.102 |
|
|
|
| 4 |
H |
0.102 |
|
|
|
| 5 |
C |
-0.521 |
|
|
|
| 6 |
C |
-0.521 |
|
|
|
| 7 |
C |
-0.521 |
|
|
|
| 8 |
C |
-0.521 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
| 10 |
H |
0.136 |
|
|
|
| 11 |
H |
0.136 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
| 13 |
H |
0.138 |
|
|
|
| 14 |
H |
0.151 |
|
|
|
| 15 |
H |
0.138 |
|
|
|
| 16 |
H |
0.151 |
|
|
|
| 17 |
H |
0.138 |
|
|
|
| 18 |
H |
0.151 |
|
|
|
| 19 |
H |
0.138 |
|
|
|
| 20 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.402 |
-0.507 |
0.000 |
| y |
-0.507 |
-42.403 |
0.000 |
| z |
0.000 |
0.000 |
-42.898 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.249 |
-0.507 |
0.000 |
| y |
-0.507 |
0.247 |
0.000 |
| z |
0.000 |
0.000 |
-0.495 |
|
| Polar |
|---|
| 3z2-r2 | -0.991 |
|---|
| x2-y2 | 0.001 |
|---|
| xy | -0.507 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.967 |
0.698 |
0.000 |
| y |
0.698 |
11.239 |
0.000 |
| z |
0.000 |
0.000 |
10.951 |
<r2> (average value of r
2) Å
2
| <r2> |
179.589 |
| (<r2>)1/2 |
13.401 |