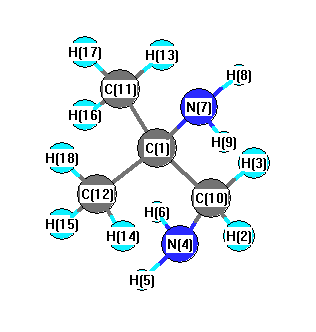
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3519 |
3429 |
1.41 |
|
|
|
| 2 |
A |
3457 |
3368 |
6.02 |
|
|
|
| 3 |
A |
3093 |
3014 |
72.08 |
|
|
|
| 4 |
A |
3062 |
2984 |
84.45 |
|
|
|
| 5 |
A |
2982 |
2906 |
1.66 |
|
|
|
| 6 |
A |
2970 |
2895 |
64.11 |
|
|
|
| 7 |
A |
1691 |
1648 |
50.00 |
|
|
|
| 8 |
A |
1689 |
1646 |
20.85 |
|
|
|
| 9 |
A |
1527 |
1488 |
11.49 |
|
|
|
| 10 |
A |
1523 |
1484 |
7.91 |
|
|
|
| 11 |
A |
1492 |
1454 |
0.86 |
|
|
|
| 12 |
A |
1427 |
1390 |
13.12 |
|
|
|
| 13 |
A |
1375 |
1340 |
2.99 |
|
|
|
| 14 |
A |
1251 |
1219 |
15.93 |
|
|
|
| 15 |
A |
1223 |
1192 |
16.61 |
|
|
|
| 16 |
A |
1095 |
1067 |
36.01 |
|
|
|
| 17 |
A |
1002 |
976 |
1.91 |
|
|
|
| 18 |
A |
908 |
885 |
4.94 |
|
|
|
| 19 |
A |
736 |
718 |
7.43 |
|
|
|
| 20 |
A |
657 |
641 |
286.54 |
|
|
|
| 21 |
A |
553 |
539 |
167.07 |
|
|
|
| 22 |
A |
479 |
467 |
166.13 |
|
|
|
| 23 |
A |
410 |
400 |
35.39 |
|
|
|
| 24 |
A |
351 |
342 |
1.07 |
|
|
|
| 25 |
A |
252 |
245 |
3.05 |
|
|
|
| 26 |
A |
238 |
232 |
3.26 |
|
|
|
| 27 |
A |
3651 |
3558 |
0.01 |
|
|
|
| 28 |
A |
3584 |
3492 |
1.61 |
|
|
|
| 29 |
A |
3086 |
3007 |
1.04 |
|
|
|
| 30 |
A |
3058 |
2980 |
27.04 |
|
|
|
| 31 |
A |
3027 |
2950 |
50.83 |
|
|
|
| 32 |
A |
2974 |
2898 |
71.08 |
|
|
|
| 33 |
A |
1511 |
1473 |
1.27 |
|
|
|
| 34 |
A |
1506 |
1467 |
0.06 |
|
|
|
| 35 |
A |
1411 |
1375 |
18.94 |
|
|
|
| 36 |
A |
1388 |
1353 |
0.02 |
|
|
|
| 37 |
A |
1327 |
1293 |
1.56 |
|
|
|
| 38 |
A |
1157 |
1127 |
0.16 |
|
|
|
| 39 |
A |
1035 |
1009 |
1.32 |
|
|
|
| 40 |
A |
1003 |
978 |
0.34 |
|
|
|
| 41 |
A |
949 |
924 |
0.08 |
|
|
|
| 42 |
A |
848 |
826 |
0.84 |
|
|
|
| 43 |
A |
441 |
429 |
1.70 |
|
|
|
| 44 |
A |
337 |
328 |
7.45 |
|
|
|
| 45 |
A |
280 |
273 |
10.59 |
|
|
|
| 46 |
A |
236 |
230 |
77.59 |
|
|
|
| 47 |
A |
203 |
198 |
0.10 |
|
|
|
| 48 |
A |
116 |
113 |
0.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36042.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 35123.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.378 |
|
|
|
| 2 |
H |
0.122 |
|
|
|
| 3 |
H |
0.122 |
|
|
|
| 4 |
N |
-0.547 |
|
|
|
| 5 |
H |
0.266 |
|
|
|
| 6 |
H |
0.266 |
|
|
|
| 7 |
N |
-0.602 |
|
|
|
| 8 |
H |
0.255 |
|
|
|
| 9 |
H |
0.255 |
|
|
|
| 10 |
C |
-0.279 |
|
|
|
| 11 |
C |
-0.539 |
|
|
|
| 12 |
C |
-0.539 |
|
|
|
| 13 |
H |
0.130 |
|
|
|
| 14 |
H |
0.130 |
|
|
|
| 15 |
H |
0.127 |
|
|
|
| 16 |
H |
0.127 |
|
|
|
| 17 |
H |
0.163 |
|
|
|
| 18 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.212 |
-0.297 |
0.000 |
0.365 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.502 |
5.285 |
0.000 |
| y |
5.285 |
-47.178 |
0.000 |
| z |
0.000 |
0.000 |
-37.390 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.782 |
5.285 |
0.000 |
| y |
5.285 |
-9.232 |
0.000 |
| z |
0.000 |
0.000 |
5.450 |
|
| Polar |
|---|
| 3z2-r2 | 10.899 |
|---|
| x2-y2 | 8.676 |
|---|
| xy | 5.285 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.286 |
0.274 |
0.000 |
| y |
0.274 |
9.536 |
0.000 |
| z |
0.000 |
0.000 |
9.568 |
<r2> (average value of r
2) Å
2
| <r2> |
158.430 |
| (<r2>)1/2 |
12.587 |