Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -44.077554 |
| Energy at 298.15K | |
| HF Energy | -44.077554 |
| Nuclear repulsion energy | 63.986998 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3230 |
3147 |
14.25 |
55.31 |
0.72 |
0.84 |
| 2 |
A' |
3130 |
3050 |
10.74 |
183.87 |
0.14 |
0.24 |
| 3 |
A' |
3110 |
3030 |
15.40 |
32.06 |
0.72 |
0.84 |
| 4 |
A' |
3007 |
2931 |
43.78 |
186.89 |
0.12 |
0.22 |
| 5 |
A' |
1684 |
1641 |
4.09 |
20.03 |
0.09 |
0.17 |
| 6 |
A' |
1508 |
1470 |
3.31 |
21.14 |
0.62 |
0.76 |
| 7 |
A' |
1451 |
1414 |
6.06 |
13.09 |
0.45 |
0.63 |
| 8 |
A' |
1395 |
1360 |
10.72 |
14.28 |
0.64 |
0.78 |
| 9 |
A' |
1316 |
1283 |
0.07 |
21.98 |
0.34 |
0.51 |
| 10 |
A' |
1102 |
1074 |
15.86 |
2.64 |
0.75 |
0.86 |
| 11 |
A' |
972 |
947 |
52.53 |
6.52 |
0.69 |
0.81 |
| 12 |
A' |
887 |
865 |
12.23 |
7.90 |
0.13 |
0.23 |
| 13 |
A' |
588 |
573 |
7.06 |
2.71 |
0.75 |
0.86 |
| 14 |
A' |
258 |
251 |
4.22 |
1.49 |
0.57 |
0.73 |
| 15 |
A" |
3068 |
2990 |
38.79 |
95.99 |
0.75 |
0.86 |
| 16 |
A" |
1210 |
1179 |
0.62 |
12.12 |
0.75 |
0.86 |
| 17 |
A" |
1040 |
1013 |
10.46 |
1.03 |
0.75 |
0.86 |
| 18 |
A" |
1019 |
993 |
23.32 |
0.15 |
0.75 |
0.86 |
| 19 |
A" |
977 |
953 |
55.93 |
1.60 |
0.75 |
0.86 |
| 20 |
A" |
565 |
551 |
15.61 |
9.24 |
0.75 |
0.86 |
| 21 |
A" |
145 |
141 |
4.47 |
4.42 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 15831.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 15427.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.973 |
-0.182 |
0.000 |
| C2 |
0.000 |
0.978 |
0.000 |
| C3 |
1.345 |
0.852 |
0.000 |
| F4 |
-0.261 |
-1.454 |
0.000 |
| H5 |
1.991 |
1.727 |
0.000 |
| H6 |
1.818 |
-0.127 |
0.000 |
| H7 |
-0.473 |
1.963 |
0.000 |
| H8 |
-1.608 |
-0.182 |
0.897 |
| H9 |
-1.608 |
-0.182 |
-0.897 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5141 | 2.5380 | 1.4575 | 3.5255 | 2.7911 | 2.2025 | 1.0987 | 1.0987 |
C2 | 1.5141 | | 1.3505 | 2.4459 | 2.1270 | 2.1268 | 1.0926 | 2.1761 | 2.1761 | C3 | 2.5380 | 1.3505 | | 2.8101 | 1.0877 | 1.0870 | 2.1302 | 3.2546 | 3.2546 | F4 | 1.4575 | 2.4459 | 2.8101 | | 3.8976 | 2.4664 | 3.4234 | 2.0582 | 2.0582 | H5 | 3.5255 | 2.1270 | 1.0877 | 3.8976 | | 1.8620 | 2.4748 | 4.1713 | 4.1713 | H6 | 2.7911 | 2.1268 | 1.0870 | 2.4664 | 1.8620 | | 3.1003 | 3.5415 | 3.5415 | H7 | 2.2025 | 1.0926 | 2.1302 | 3.4234 | 2.4748 | 3.1003 | | 2.5870 | 2.5870 | H8 | 1.0987 | 2.1761 | 3.2546 | 2.0582 | 4.1713 | 3.5415 | 2.5870 | | 1.7932 | H9 | 1.0987 | 2.1761 | 3.2546 | 2.0582 | 4.1713 | 3.5415 | 2.5870 | 1.7932 | |
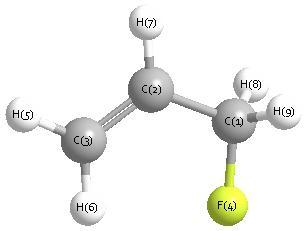 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.645 |
|
C1 |
C2 |
H7 |
114.358 |
| C2 |
C1 |
F4 |
110.774 |
|
C2 |
C1 |
H8 |
111.800 |
| C2 |
C1 |
H9 |
111.800 |
|
C2 |
C3 |
H5 |
121.084 |
| C2 |
C3 |
H6 |
121.132 |
|
C3 |
C2 |
H7 |
120.997 |
| F4 |
C1 |
H8 |
106.405 |
|
F4 |
C1 |
H9 |
106.405 |
| H5 |
C3 |
H6 |
117.784 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.140 |
|
|
|
| 2 |
C |
0.021 |
|
|
|
| 3 |
C |
-0.339 |
|
|
|
| 4 |
F |
-0.277 |
|
|
|
| 5 |
H |
0.124 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
H |
0.140 |
|
|
|
| 8 |
H |
0.154 |
|
|
|
| 9 |
H |
0.154 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.110 |
2.057 |
0.000 |
2.337 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.603 |
-0.301 |
0.000 |
| y |
-0.301 |
-26.403 |
0.000 |
| z |
0.000 |
0.000 |
-25.379 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.288 |
-0.301 |
0.000 |
| y |
-0.301 |
-3.412 |
0.000 |
| z |
0.000 |
0.000 |
-1.876 |
|
| Polar |
|---|
| 3z2-r2 | -3.752 |
|---|
| x2-y2 | 5.800 |
|---|
| xy | -0.301 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.114 |
0.148 |
0.000 |
| y |
0.148 |
4.908 |
0.000 |
| z |
0.000 |
0.000 |
3.341 |
<r2> (average value of r
2) Å
2
| <r2> |
69.416 |
| (<r2>)1/2 |
8.332 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -44.078166 |
| Energy at 298.15K | |
| HF Energy | -44.078166 |
| Nuclear repulsion energy | 62.686937 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3209 |
3127 |
28.92 |
68.56 |
0.57 |
0.73 |
| 2 |
A |
3141 |
3061 |
13.50 |
108.81 |
0.24 |
0.38 |
| 3 |
A |
3104 |
3025 |
31.95 |
117.25 |
0.32 |
0.48 |
| 4 |
A |
3100 |
3021 |
7.26 |
51.83 |
0.53 |
0.69 |
| 5 |
A |
3027 |
2950 |
43.70 |
132.96 |
0.09 |
0.16 |
| 6 |
A |
1679 |
1636 |
0.47 |
26.43 |
0.12 |
0.22 |
| 7 |
A |
1513 |
1475 |
4.26 |
8.68 |
0.66 |
0.80 |
| 8 |
A |
1467 |
1430 |
16.12 |
17.17 |
0.50 |
0.67 |
| 9 |
A |
1384 |
1349 |
13.07 |
6.38 |
0.66 |
0.80 |
| 10 |
A |
1318 |
1284 |
0.06 |
18.48 |
0.39 |
0.56 |
| 11 |
A |
1235 |
1203 |
4.59 |
17.98 |
0.71 |
0.83 |
| 12 |
A |
1173 |
1143 |
1.07 |
3.64 |
0.67 |
0.80 |
| 13 |
A |
1034 |
1007 |
21.89 |
1.75 |
0.73 |
0.85 |
| 14 |
A |
988 |
963 |
54.40 |
1.25 |
0.75 |
0.86 |
| 15 |
A |
979 |
954 |
11.00 |
1.79 |
0.29 |
0.45 |
| 16 |
A |
963 |
938 |
84.65 |
4.54 |
0.62 |
0.77 |
| 17 |
A |
910 |
887 |
45.22 |
11.62 |
0.25 |
0.40 |
| 18 |
A |
655 |
638 |
8.08 |
5.40 |
0.53 |
0.69 |
| 19 |
A |
423 |
412 |
2.31 |
4.94 |
0.45 |
0.62 |
| 20 |
A |
323 |
315 |
10.63 |
4.44 |
0.74 |
0.85 |
| 21 |
A |
106 |
104 |
1.65 |
6.42 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 15865.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 15460.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.597 |
0.463 |
0.336 |
| C2 |
0.661 |
-0.368 |
0.286 |
| C3 |
1.825 |
0.078 |
-0.241 |
| F4 |
-1.679 |
-0.249 |
-0.363 |
| H5 |
2.723 |
-0.536 |
-0.239 |
| H6 |
1.910 |
1.069 |
-0.687 |
| H7 |
0.589 |
-1.367 |
0.717 |
| H8 |
-0.963 |
0.606 |
1.360 |
| H9 |
-0.478 |
1.429 |
-0.167 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5085 | 2.5193 | 1.4715 | 3.5146 | 2.7743 | 2.2137 | 1.0973 | 1.0957 |
C2 | 1.5085 | | 1.3529 | 2.4317 | 2.1344 | 2.1377 | 1.0905 | 2.1769 | 2.1755 | C3 | 2.5193 | 1.3529 | | 3.5213 | 1.0879 | 1.0903 | 2.1288 | 3.2579 | 2.6715 | F4 | 1.4715 | 2.4317 | 3.5213 | | 4.4135 | 3.8370 | 2.7497 | 2.0526 | 2.0725 | H5 | 3.5146 | 2.1344 | 1.0879 | 4.4135 | | 1.8544 | 2.4816 | 4.1770 | 3.7573 | H6 | 2.7743 | 2.1377 | 1.0903 | 3.8370 | 1.8544 | | 3.1062 | 3.5574 | 2.4704 | H7 | 2.2137 | 1.0905 | 2.1288 | 2.7497 | 2.4816 | 3.1062 | | 2.5911 | 3.1207 | H8 | 1.0973 | 2.1769 | 3.2579 | 2.0526 | 4.1770 | 3.5574 | 2.5911 | | 1.8016 | H9 | 1.0957 | 2.1755 | 2.6715 | 2.0725 | 3.7573 | 2.4704 | 3.1207 | 1.8016 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.298 |
|
C1 |
C2 |
H7 |
115.868 |
| C2 |
C1 |
F4 |
109.366 |
|
C2 |
C1 |
H8 |
112.350 |
| C2 |
C1 |
H9 |
112.340 |
|
C2 |
C3 |
H5 |
121.582 |
| C2 |
C3 |
H6 |
121.705 |
|
C3 |
C2 |
H7 |
120.830 |
| F4 |
C1 |
H8 |
105.145 |
|
F4 |
C1 |
H9 |
106.753 |
| H5 |
C3 |
H6 |
116.713 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.177 |
|
|
|
| 2 |
C |
0.123 |
|
|
|
| 3 |
C |
-0.391 |
|
|
|
| 4 |
F |
-0.286 |
|
|
|
| 5 |
H |
0.134 |
|
|
|
| 6 |
H |
0.145 |
|
|
|
| 7 |
H |
0.159 |
|
|
|
| 8 |
H |
0.144 |
|
|
|
| 9 |
H |
0.150 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.066 |
1.046 |
1.148 |
2.584 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.716 |
-1.673 |
-2.490 |
| y |
-1.673 |
-22.521 |
-1.216 |
| z |
-2.490 |
-1.216 |
-24.825 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.043 |
-1.673 |
-2.490 |
| y |
-1.673 |
3.250 |
-1.216 |
| z |
-2.490 |
-1.216 |
-0.207 |
|
| Polar |
|---|
| 3z2-r2 | -0.413 |
|---|
| x2-y2 | -4.196 |
|---|
| xy | -1.673 |
|---|
| xz | -2.490 |
|---|
| yz | -1.216 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.989 |
0.308 |
-0.927 |
| y |
0.308 |
4.670 |
-0.540 |
| z |
-0.927 |
-0.540 |
3.920 |
<r2> (average value of r
2) Å
2
| <r2> |
76.635 |
| (<r2>)1/2 |
8.754 |
