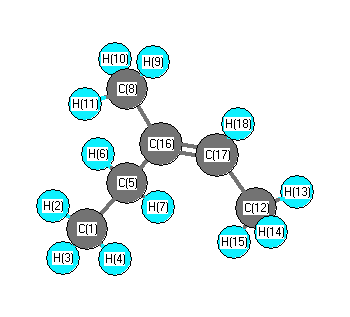
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3098 |
3019 |
99.56 |
|
|
|
| 2 |
A |
3081 |
3003 |
34.67 |
|
|
|
| 3 |
A |
3076 |
2998 |
49.67 |
|
|
|
| 4 |
A |
3074 |
2995 |
13.91 |
|
|
|
| 5 |
A |
3068 |
2990 |
70.92 |
|
|
|
| 6 |
A |
3039 |
2961 |
12.80 |
|
|
|
| 7 |
A |
3030 |
2952 |
26.73 |
|
|
|
| 8 |
A |
3029 |
2951 |
65.15 |
|
|
|
| 9 |
A |
2986 |
2910 |
36.80 |
|
|
|
| 10 |
A |
2980 |
2904 |
53.29 |
|
|
|
| 11 |
A |
2972 |
2896 |
62.46 |
|
|
|
| 12 |
A |
2967 |
2891 |
49.13 |
|
|
|
| 13 |
A |
1705 |
1661 |
0.86 |
|
|
|
| 14 |
A |
1536 |
1497 |
14.39 |
|
|
|
| 15 |
A |
1523 |
1484 |
1.16 |
|
|
|
| 16 |
A |
1520 |
1481 |
6.79 |
|
|
|
| 17 |
A |
1519 |
1480 |
10.18 |
|
|
|
| 18 |
A |
1511 |
1473 |
8.30 |
|
|
|
| 19 |
A |
1507 |
1469 |
4.32 |
|
|
|
| 20 |
A |
1502 |
1464 |
5.59 |
|
|
|
| 21 |
A |
1436 |
1399 |
3.57 |
|
|
|
| 22 |
A |
1433 |
1397 |
5.68 |
|
|
|
| 23 |
A |
1426 |
1389 |
6.88 |
|
|
|
| 24 |
A |
1389 |
1353 |
1.51 |
|
|
|
| 25 |
A |
1343 |
1308 |
3.39 |
|
|
|
| 26 |
A |
1295 |
1262 |
1.62 |
|
|
|
| 27 |
A |
1232 |
1200 |
1.99 |
|
|
|
| 28 |
A |
1141 |
1112 |
3.79 |
|
|
|
| 29 |
A |
1105 |
1077 |
3.67 |
|
|
|
| 30 |
A |
1076 |
1048 |
0.92 |
|
|
|
| 31 |
A |
1074 |
1047 |
3.50 |
|
|
|
| 32 |
A |
1057 |
1030 |
6.74 |
|
|
|
| 33 |
A |
1022 |
996 |
5.60 |
|
|
|
| 34 |
A |
997 |
972 |
0.61 |
|
|
|
| 35 |
A |
922 |
899 |
6.12 |
|
|
|
| 36 |
A |
849 |
827 |
17.84 |
|
|
|
| 37 |
A |
800 |
780 |
2.09 |
|
|
|
| 38 |
A |
730 |
711 |
0.90 |
|
|
|
| 39 |
A |
559 |
545 |
4.23 |
|
|
|
| 40 |
A |
517 |
504 |
2.61 |
|
|
|
| 41 |
A |
388 |
378 |
0.41 |
|
|
|
| 42 |
A |
297 |
289 |
0.26 |
|
|
|
| 43 |
A |
264 |
258 |
1.54 |
|
|
|
| 44 |
A |
235 |
229 |
1.44 |
|
|
|
| 45 |
A |
211 |
205 |
0.12 |
|
|
|
| 46 |
A |
169 |
165 |
1.22 |
|
|
|
| 47 |
A |
116 |
113 |
0.33 |
|
|
|
| 48 |
A |
46 |
44 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35923.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 35007.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.484 |
|
|
|
| 2 |
H |
0.143 |
|
|
|
| 3 |
H |
0.146 |
|
|
|
| 4 |
H |
0.147 |
|
|
|
| 5 |
C |
-0.313 |
|
|
|
| 6 |
H |
0.127 |
|
|
|
| 7 |
H |
0.142 |
|
|
|
| 8 |
C |
-0.673 |
|
|
|
| 9 |
H |
0.161 |
|
|
|
| 10 |
H |
0.151 |
|
|
|
| 11 |
H |
0.151 |
|
|
|
| 12 |
C |
-0.567 |
|
|
|
| 13 |
H |
0.152 |
|
|
|
| 14 |
H |
0.153 |
|
|
|
| 15 |
H |
0.151 |
|
|
|
| 16 |
C |
0.507 |
|
|
|
| 17 |
C |
-0.229 |
|
|
|
| 18 |
H |
0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.153 |
-0.155 |
0.044 |
0.222 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.244 |
0.424 |
0.715 |
| y |
0.424 |
-39.802 |
-0.619 |
| z |
0.715 |
-0.619 |
-41.480 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.397 |
0.424 |
0.715 |
| y |
0.424 |
0.560 |
-0.619 |
| z |
0.715 |
-0.619 |
-1.957 |
|
| Polar |
|---|
| 3z2-r2 | -3.914 |
|---|
| x2-y2 | 0.558 |
|---|
| xy | 0.424 |
|---|
| xz | 0.715 |
|---|
| yz | -0.619 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.392 |
0.301 |
0.691 |
| y |
0.301 |
10.331 |
-0.159 |
| z |
0.691 |
-0.159 |
8.409 |
<r2> (average value of r
2) Å
2
| <r2> |
179.328 |
| (<r2>)1/2 |
13.391 |