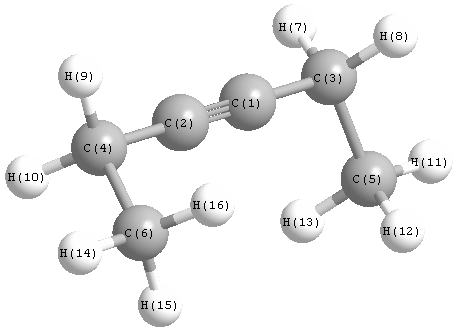
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3089 |
3011 |
0.00 |
|
|
|
| 2 |
A |
3080 |
3002 |
20.84 |
|
|
|
| 3 |
A |
3020 |
2943 |
0.00 |
|
|
|
| 4 |
A |
2993 |
2916 |
18.42 |
|
|
|
| 5 |
A |
2983 |
2907 |
77.54 |
|
|
|
| 6 |
A |
2299 |
2240 |
0.00 |
|
|
|
| 7 |
A |
1528 |
1489 |
2.90 |
|
|
|
| 8 |
A |
1515 |
1477 |
0.00 |
|
|
|
| 9 |
A |
1511 |
1472 |
5.99 |
|
|
|
| 10 |
A |
1428 |
1391 |
7.75 |
|
|
|
| 11 |
A |
1356 |
1322 |
0.08 |
|
|
|
| 12 |
A |
1284 |
1251 |
0.00 |
|
|
|
| 13 |
A |
1109 |
1080 |
0.00 |
|
|
|
| 14 |
A |
1097 |
1069 |
1.59 |
|
|
|
| 15 |
A |
990 |
965 |
0.28 |
|
|
|
| 16 |
A |
798 |
777 |
0.00 |
|
|
|
| 17 |
A |
679 |
662 |
0.23 |
|
|
|
| 18 |
A |
418 |
408 |
8.40 |
|
|
|
| 19 |
A |
406 |
395 |
0.00 |
|
|
|
| 20 |
A |
221 |
215 |
0.00 |
|
|
|
| 21 |
A |
89 |
87 |
2.21 |
|
|
|
| 22 |
A |
21i |
21i |
0.00 |
|
|
|
| 23 |
B |
3090 |
3011 |
108.07 |
|
|
|
| 24 |
B |
3079 |
3001 |
107.22 |
|
|
|
| 25 |
B |
3020 |
2943 |
34.22 |
|
|
|
| 26 |
B |
2991 |
2915 |
77.32 |
|
|
|
| 27 |
B |
2983 |
2907 |
20.82 |
|
|
|
| 28 |
B |
1527 |
1488 |
5.99 |
|
|
|
| 29 |
B |
1516 |
1477 |
16.32 |
|
|
|
| 30 |
B |
1510 |
1471 |
1.52 |
|
|
|
| 31 |
B |
1427 |
1391 |
0.05 |
|
|
|
| 32 |
B |
1351 |
1316 |
40.26 |
|
|
|
| 33 |
B |
1289 |
1256 |
0.13 |
|
|
|
| 34 |
B |
1162 |
1132 |
5.40 |
|
|
|
| 35 |
B |
1128 |
1099 |
0.52 |
|
|
|
| 36 |
B |
1076 |
1049 |
3.67 |
|
|
|
| 37 |
B |
918 |
895 |
0.41 |
|
|
|
| 38 |
B |
799 |
779 |
4.88 |
|
|
|
| 39 |
B |
556 |
542 |
1.14 |
|
|
|
| 40 |
B |
278 |
271 |
5.58 |
|
|
|
| 41 |
B |
217 |
212 |
0.15 |
|
|
|
| 42 |
B |
157 |
153 |
5.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30971.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30182.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.028 |
|
|
|
| 2 |
C |
0.028 |
|
|
|
| 3 |
C |
-0.314 |
|
|
|
| 4 |
C |
-0.314 |
|
|
|
| 5 |
C |
-0.485 |
|
|
|
| 6 |
C |
-0.485 |
|
|
|
| 7 |
H |
0.156 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
| 9 |
H |
0.156 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
| 11 |
H |
0.156 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.151 |
|
|
|
| 14 |
H |
0.156 |
|
|
|
| 15 |
H |
0.151 |
|
|
|
| 16 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.092 |
0.092 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.814 |
0.000 |
0.000 |
| y |
0.000 |
-32.547 |
0.000 |
| z |
0.000 |
0.000 |
-39.596 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.743 |
0.000 |
0.000 |
| y |
0.000 |
6.658 |
0.000 |
| z |
0.000 |
0.000 |
-3.915 |
|
| Polar |
|---|
| 3z2-r2 | -7.830 |
|---|
| x2-y2 | -6.267 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.667 |
-0.001 |
0.000 |
| y |
-0.001 |
14.505 |
0.000 |
| z |
0.000 |
0.000 |
8.422 |
<r2> (average value of r
2) Å
2
| <r2> |
227.348 |
| (<r2>)1/2 |
15.078 |