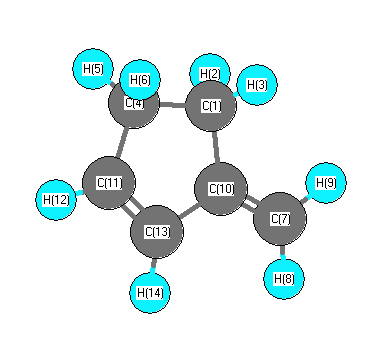
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3200 |
3118 |
30.24 |
|
|
|
| 2 |
A |
3187 |
3106 |
31.51 |
|
|
|
| 3 |
A |
3157 |
3076 |
5.68 |
|
|
|
| 4 |
A |
3103 |
3024 |
12.18 |
|
|
|
| 5 |
A |
3061 |
2983 |
45.64 |
|
|
|
| 6 |
A |
3016 |
2939 |
21.18 |
|
|
|
| 7 |
A |
3015 |
2939 |
41.61 |
|
|
|
| 8 |
A |
2983 |
2907 |
56.87 |
|
|
|
| 9 |
A |
1672 |
1629 |
34.37 |
|
|
|
| 10 |
A |
1612 |
1570 |
0.90 |
|
|
|
| 11 |
A |
1519 |
1480 |
6.48 |
|
|
|
| 12 |
A |
1496 |
1458 |
2.06 |
|
|
|
| 13 |
A |
1466 |
1428 |
0.82 |
|
|
|
| 14 |
A |
1367 |
1332 |
1.09 |
|
|
|
| 15 |
A |
1329 |
1295 |
2.49 |
|
|
|
| 16 |
A |
1291 |
1258 |
0.16 |
|
|
|
| 17 |
A |
1260 |
1228 |
3.47 |
|
|
|
| 18 |
A |
1240 |
1208 |
0.22 |
|
|
|
| 19 |
A |
1168 |
1138 |
1.44 |
|
|
|
| 20 |
A |
1127 |
1098 |
2.02 |
|
|
|
| 21 |
A |
1046 |
1019 |
6.64 |
|
|
|
| 22 |
A |
1016 |
990 |
2.80 |
|
|
|
| 23 |
A |
996 |
971 |
6.03 |
|
|
|
| 24 |
A |
957 |
933 |
2.33 |
|
|
|
| 25 |
A |
919 |
895 |
62.71 |
|
|
|
| 26 |
A |
908 |
885 |
9.82 |
|
|
|
| 27 |
A |
851 |
830 |
14.59 |
|
|
|
| 28 |
A |
841 |
820 |
0.63 |
|
|
|
| 29 |
A |
803 |
783 |
12.94 |
|
|
|
| 30 |
A |
773 |
754 |
0.94 |
|
|
|
| 31 |
A |
679 |
662 |
12.73 |
|
|
|
| 32 |
A |
622 |
606 |
0.67 |
|
|
|
| 33 |
A |
513 |
500 |
18.07 |
|
|
|
| 34 |
A |
358 |
348 |
0.51 |
|
|
|
| 35 |
A |
300 |
292 |
1.04 |
|
|
|
| 36 |
A |
63 |
62 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26456.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 25782.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.462 |
|
|
|
| 2 |
H |
0.157 |
|
|
|
| 3 |
H |
0.157 |
|
|
|
| 4 |
C |
-0.352 |
|
|
|
| 5 |
H |
0.148 |
|
|
|
| 6 |
H |
0.148 |
|
|
|
| 7 |
C |
-0.607 |
|
|
|
| 8 |
H |
0.128 |
|
|
|
| 9 |
H |
0.130 |
|
|
|
| 10 |
C |
0.475 |
|
|
|
| 11 |
C |
-0.155 |
|
|
|
| 12 |
H |
0.156 |
|
|
|
| 13 |
C |
-0.096 |
|
|
|
| 14 |
H |
0.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.860 |
-0.276 |
0.000 |
0.903 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.989 |
0.127 |
0.000 |
| y |
0.127 |
-34.753 |
0.000 |
| z |
0.000 |
0.000 |
-39.782 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.279 |
0.127 |
0.000 |
| y |
0.127 |
3.132 |
0.000 |
| z |
0.000 |
0.000 |
-4.411 |
|
| Polar |
|---|
| 3z2-r2 | -8.822 |
|---|
| x2-y2 | -1.236 |
|---|
| xy | 0.127 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
15.388 |
-0.074 |
0.001 |
| y |
-0.074 |
9.439 |
0.002 |
| z |
0.001 |
0.002 |
6.003 |
<r2> (average value of r
2) Å
2
| <r2> |
129.001 |
| (<r2>)1/2 |
11.358 |