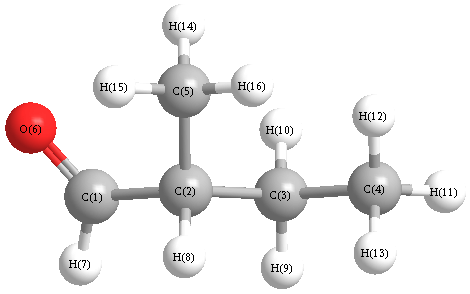
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3097 |
3018 |
31.45 |
|
|
|
| 2 |
A |
3088 |
3010 |
43.23 |
|
|
|
| 3 |
A |
3079 |
3000 |
66.19 |
|
|
|
| 4 |
A |
3068 |
2990 |
60.40 |
|
|
|
| 5 |
A |
3022 |
2945 |
25.94 |
|
|
|
| 6 |
A |
3003 |
2926 |
19.25 |
|
|
|
| 7 |
A |
2986 |
2909 |
29.89 |
|
|
|
| 8 |
A |
2974 |
2899 |
58.77 |
|
|
|
| 9 |
A |
2940 |
2865 |
35.31 |
|
|
|
| 10 |
A |
2893 |
2819 |
102.56 |
|
|
|
| 11 |
A |
1668 |
1625 |
151.52 |
|
|
|
| 12 |
A |
1531 |
1492 |
5.63 |
|
|
|
| 13 |
A |
1525 |
1486 |
12.97 |
|
|
|
| 14 |
A |
1519 |
1480 |
5.42 |
|
|
|
| 15 |
A |
1514 |
1475 |
9.12 |
|
|
|
| 16 |
A |
1509 |
1470 |
0.51 |
|
|
|
| 17 |
A |
1439 |
1402 |
9.87 |
|
|
|
| 18 |
A |
1432 |
1396 |
9.34 |
|
|
|
| 19 |
A |
1409 |
1373 |
1.39 |
|
|
|
| 20 |
A |
1377 |
1342 |
5.17 |
|
|
|
| 21 |
A |
1347 |
1312 |
1.54 |
|
|
|
| 22 |
A |
1312 |
1279 |
3.41 |
|
|
|
| 23 |
A |
1280 |
1247 |
0.61 |
|
|
|
| 24 |
A |
1198 |
1167 |
1.23 |
|
|
|
| 25 |
A |
1159 |
1130 |
2.97 |
|
|
|
| 26 |
A |
1119 |
1090 |
4.09 |
|
|
|
| 27 |
A |
1043 |
1017 |
0.12 |
|
|
|
| 28 |
A |
1018 |
992 |
11.11 |
|
|
|
| 29 |
A |
985 |
960 |
12.21 |
|
|
|
| 30 |
A |
930 |
906 |
2.26 |
|
|
|
| 31 |
A |
899 |
876 |
19.52 |
|
|
|
| 32 |
A |
794 |
774 |
5.30 |
|
|
|
| 33 |
A |
768 |
749 |
3.45 |
|
|
|
| 34 |
A |
648 |
632 |
9.23 |
|
|
|
| 35 |
A |
400 |
390 |
1.18 |
|
|
|
| 36 |
A |
381 |
372 |
0.85 |
|
|
|
| 37 |
A |
278 |
271 |
6.28 |
|
|
|
| 38 |
A |
259 |
253 |
0.75 |
|
|
|
| 39 |
A |
218 |
212 |
2.51 |
|
|
|
| 40 |
A |
180 |
175 |
0.71 |
|
|
|
| 41 |
A |
84 |
82 |
3.37 |
|
|
|
| 42 |
A |
70 |
68 |
4.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30721.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29938.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.069 |
|
|
|
| 2 |
C |
0.060 |
|
|
|
| 3 |
C |
-0.238 |
|
|
|
| 4 |
C |
-0.491 |
|
|
|
| 5 |
C |
-0.539 |
|
|
|
| 6 |
O |
-0.146 |
|
|
|
| 7 |
H |
0.088 |
|
|
|
| 8 |
H |
0.139 |
|
|
|
| 9 |
H |
0.137 |
|
|
|
| 10 |
H |
0.141 |
|
|
|
| 11 |
H |
0.153 |
|
|
|
| 12 |
H |
0.154 |
|
|
|
| 13 |
H |
0.144 |
|
|
|
| 14 |
H |
0.154 |
|
|
|
| 15 |
H |
0.166 |
|
|
|
| 16 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.065 |
-0.561 |
0.944 |
3.256 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.224 |
0.070 |
-2.191 |
| y |
0.070 |
-37.399 |
-0.381 |
| z |
-2.191 |
-0.381 |
-37.958 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.546 |
0.070 |
-2.191 |
| y |
0.070 |
5.692 |
-0.381 |
| z |
-2.191 |
-0.381 |
4.854 |
|
| Polar |
|---|
| 3z2-r2 | 9.707 |
|---|
| x2-y2 | -10.825 |
|---|
| xy | 0.070 |
|---|
| xz | -2.191 |
|---|
| yz | -0.381 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.446 |
-0.350 |
0.336 |
| y |
-0.350 |
8.911 |
-0.289 |
| z |
0.336 |
-0.289 |
7.658 |
<r2> (average value of r
2) Å
2
| <r2> |
171.844 |
| (<r2>)1/2 |
13.109 |