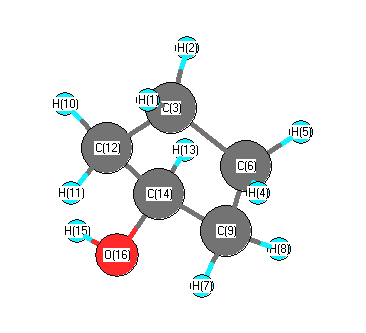
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3653 |
3560 |
4.86 |
|
|
|
| 2 |
A |
3084 |
3005 |
122.44 |
|
|
|
| 3 |
A |
3072 |
2993 |
56.75 |
|
|
|
| 4 |
A |
3058 |
2980 |
8.91 |
|
|
|
| 5 |
A |
3041 |
2963 |
19.82 |
|
|
|
| 6 |
A |
3020 |
2943 |
38.58 |
|
|
|
| 7 |
A |
3011 |
2934 |
56.38 |
|
|
|
| 8 |
A |
3000 |
2923 |
86.78 |
|
|
|
| 9 |
A |
2995 |
2919 |
13.82 |
|
|
|
| 10 |
A |
2958 |
2882 |
52.66 |
|
|
|
| 11 |
A |
1540 |
1501 |
3.31 |
|
|
|
| 12 |
A |
1522 |
1483 |
6.63 |
|
|
|
| 13 |
A |
1519 |
1481 |
5.58 |
|
|
|
| 14 |
A |
1511 |
1473 |
2.93 |
|
|
|
| 15 |
A |
1413 |
1377 |
3.34 |
|
|
|
| 16 |
A |
1363 |
1328 |
16.22 |
|
|
|
| 17 |
A |
1340 |
1306 |
0.89 |
|
|
|
| 18 |
A |
1332 |
1298 |
2.89 |
|
|
|
| 19 |
A |
1318 |
1284 |
2.49 |
|
|
|
| 20 |
A |
1298 |
1265 |
1.03 |
|
|
|
| 21 |
A |
1283 |
1250 |
5.92 |
|
|
|
| 22 |
A |
1245 |
1214 |
10.20 |
|
|
|
| 23 |
A |
1216 |
1185 |
17.04 |
|
|
|
| 24 |
A |
1203 |
1172 |
1.37 |
|
|
|
| 25 |
A |
1178 |
1148 |
9.52 |
|
|
|
| 26 |
A |
1080 |
1053 |
1.51 |
|
|
|
| 27 |
A |
1063 |
1036 |
38.07 |
|
|
|
| 28 |
A |
1030 |
1003 |
74.20 |
|
|
|
| 29 |
A |
1000 |
974 |
0.73 |
|
|
|
| 30 |
A |
979 |
954 |
4.99 |
|
|
|
| 31 |
A |
945 |
921 |
11.61 |
|
|
|
| 32 |
A |
892 |
869 |
1.29 |
|
|
|
| 33 |
A |
862 |
840 |
0.66 |
|
|
|
| 34 |
A |
820 |
799 |
0.01 |
|
|
|
| 35 |
A |
777 |
757 |
2.81 |
|
|
|
| 36 |
A |
624 |
608 |
0.66 |
|
|
|
| 37 |
A |
532 |
518 |
4.37 |
|
|
|
| 38 |
A |
456 |
445 |
10.06 |
|
|
|
| 39 |
A |
342 |
334 |
20.69 |
|
|
|
| 40 |
A |
272 |
266 |
149.81 |
|
|
|
| 41 |
A |
166 |
162 |
1.60 |
|
|
|
| 42 |
A |
35 |
34 |
0.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31523.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30719.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.134 |
|
|
|
| 2 |
H |
0.130 |
|
|
|
| 3 |
C |
-0.263 |
|
|
|
| 4 |
H |
0.135 |
|
|
|
| 5 |
H |
0.130 |
|
|
|
| 6 |
C |
-0.269 |
|
|
|
| 7 |
H |
0.144 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
C |
-0.273 |
|
|
|
| 10 |
H |
0.123 |
|
|
|
| 11 |
H |
0.144 |
|
|
|
| 12 |
C |
-0.310 |
|
|
|
| 13 |
H |
0.110 |
|
|
|
| 14 |
C |
0.057 |
|
|
|
| 15 |
H |
0.329 |
|
|
|
| 16 |
O |
-0.467 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.848 |
1.313 |
1.159 |
1.946 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.589 |
4.201 |
2.975 |
| y |
4.201 |
-38.266 |
0.197 |
| z |
2.975 |
0.197 |
-38.054 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.429 |
4.201 |
2.975 |
| y |
4.201 |
0.556 |
0.197 |
| z |
2.975 |
0.197 |
0.873 |
|
| Polar |
|---|
| 3z2-r2 | 1.746 |
|---|
| x2-y2 | -1.323 |
|---|
| xy | 4.201 |
|---|
| xz | 2.975 |
|---|
| yz | 0.197 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.550 |
0.235 |
-0.004 |
| y |
0.235 |
8.898 |
0.084 |
| z |
-0.004 |
0.084 |
7.963 |
<r2> (average value of r
2) Å
2
| <r2> |
142.881 |
| (<r2>)1/2 |
11.953 |