Jump to
S1C2
S1C3
Energy calculated at wB97X-D/CEP-121G*
| | hartrees |
|---|
| Energy at 0K | -34.353740 |
| Energy at 298.15K | |
| HF Energy | -34.353740 |
| Nuclear repulsion energy | 25.127774 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/CEP-121G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
4004 |
4004 |
42.98 |
|
|
|
| 2 |
A1 |
730 |
730 |
3.07 |
|
|
|
| 3 |
A1 |
610 |
610 |
177.18 |
|
|
|
| 4 |
A1 |
294 |
294 |
10.35 |
|
|
|
| 5 |
A2 |
291i |
291i |
0.00 |
|
|
|
| 6 |
B1 |
361 |
361 |
86.76 |
|
|
|
| 7 |
B2 |
4003 |
4003 |
103.96 |
|
|
|
| 8 |
B2 |
1513 |
1513 |
354.08 |
|
|
|
| 9 |
B2 |
483 |
483 |
353.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5853.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5853.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/CEP-121G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
0.018 |
| O2 |
0.000 |
1.425 |
0.077 |
| O3 |
0.000 |
-1.425 |
0.077 |
| H4 |
0.000 |
2.046 |
-0.651 |
| H5 |
0.000 |
-2.046 |
-0.651 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4265 | 1.4265 | 2.1527 | 2.1527 |
O2 | 1.4265 | | 2.8506 | 0.9565 | 3.5470 | O3 | 1.4265 | 2.8506 | | 3.5470 | 0.9565 | H4 | 2.1527 | 0.9565 | 3.5470 | | 4.0925 | H5 | 2.1527 | 3.5470 | 0.9565 | 4.0925 | |
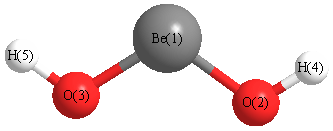 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
128.116 |
|
Be1 |
O3 |
H5 |
128.116 |
| O2 |
Be1 |
O3 |
175.274 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/CEP-121G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.470 |
|
|
|
| 2 |
O |
-0.664 |
|
|
|
| 3 |
O |
-0.664 |
|
|
|
| 4 |
H |
0.429 |
|
|
|
| 5 |
H |
0.429 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.803 |
2.803 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.813 |
0.000 |
0.000 |
| y |
0.000 |
-14.994 |
0.000 |
| z |
0.000 |
0.000 |
-14.101 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.266 |
0.000 |
0.000 |
| y |
0.000 |
0.463 |
0.000 |
| z |
0.000 |
0.000 |
1.803 |
|
| Polar |
|---|
| 3z2-r2 | 3.606 |
|---|
| x2-y2 | -1.820 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.222 |
0.000 |
0.000 |
| y |
0.000 |
3.597 |
0.000 |
| z |
0.000 |
0.000 |
2.400 |
<r2> (average value of r
2) Å
2
| <r2> |
43.228 |
| (<r2>)1/2 |
6.575 |
Jump to
S1C1
S1C3
Energy calculated at wB97X-D/CEP-121G*
| | hartrees |
|---|
| Energy at 0K | -34.354698 |
| Energy at 298.15K | -34.356237 |
| HF Energy | -34.354698 |
| Nuclear repulsion energy | 25.125828 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/CEP-121G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3988 |
3988 |
21.36 |
|
|
|
| 2 |
A |
727 |
727 |
0.61 |
|
|
|
| 3 |
A |
607 |
607 |
90.98 |
|
|
|
| 4 |
A |
321 |
321 |
57.87 |
|
|
|
| 5 |
A |
224 |
224 |
147.96 |
|
|
|
| 6 |
B |
3987 |
3987 |
110.89 |
|
|
|
| 7 |
B |
1512 |
1512 |
351.02 |
|
|
|
| 8 |
B |
600 |
600 |
392.64 |
|
|
|
| 9 |
B |
311 |
311 |
100.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6139.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 6139.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/CEP-121G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
-0.022 |
| O2 |
0.000 |
1.428 |
-0.053 |
| O3 |
0.000 |
-1.428 |
-0.053 |
| H4 |
0.563 |
1.998 |
0.470 |
| H5 |
-0.563 |
-1.998 |
0.470 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4279 | 1.4279 | 2.1337 | 2.1337 |
O2 | 1.4279 | | 2.8551 | 0.9576 | 3.5112 | O3 | 1.4279 | 2.8551 | | 3.5112 | 0.9576 | H4 | 2.1337 | 0.9576 | 3.5112 | | 4.1526 | H5 | 2.1337 | 3.5112 | 0.9576 | 4.1526 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
125.721 |
|
Be1 |
O3 |
H5 |
125.721 |
| O2 |
Be1 |
O3 |
177.460 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/CEP-121G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.464 |
|
|
|
| 2 |
O |
-0.661 |
|
|
|
| 3 |
O |
-0.661 |
|
|
|
| 4 |
H |
0.429 |
|
|
|
| 5 |
H |
0.429 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.972 |
1.972 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.923 |
4.926 |
0.000 |
| y |
4.926 |
-15.899 |
0.000 |
| z |
0.000 |
0.000 |
-15.419 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.735 |
4.926 |
0.000 |
| y |
4.926 |
-0.727 |
0.000 |
| z |
0.000 |
0.000 |
-0.008 |
|
| Polar |
|---|
| 3z2-r2 | -0.016 |
|---|
| x2-y2 | 0.975 |
|---|
| xy | 4.926 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.334 |
0.132 |
0.000 |
| y |
0.132 |
3.639 |
0.000 |
| z |
0.000 |
0.000 |
2.341 |
<r2> (average value of r
2) Å
2
| <r2> |
43.181 |
| (<r2>)1/2 |
6.571 |
Jump to
S1C1
S1C2
Energy calculated at wB97X-D/CEP-121G*
| | hartrees |
|---|
| Energy at 0K | -34.346053 |
| Energy at 298.15K | |
| HF Energy | -34.346053 |
| Nuclear repulsion energy | 25.294863 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/CEP-121G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
4160 |
4160 |
0.00 |
|
|
|
| 2 |
Σg |
766 |
766 |
0.00 |
|
|
|
| 3 |
Σu |
4158 |
4158 |
380.46 |
|
|
|
| 4 |
Σu |
1604 |
1604 |
483.98 |
|
|
|
| 5 |
Πg |
453i |
453i |
0.00 |
|
|
|
| 5 |
Πg |
453i |
453i |
0.00 |
|
|
|
| 6 |
Πu |
344 |
344 |
15.73 |
|
|
|
| 6 |
Πu |
344 |
344 |
15.73 |
|
|
|
| 7 |
Πu |
472i |
472i |
464.19 |
|
|
|
| 7 |
Πu |
472i |
472i |
464.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4762.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4762.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/CEP-121G*
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
0.000 |
| O2 |
0.000 |
0.000 |
1.400 |
| O3 |
0.000 |
0.000 |
-1.400 |
| H4 |
0.000 |
0.000 |
2.347 |
| H5 |
0.000 |
0.000 |
-2.347 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.3998 | 1.3998 | 2.3465 | 2.3465 |
O2 | 1.3998 | | 2.7997 | 0.9467 | 3.7463 | O3 | 1.3998 | 2.7997 | | 3.7463 | 0.9467 | H4 | 2.3465 | 0.9467 | 3.7463 | | 4.6930 | H5 | 2.3465 | 3.7463 | 0.9467 | 4.6930 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
180.000 |
|
Be1 |
O3 |
H5 |
180.000 |
| O2 |
Be1 |
O3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/CEP-121G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.566 |
|
|
|
| 2 |
O |
-0.716 |
|
|
|
| 3 |
O |
-0.716 |
|
|
|
| 4 |
H |
0.433 |
|
|
|
| 5 |
H |
0.433 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.665 |
0.000 |
0.000 |
| y |
0.000 |
-16.665 |
0.000 |
| z |
0.000 |
0.000 |
-7.167 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.749 |
0.000 |
0.000 |
| y |
0.000 |
-4.749 |
0.000 |
| z |
0.000 |
0.000 |
9.499 |
|
| Polar |
|---|
| 3z2-r2 | 18.998 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.055 |
0.000 |
0.000 |
| y |
0.000 |
2.055 |
0.000 |
| z |
0.000 |
0.000 |
3.757 |
<r2> (average value of r
2) Å
2
| <r2> |
42.958 |
| (<r2>)1/2 |
6.554 |
