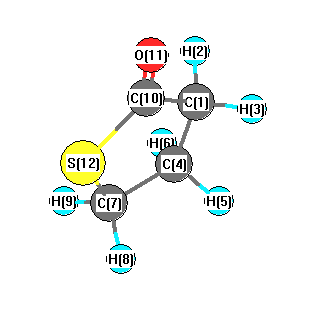
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3186 |
3062 |
19.26 |
|
|
|
| 2 |
A |
3143 |
3021 |
20.64 |
|
|
|
| 3 |
A |
3131 |
3010 |
20.89 |
|
|
|
| 4 |
A |
3105 |
2984 |
24.10 |
|
|
|
| 5 |
A |
3073 |
2954 |
28.47 |
|
|
|
| 6 |
A |
3054 |
2936 |
11.72 |
|
|
|
| 7 |
A |
1707 |
1641 |
369.09 |
|
|
|
| 8 |
A |
1517 |
1458 |
5.04 |
|
|
|
| 9 |
A |
1506 |
1447 |
8.87 |
|
|
|
| 10 |
A |
1484 |
1427 |
14.85 |
|
|
|
| 11 |
A |
1378 |
1325 |
1.41 |
|
|
|
| 12 |
A |
1337 |
1285 |
7.25 |
|
|
|
| 13 |
A |
1321 |
1269 |
7.68 |
|
|
|
| 14 |
A |
1258 |
1209 |
1.55 |
|
|
|
| 15 |
A |
1201 |
1154 |
3.24 |
|
|
|
| 16 |
A |
1135 |
1091 |
5.86 |
|
|
|
| 17 |
A |
1085 |
1043 |
16.02 |
|
|
|
| 18 |
A |
1050 |
1009 |
7.52 |
|
|
|
| 19 |
A |
1033 |
993 |
81.93 |
|
|
|
| 20 |
A |
921 |
886 |
23.57 |
|
|
|
| 21 |
A |
883 |
848 |
8.31 |
|
|
|
| 22 |
A |
847 |
814 |
18.92 |
|
|
|
| 23 |
A |
652 |
626 |
3.77 |
|
|
|
| 24 |
A |
600 |
576 |
20.19 |
|
|
|
| 25 |
A |
574 |
552 |
35.17 |
|
|
|
| 26 |
A |
475 |
457 |
2.20 |
|
|
|
| 27 |
A |
463 |
445 |
0.99 |
|
|
|
| 28 |
A |
390 |
375 |
3.95 |
|
|
|
| 29 |
A |
224 |
215 |
1.88 |
|
|
|
| 30 |
A |
124 |
119 |
1.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20926.3 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 20114.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.381 |
|
|
|
| 2 |
H |
0.248 |
|
|
|
| 3 |
H |
0.227 |
|
|
|
| 4 |
C |
-0.349 |
|
|
|
| 5 |
H |
0.218 |
|
|
|
| 6 |
H |
0.219 |
|
|
|
| 7 |
C |
-0.637 |
|
|
|
| 8 |
H |
0.254 |
|
|
|
| 9 |
H |
0.257 |
|
|
|
| 10 |
C |
-0.049 |
|
|
|
| 11 |
O |
-0.191 |
|
|
|
| 12 |
S |
0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.239 |
-2.265 |
0.455 |
4.828 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.591 |
-1.391 |
0.068 |
| y |
-1.391 |
-43.779 |
0.067 |
| z |
0.068 |
0.067 |
-41.653 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.875 |
-1.391 |
0.068 |
| y |
-1.391 |
0.843 |
0.067 |
| z |
0.068 |
0.067 |
4.033 |
|
| Polar |
|---|
| 3z2-r2 | 8.065 |
|---|
| x2-y2 | -3.812 |
|---|
| xy | -1.391 |
|---|
| xz | 0.068 |
|---|
| yz | 0.067 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.714 |
-0.531 |
-0.234 |
| y |
-0.531 |
9.289 |
-0.334 |
| z |
-0.234 |
-0.334 |
5.431 |
<r2> (average value of r
2) Å
2
| <r2> |
163.253 |
| (<r2>)1/2 |
12.777 |