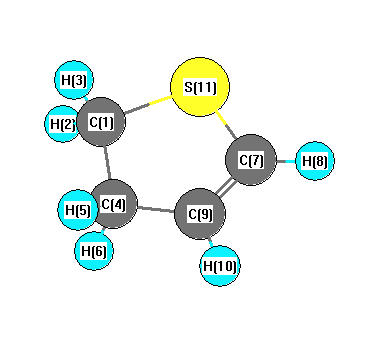
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3270 |
3143 |
10.44 |
|
|
|
| 2 |
A |
3215 |
3090 |
3.85 |
|
|
|
| 3 |
A |
3185 |
3061 |
21.72 |
|
|
|
| 4 |
A |
3109 |
2988 |
49.86 |
|
|
|
| 5 |
A |
3089 |
2969 |
13.54 |
|
|
|
| 6 |
A |
3031 |
2913 |
45.19 |
|
|
|
| 7 |
A |
1646 |
1582 |
14.48 |
|
|
|
| 8 |
A |
1513 |
1454 |
4.26 |
|
|
|
| 9 |
A |
1498 |
1440 |
4.30 |
|
|
|
| 10 |
A |
1348 |
1296 |
5.47 |
|
|
|
| 11 |
A |
1328 |
1277 |
4.22 |
|
|
|
| 12 |
A |
1291 |
1241 |
21.45 |
|
|
|
| 13 |
A |
1221 |
1174 |
2.18 |
|
|
|
| 14 |
A |
1155 |
1111 |
0.52 |
|
|
|
| 15 |
A |
1136 |
1092 |
0.67 |
|
|
|
| 16 |
A |
1035 |
995 |
7.77 |
|
|
|
| 17 |
A |
1016 |
977 |
4.17 |
|
|
|
| 18 |
A |
940 |
903 |
1.54 |
|
|
|
| 19 |
A |
917 |
882 |
6.46 |
|
|
|
| 20 |
A |
856 |
823 |
2.35 |
|
|
|
| 21 |
A |
774 |
744 |
21.22 |
|
|
|
| 22 |
A |
698 |
671 |
86.56 |
|
|
|
| 23 |
A |
669 |
643 |
18.79 |
|
|
|
| 24 |
A |
619 |
595 |
2.90 |
|
|
|
| 25 |
A |
493 |
474 |
0.58 |
|
|
|
| 26 |
A |
403 |
388 |
2.23 |
|
|
|
| 27 |
A |
131 |
126 |
0.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19792.5 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 19024.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.628 |
|
|
|
| 2 |
H |
0.256 |
|
|
|
| 3 |
H |
0.245 |
|
|
|
| 4 |
C |
-0.347 |
|
|
|
| 5 |
H |
0.207 |
|
|
|
| 6 |
H |
0.211 |
|
|
|
| 7 |
C |
-0.436 |
|
|
|
| 8 |
H |
0.243 |
|
|
|
| 9 |
C |
-0.130 |
|
|
|
| 10 |
H |
0.231 |
|
|
|
| 11 |
S |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.725 |
-1.952 |
0.228 |
2.095 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.295 |
0.320 |
-0.123 |
| y |
0.320 |
-39.424 |
0.196 |
| z |
-0.123 |
0.196 |
-39.387 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.111 |
0.320 |
-0.123 |
| y |
0.320 |
-3.582 |
0.196 |
| z |
-0.123 |
0.196 |
-3.528 |
|
| Polar |
|---|
| 3z2-r2 | -7.056 |
|---|
| x2-y2 | 7.129 |
|---|
| xy | 0.320 |
|---|
| xz | -0.123 |
|---|
| yz | 0.196 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.830 |
-0.429 |
-0.019 |
| y |
-0.429 |
10.004 |
-0.413 |
| z |
-0.019 |
-0.413 |
4.481 |
<r2> (average value of r
2) Å
2
| <r2> |
110.177 |
| (<r2>)1/2 |
10.497 |