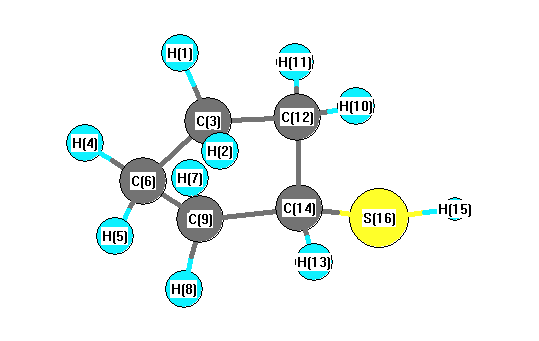
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3146 |
3024 |
125.85 |
|
|
|
| 2 |
A |
3129 |
3008 |
21.43 |
|
|
|
| 3 |
A |
3124 |
3002 |
5.21 |
|
|
|
| 4 |
A |
3103 |
2983 |
18.21 |
|
|
|
| 5 |
A |
3092 |
2972 |
9.21 |
|
|
|
| 6 |
A |
3077 |
2958 |
46.35 |
|
|
|
| 7 |
A |
3064 |
2945 |
63.79 |
|
|
|
| 8 |
A |
3061 |
2942 |
43.07 |
|
|
|
| 9 |
A |
3056 |
2937 |
3.35 |
|
|
|
| 10 |
A |
2498 |
2401 |
39.11 |
|
|
|
| 11 |
A |
1539 |
1479 |
2.27 |
|
|
|
| 12 |
A |
1523 |
1464 |
11.25 |
|
|
|
| 13 |
A |
1521 |
1462 |
8.80 |
|
|
|
| 14 |
A |
1513 |
1454 |
1.88 |
|
|
|
| 15 |
A |
1367 |
1314 |
10.19 |
|
|
|
| 16 |
A |
1356 |
1303 |
1.02 |
|
|
|
| 17 |
A |
1341 |
1289 |
2.08 |
|
|
|
| 18 |
A |
1330 |
1279 |
0.82 |
|
|
|
| 19 |
A |
1317 |
1265 |
0.88 |
|
|
|
| 20 |
A |
1288 |
1238 |
10.48 |
|
|
|
| 21 |
A |
1265 |
1216 |
2.06 |
|
|
|
| 22 |
A |
1240 |
1191 |
5.63 |
|
|
|
| 23 |
A |
1209 |
1163 |
2.08 |
|
|
|
| 24 |
A |
1184 |
1138 |
3.22 |
|
|
|
| 25 |
A |
1130 |
1086 |
0.25 |
|
|
|
| 26 |
A |
1076 |
1035 |
2.02 |
|
|
|
| 27 |
A |
1002 |
963 |
1.54 |
|
|
|
| 28 |
A |
992 |
954 |
0.97 |
|
|
|
| 29 |
A |
969 |
931 |
4.80 |
|
|
|
| 30 |
A |
938 |
902 |
1.28 |
|
|
|
| 31 |
A |
893 |
858 |
4.31 |
|
|
|
| 32 |
A |
848 |
816 |
5.65 |
|
|
|
| 33 |
A |
819 |
787 |
2.00 |
|
|
|
| 34 |
A |
806 |
775 |
10.26 |
|
|
|
| 35 |
A |
738 |
709 |
7.98 |
|
|
|
| 36 |
A |
638 |
613 |
0.95 |
|
|
|
| 37 |
A |
461 |
443 |
0.52 |
|
|
|
| 38 |
A |
351 |
337 |
3.04 |
|
|
|
| 39 |
A |
241 |
231 |
3.01 |
|
|
|
| 40 |
A |
190 |
183 |
25.85 |
|
|
|
| 41 |
A |
150 |
144 |
1.03 |
|
|
|
| 42 |
A |
34 |
32 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30808.0 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 29612.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.197 |
|
|
|
| 2 |
H |
0.201 |
|
|
|
| 3 |
C |
-0.394 |
|
|
|
| 4 |
H |
0.195 |
|
|
|
| 5 |
H |
0.203 |
|
|
|
| 6 |
C |
-0.388 |
|
|
|
| 7 |
H |
0.204 |
|
|
|
| 8 |
H |
0.217 |
|
|
|
| 9 |
C |
-0.362 |
|
|
|
| 10 |
H |
0.205 |
|
|
|
| 11 |
H |
0.207 |
|
|
|
| 12 |
C |
-0.346 |
|
|
|
| 13 |
H |
0.240 |
|
|
|
| 14 |
C |
-0.407 |
|
|
|
| 15 |
H |
0.072 |
|
|
|
| 16 |
S |
-0.045 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.773 |
0.952 |
0.823 |
2.175 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.272 |
3.743 |
2.072 |
| y |
3.743 |
-44.319 |
0.757 |
| z |
2.072 |
0.757 |
-44.732 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.747 |
3.743 |
2.072 |
| y |
3.743 |
2.684 |
0.757 |
| z |
2.072 |
0.757 |
2.063 |
|
| Polar |
|---|
| 3z2-r2 | 4.126 |
|---|
| x2-y2 | -4.953 |
|---|
| xy | 3.743 |
|---|
| xz | 2.072 |
|---|
| yz | 0.757 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.236 |
0.256 |
-0.458 |
| y |
0.256 |
9.148 |
0.593 |
| z |
-0.458 |
0.593 |
7.414 |
<r2> (average value of r
2) Å
2
| <r2> |
180.851 |
| (<r2>)1/2 |
13.448 |