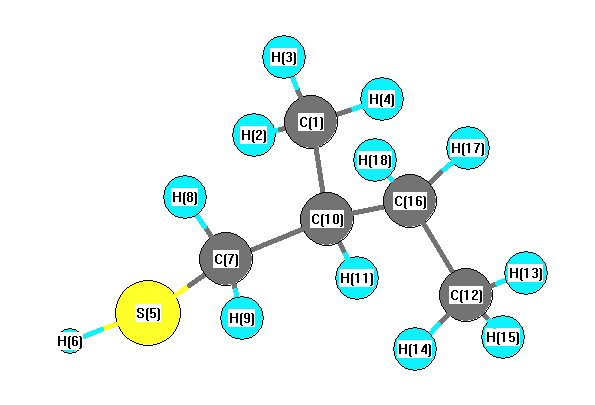
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3166 |
3043 |
28.27 |
|
|
|
| 2 |
A |
3137 |
3015 |
42.56 |
|
|
|
| 3 |
A |
3127 |
3006 |
118.69 |
|
|
|
| 4 |
A |
3123 |
3002 |
55.25 |
|
|
|
| 5 |
A |
3120 |
2999 |
16.84 |
|
|
|
| 6 |
A |
3086 |
2967 |
29.37 |
|
|
|
| 7 |
A |
3076 |
2957 |
14.35 |
|
|
|
| 8 |
A |
3043 |
2925 |
31.18 |
|
|
|
| 9 |
A |
3037 |
2919 |
14.08 |
|
|
|
| 10 |
A |
3035 |
2917 |
44.28 |
|
|
|
| 11 |
A |
3025 |
2908 |
23.10 |
|
|
|
| 12 |
A |
2501 |
2404 |
42.76 |
|
|
|
| 13 |
A |
1532 |
1472 |
15.30 |
|
|
|
| 14 |
A |
1525 |
1466 |
20.36 |
|
|
|
| 15 |
A |
1523 |
1464 |
5.36 |
|
|
|
| 16 |
A |
1523 |
1464 |
5.45 |
|
|
|
| 17 |
A |
1513 |
1454 |
3.36 |
|
|
|
| 18 |
A |
1494 |
1436 |
7.61 |
|
|
|
| 19 |
A |
1440 |
1384 |
24.32 |
|
|
|
| 20 |
A |
1439 |
1383 |
6.11 |
|
|
|
| 21 |
A |
1397 |
1342 |
2.11 |
|
|
|
| 22 |
A |
1376 |
1322 |
1.77 |
|
|
|
| 23 |
A |
1344 |
1292 |
0.35 |
|
|
|
| 24 |
A |
1318 |
1267 |
13.26 |
|
|
|
| 25 |
A |
1291 |
1241 |
3.52 |
|
|
|
| 26 |
A |
1242 |
1194 |
5.00 |
|
|
|
| 27 |
A |
1196 |
1150 |
0.78 |
|
|
|
| 28 |
A |
1160 |
1115 |
3.01 |
|
|
|
| 29 |
A |
1097 |
1054 |
1.38 |
|
|
|
| 30 |
A |
1051 |
1010 |
0.81 |
|
|
|
| 31 |
A |
1048 |
1008 |
5.87 |
|
|
|
| 32 |
A |
998 |
959 |
13.15 |
|
|
|
| 33 |
A |
941 |
904 |
0.72 |
|
|
|
| 34 |
A |
893 |
859 |
7.08 |
|
|
|
| 35 |
A |
871 |
838 |
1.24 |
|
|
|
| 36 |
A |
806 |
775 |
3.30 |
|
|
|
| 37 |
A |
767 |
737 |
12.05 |
|
|
|
| 38 |
A |
688 |
662 |
4.64 |
|
|
|
| 39 |
A |
457 |
439 |
0.36 |
|
|
|
| 40 |
A |
392 |
377 |
0.19 |
|
|
|
| 41 |
A |
377 |
363 |
1.29 |
|
|
|
| 42 |
A |
248 |
239 |
0.08 |
|
|
|
| 43 |
A |
234 |
225 |
1.13 |
|
|
|
| 44 |
A |
208 |
200 |
0.29 |
|
|
|
| 45 |
A |
194 |
186 |
1.84 |
|
|
|
| 46 |
A |
132 |
126 |
19.13 |
|
|
|
| 47 |
A |
92 |
89 |
1.20 |
|
|
|
| 48 |
A |
50 |
48 |
11.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35166.2 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 33801.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.658 |
|
|
|
| 2 |
H |
0.221 |
|
|
|
| 3 |
H |
0.188 |
|
|
|
| 4 |
H |
0.196 |
|
|
|
| 5 |
S |
-0.059 |
|
|
|
| 6 |
H |
0.075 |
|
|
|
| 7 |
C |
-0.676 |
|
|
|
| 8 |
H |
0.234 |
|
|
|
| 9 |
H |
0.253 |
|
|
|
| 10 |
C |
0.066 |
|
|
|
| 11 |
H |
0.186 |
|
|
|
| 12 |
C |
-0.658 |
|
|
|
| 13 |
H |
0.202 |
|
|
|
| 14 |
H |
0.195 |
|
|
|
| 15 |
H |
0.198 |
|
|
|
| 16 |
C |
-0.320 |
|
|
|
| 17 |
H |
0.183 |
|
|
|
| 18 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.728 |
-0.509 |
0.954 |
2.039 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.456 |
1.936 |
-2.121 |
| y |
1.936 |
-44.127 |
-1.839 |
| z |
-2.121 |
-1.839 |
-47.204 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.790 |
1.936 |
-2.121 |
| y |
1.936 |
5.203 |
-1.839 |
| z |
-2.121 |
-1.839 |
0.587 |
|
| Polar |
|---|
| 3z2-r2 | 1.174 |
|---|
| x2-y2 | -7.329 |
|---|
| xy | 1.936 |
|---|
| xz | -2.121 |
|---|
| yz | -1.839 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.230 |
0.857 |
0.771 |
| y |
0.857 |
9.975 |
-0.454 |
| z |
0.771 |
-0.454 |
7.990 |
<r2> (average value of r
2) Å
2
| <r2> |
231.305 |
| (<r2>)1/2 |
15.209 |