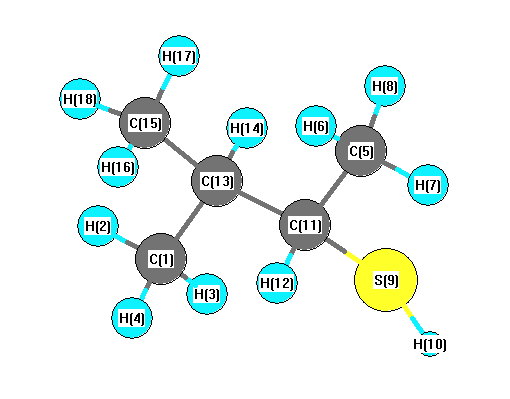
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
3023 |
37.76 |
|
|
|
| 2 |
A |
3141 |
3020 |
48.85 |
|
|
|
| 3 |
A |
3138 |
3016 |
44.37 |
|
|
|
| 4 |
A |
3126 |
3005 |
92.23 |
|
|
|
| 5 |
A |
3124 |
3003 |
41.15 |
|
|
|
| 6 |
A |
3116 |
2995 |
0.99 |
|
|
|
| 7 |
A |
3084 |
2964 |
18.06 |
|
|
|
| 8 |
A |
3042 |
2924 |
24.13 |
|
|
|
| 9 |
A |
3040 |
2922 |
30.04 |
|
|
|
| 10 |
A |
3037 |
2919 |
22.20 |
|
|
|
| 11 |
A |
3034 |
2916 |
18.54 |
|
|
|
| 12 |
A |
2505 |
2408 |
44.02 |
|
|
|
| 13 |
A |
1540 |
1480 |
8.67 |
|
|
|
| 14 |
A |
1528 |
1469 |
9.91 |
|
|
|
| 15 |
A |
1525 |
1465 |
24.30 |
|
|
|
| 16 |
A |
1516 |
1457 |
4.93 |
|
|
|
| 17 |
A |
1514 |
1455 |
8.17 |
|
|
|
| 18 |
A |
1509 |
1451 |
6.04 |
|
|
|
| 19 |
A |
1448 |
1392 |
14.55 |
|
|
|
| 20 |
A |
1439 |
1383 |
22.03 |
|
|
|
| 21 |
A |
1428 |
1373 |
10.80 |
|
|
|
| 22 |
A |
1386 |
1333 |
0.36 |
|
|
|
| 23 |
A |
1354 |
1301 |
0.45 |
|
|
|
| 24 |
A |
1331 |
1279 |
0.75 |
|
|
|
| 25 |
A |
1259 |
1211 |
25.25 |
|
|
|
| 26 |
A |
1216 |
1169 |
4.16 |
|
|
|
| 27 |
A |
1189 |
1143 |
13.10 |
|
|
|
| 28 |
A |
1159 |
1115 |
1.31 |
|
|
|
| 29 |
A |
1096 |
1054 |
3.56 |
|
|
|
| 30 |
A |
1054 |
1014 |
19.88 |
|
|
|
| 31 |
A |
1003 |
964 |
7.39 |
|
|
|
| 32 |
A |
981 |
943 |
0.98 |
|
|
|
| 33 |
A |
949 |
912 |
2.22 |
|
|
|
| 34 |
A |
929 |
893 |
3.99 |
|
|
|
| 35 |
A |
868 |
834 |
3.36 |
|
|
|
| 36 |
A |
778 |
748 |
4.53 |
|
|
|
| 37 |
A |
650 |
624 |
5.79 |
|
|
|
| 38 |
A |
472 |
454 |
0.97 |
|
|
|
| 39 |
A |
403 |
387 |
0.40 |
|
|
|
| 40 |
A |
363 |
349 |
0.21 |
|
|
|
| 41 |
A |
346 |
332 |
0.11 |
|
|
|
| 42 |
A |
325 |
313 |
2.78 |
|
|
|
| 43 |
A |
246 |
237 |
1.95 |
|
|
|
| 44 |
A |
229 |
220 |
1.07 |
|
|
|
| 45 |
A |
214 |
206 |
1.19 |
|
|
|
| 46 |
A |
188 |
181 |
2.45 |
|
|
|
| 47 |
A |
166 |
160 |
21.35 |
|
|
|
| 48 |
A |
61 |
58 |
2.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35096.7 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 33734.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.652 |
|
|
|
| 2 |
H |
0.194 |
|
|
|
| 3 |
H |
0.226 |
|
|
|
| 4 |
H |
0.188 |
|
|
|
| 5 |
C |
-0.627 |
|
|
|
| 6 |
H |
0.211 |
|
|
|
| 7 |
H |
0.199 |
|
|
|
| 8 |
H |
0.218 |
|
|
|
| 9 |
S |
-0.068 |
|
|
|
| 10 |
H |
0.070 |
|
|
|
| 11 |
C |
-0.357 |
|
|
|
| 12 |
H |
0.232 |
|
|
|
| 13 |
C |
0.050 |
|
|
|
| 14 |
H |
0.192 |
|
|
|
| 15 |
C |
-0.678 |
|
|
|
| 16 |
H |
0.192 |
|
|
|
| 17 |
H |
0.211 |
|
|
|
| 18 |
H |
0.201 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.284 |
-1.897 |
0.761 |
2.064 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.137 |
1.940 |
-2.227 |
| y |
1.940 |
-50.631 |
0.622 |
| z |
-2.227 |
0.622 |
-47.044 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.701 |
1.940 |
-2.227 |
| y |
1.940 |
-5.041 |
0.622 |
| z |
-2.227 |
0.622 |
0.340 |
|
| Polar |
|---|
| 3z2-r2 | 0.680 |
|---|
| x2-y2 | 6.494 |
|---|
| xy | 1.940 |
|---|
| xz | -2.227 |
|---|
| yz | 0.622 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.533 |
-0.985 |
-0.170 |
| y |
-0.985 |
11.052 |
-0.694 |
| z |
-0.170 |
-0.694 |
7.775 |
<r2> (average value of r
2) Å
2
| <r2> |
212.277 |
| (<r2>)1/2 |
14.570 |