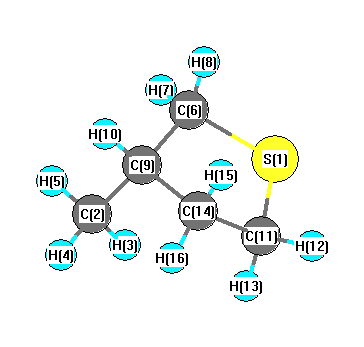
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3171 |
3048 |
33.80 |
|
|
|
| 2 |
A |
3164 |
3041 |
19.42 |
|
|
|
| 3 |
A |
3136 |
3014 |
46.17 |
|
|
|
| 4 |
A |
3124 |
3003 |
49.21 |
|
|
|
| 5 |
A |
3112 |
2991 |
31.94 |
|
|
|
| 6 |
A |
3099 |
2979 |
6.69 |
|
|
|
| 7 |
A |
3095 |
2975 |
63.53 |
|
|
|
| 8 |
A |
3059 |
2940 |
51.62 |
|
|
|
| 9 |
A |
3042 |
2924 |
17.61 |
|
|
|
| 10 |
A |
3039 |
2921 |
27.30 |
|
|
|
| 11 |
A |
1532 |
1473 |
9.63 |
|
|
|
| 12 |
A |
1524 |
1465 |
3.21 |
|
|
|
| 13 |
A |
1515 |
1456 |
9.75 |
|
|
|
| 14 |
A |
1506 |
1447 |
3.54 |
|
|
|
| 15 |
A |
1503 |
1444 |
9.54 |
|
|
|
| 16 |
A |
1438 |
1383 |
17.49 |
|
|
|
| 17 |
A |
1382 |
1328 |
2.48 |
|
|
|
| 18 |
A |
1358 |
1306 |
9.57 |
|
|
|
| 19 |
A |
1332 |
1280 |
1.58 |
|
|
|
| 20 |
A |
1312 |
1261 |
8.96 |
|
|
|
| 21 |
A |
1271 |
1221 |
11.84 |
|
|
|
| 22 |
A |
1247 |
1198 |
4.21 |
|
|
|
| 23 |
A |
1193 |
1146 |
4.56 |
|
|
|
| 24 |
A |
1152 |
1107 |
0.08 |
|
|
|
| 25 |
A |
1098 |
1055 |
0.95 |
|
|
|
| 26 |
A |
1074 |
1032 |
1.39 |
|
|
|
| 27 |
A |
1059 |
1018 |
1.21 |
|
|
|
| 28 |
A |
1011 |
971 |
4.13 |
|
|
|
| 29 |
A |
975 |
937 |
0.36 |
|
|
|
| 30 |
A |
942 |
905 |
1.50 |
|
|
|
| 31 |
A |
911 |
875 |
5.50 |
|
|
|
| 32 |
A |
864 |
831 |
2.09 |
|
|
|
| 33 |
A |
797 |
766 |
3.59 |
|
|
|
| 34 |
A |
702 |
675 |
7.01 |
|
|
|
| 35 |
A |
647 |
622 |
4.00 |
|
|
|
| 36 |
A |
584 |
561 |
0.78 |
|
|
|
| 37 |
A |
486 |
467 |
0.25 |
|
|
|
| 38 |
A |
378 |
363 |
0.30 |
|
|
|
| 39 |
A |
353 |
339 |
0.39 |
|
|
|
| 40 |
A |
226 |
217 |
0.07 |
|
|
|
| 41 |
A |
193 |
186 |
0.35 |
|
|
|
| 42 |
A |
108 |
104 |
2.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31355.0 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 30138.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.143 |
|
|
|
| 2 |
C |
-0.673 |
|
|
|
| 3 |
H |
0.226 |
|
|
|
| 4 |
H |
0.197 |
|
|
|
| 5 |
H |
0.201 |
|
|
|
| 6 |
C |
-0.667 |
|
|
|
| 7 |
H |
0.236 |
|
|
|
| 8 |
H |
0.245 |
|
|
|
| 9 |
C |
-0.003 |
|
|
|
| 10 |
H |
0.199 |
|
|
|
| 11 |
C |
-0.663 |
|
|
|
| 12 |
H |
0.243 |
|
|
|
| 13 |
H |
0.246 |
|
|
|
| 14 |
C |
-0.333 |
|
|
|
| 15 |
H |
0.198 |
|
|
|
| 16 |
H |
0.204 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.619 |
-1.561 |
1.178 |
2.539 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.949 |
-4.388 |
0.733 |
| y |
-4.388 |
-44.970 |
1.901 |
| z |
0.733 |
1.901 |
-45.549 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.689 |
-4.388 |
0.733 |
| y |
-4.388 |
1.279 |
1.901 |
| z |
0.733 |
1.901 |
0.411 |
|
| Polar |
|---|
| 3z2-r2 | 0.821 |
|---|
| x2-y2 | -1.979 |
|---|
| xy | -4.388 |
|---|
| xz | 0.733 |
|---|
| yz | 1.901 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.162 |
-0.139 |
-1.027 |
| y |
-0.139 |
10.377 |
-0.165 |
| z |
-1.027 |
-0.165 |
8.205 |
<r2> (average value of r
2) Å
2
| <r2> |
169.251 |
| (<r2>)1/2 |
13.010 |