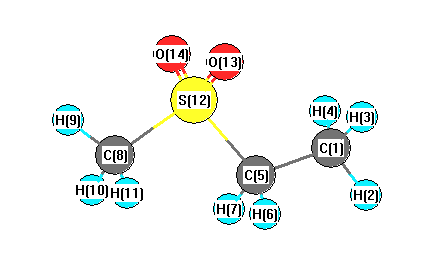
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3239 |
3114 |
2.35 |
|
|
|
| 2 |
A |
3234 |
3108 |
0.50 |
|
|
|
| 3 |
A |
3194 |
3070 |
6.39 |
|
|
|
| 4 |
A |
3157 |
3035 |
4.54 |
|
|
|
| 5 |
A |
3139 |
3017 |
19.93 |
|
|
|
| 6 |
A |
3109 |
2989 |
4.68 |
|
|
|
| 7 |
A |
3095 |
2975 |
2.57 |
|
|
|
| 8 |
A |
3054 |
2935 |
16.87 |
|
|
|
| 9 |
A |
1527 |
1468 |
13.17 |
|
|
|
| 10 |
A |
1511 |
1452 |
18.24 |
|
|
|
| 11 |
A |
1478 |
1420 |
22.16 |
|
|
|
| 12 |
A |
1474 |
1417 |
2.78 |
|
|
|
| 13 |
A |
1457 |
1401 |
9.85 |
|
|
|
| 14 |
A |
1445 |
1389 |
21.16 |
|
|
|
| 15 |
A |
1368 |
1315 |
4.42 |
|
|
|
| 16 |
A |
1313 |
1262 |
8.18 |
|
|
|
| 17 |
A |
1268 |
1219 |
2.01 |
|
|
|
| 18 |
A |
1100 |
1057 |
17.16 |
|
|
|
| 19 |
A |
1092 |
1049 |
13.09 |
|
|
|
| 20 |
A |
1034 |
994 |
34.17 |
|
|
|
| 21 |
A |
1012 |
972 |
9.36 |
|
|
|
| 22 |
A |
995 |
957 |
28.19 |
|
|
|
| 23 |
A |
885 |
851 |
41.76 |
|
|
|
| 24 |
A |
807 |
775 |
44.57 |
|
|
|
| 25 |
A |
774 |
744 |
25.07 |
|
|
|
| 26 |
A |
633 |
608 |
34.37 |
|
|
|
| 27 |
A |
529 |
508 |
3.76 |
|
|
|
| 28 |
A |
389 |
374 |
8.39 |
|
|
|
| 29 |
A |
320 |
307 |
27.84 |
|
|
|
| 30 |
A |
295 |
284 |
29.95 |
|
|
|
| 31 |
A |
247 |
238 |
1.85 |
|
|
|
| 32 |
A |
230 |
221 |
1.65 |
|
|
|
| 33 |
A |
182 |
175 |
2.76 |
|
|
|
| 34 |
A |
172 |
166 |
2.04 |
|
|
|
| 35 |
A |
140 |
135 |
0.21 |
|
|
|
| 36 |
A |
67 |
64 |
3.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24482.0 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 23532.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.595 |
|
|
|
| 2 |
H |
0.263 |
|
|
|
| 3 |
H |
0.223 |
|
|
|
| 4 |
H |
0.204 |
|
|
|
| 5 |
C |
-0.501 |
|
|
|
| 6 |
H |
0.285 |
|
|
|
| 7 |
H |
0.244 |
|
|
|
| 8 |
C |
-0.708 |
|
|
|
| 9 |
H |
0.251 |
|
|
|
| 10 |
H |
0.289 |
|
|
|
| 11 |
H |
0.253 |
|
|
|
| 12 |
S |
1.013 |
|
|
|
| 13 |
O |
-0.613 |
|
|
|
| 14 |
O |
-0.609 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.945 |
3.306 |
-3.797 |
6.396 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.284 |
-7.847 |
-0.283 |
| y |
-7.847 |
-43.453 |
-2.092 |
| z |
-0.283 |
-2.092 |
-50.311 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.598 |
-7.847 |
-0.283 |
| y |
-7.847 |
4.345 |
-2.092 |
| z |
-0.283 |
-2.092 |
-5.942 |
|
| Polar |
|---|
| 3z2-r2 | -11.885 |
|---|
| x2-y2 | -1.831 |
|---|
| xy | -7.847 |
|---|
| xz | -0.283 |
|---|
| yz | -2.092 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.087 |
-0.030 |
0.269 |
| y |
-0.030 |
8.793 |
0.211 |
| z |
0.269 |
0.211 |
8.638 |
<r2> (average value of r
2) Å
2
| <r2> |
208.633 |
| (<r2>)1/2 |
14.444 |