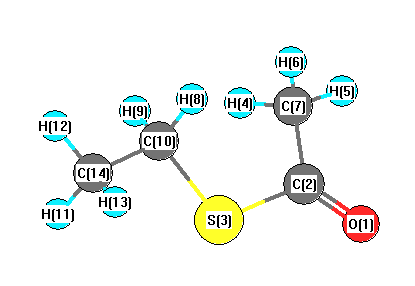
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3178 |
3055 |
11.92 |
|
|
|
| 2 |
A |
3171 |
3047 |
49.47 |
|
|
|
| 3 |
A |
3144 |
3022 |
7.73 |
|
|
|
| 4 |
A |
3142 |
3020 |
7.59 |
|
|
|
| 5 |
A |
3136 |
3015 |
31.84 |
|
|
|
| 6 |
A |
3089 |
2969 |
22.09 |
|
|
|
| 7 |
A |
3057 |
2938 |
2.79 |
|
|
|
| 8 |
A |
3049 |
2930 |
31.41 |
|
|
|
| 9 |
A |
1683 |
1618 |
307.70 |
|
|
|
| 10 |
A |
1526 |
1467 |
8.26 |
|
|
|
| 11 |
A |
1515 |
1457 |
15.68 |
|
|
|
| 12 |
A |
1503 |
1445 |
3.75 |
|
|
|
| 13 |
A |
1503 |
1445 |
21.92 |
|
|
|
| 14 |
A |
1488 |
1430 |
38.10 |
|
|
|
| 15 |
A |
1447 |
1391 |
16.97 |
|
|
|
| 16 |
A |
1419 |
1364 |
31.95 |
|
|
|
| 17 |
A |
1322 |
1271 |
26.71 |
|
|
|
| 18 |
A |
1285 |
1235 |
0.33 |
|
|
|
| 19 |
A |
1151 |
1107 |
141.11 |
|
|
|
| 20 |
A |
1091 |
1049 |
6.13 |
|
|
|
| 21 |
A |
1064 |
1023 |
0.00 |
|
|
|
| 22 |
A |
1045 |
1005 |
3.70 |
|
|
|
| 23 |
A |
1012 |
973 |
11.49 |
|
|
|
| 24 |
A |
956 |
919 |
53.92 |
|
|
|
| 25 |
A |
808 |
777 |
11.35 |
|
|
|
| 26 |
A |
643 |
618 |
0.18 |
|
|
|
| 27 |
A |
602 |
579 |
76.56 |
|
|
|
| 28 |
A |
506 |
487 |
1.40 |
|
|
|
| 29 |
A |
436 |
419 |
5.01 |
|
|
|
| 30 |
A |
359 |
345 |
1.78 |
|
|
|
| 31 |
A |
297 |
285 |
1.33 |
|
|
|
| 32 |
A |
245 |
235 |
0.04 |
|
|
|
| 33 |
A |
156 |
150 |
1.06 |
|
|
|
| 34 |
A |
138 |
132 |
0.10 |
|
|
|
| 35 |
A |
72 |
69 |
0.85 |
|
|
|
| 36 |
A |
32 |
31 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25134.9 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 24159.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.212 |
|
|
|
| 2 |
C |
0.012 |
|
|
|
| 3 |
S |
0.165 |
|
|
|
| 4 |
H |
0.225 |
|
|
|
| 5 |
H |
0.247 |
|
|
|
| 6 |
H |
0.225 |
|
|
|
| 7 |
C |
-0.627 |
|
|
|
| 8 |
H |
0.238 |
|
|
|
| 9 |
H |
0.238 |
|
|
|
| 10 |
C |
-0.552 |
|
|
|
| 11 |
H |
0.220 |
|
|
|
| 12 |
H |
0.208 |
|
|
|
| 13 |
H |
0.220 |
|
|
|
| 14 |
C |
-0.607 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.024 |
3.971 |
0.000 |
4.991 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.064 |
4.888 |
0.002 |
| y |
4.888 |
-45.906 |
0.000 |
| z |
0.002 |
0.000 |
-43.608 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.307 |
4.888 |
0.002 |
| y |
4.888 |
-0.570 |
0.000 |
| z |
0.002 |
0.000 |
2.877 |
|
| Polar |
|---|
| 3z2-r2 | 5.754 |
|---|
| x2-y2 | -1.158 |
|---|
| xy | 4.888 |
|---|
| xz | 0.002 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.335 |
-0.369 |
0.006 |
| y |
-0.369 |
8.734 |
0.001 |
| z |
0.006 |
0.001 |
5.681 |
<r2> (average value of r
2) Å
2
| <r2> |
252.848 |
| (<r2>)1/2 |
15.901 |