Jump to
S1C2
Energy calculated at B3LYP/LANL2DZ
| | hartrees |
|---|
| Energy at 0K | -166.051642 |
| Energy at 298.15K | -166.057466 |
| Nuclear repulsion energy | 126.142248 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3272 |
3145 |
23.18 |
|
|
|
| 2 |
A |
3200 |
3075 |
13.41 |
|
|
|
| 3 |
A |
3164 |
3041 |
4.01 |
|
|
|
| 4 |
A |
1666 |
1601 |
4.64 |
|
|
|
| 5 |
A |
1442 |
1386 |
14.97 |
|
|
|
| 6 |
A |
1330 |
1279 |
0.17 |
|
|
|
| 7 |
A |
1074 |
1033 |
3.78 |
|
|
|
| 8 |
A |
975 |
937 |
0.00 |
|
|
|
| 9 |
A |
957 |
920 |
0.00 |
|
|
|
| 10 |
A |
678 |
651 |
0.00 |
|
|
|
| 11 |
A |
612 |
588 |
0.00 |
|
|
|
| 12 |
A |
392 |
376 |
1.55 |
|
|
|
| 13 |
A |
172 |
165 |
0.00 |
|
|
|
| 14 |
A |
3272 |
3145 |
2.75 |
|
|
|
| 15 |
A |
3200 |
3076 |
3.49 |
|
|
|
| 16 |
A |
3164 |
3041 |
0.20 |
|
|
|
| 17 |
A |
1641 |
1578 |
188.46 |
|
|
|
| 18 |
A |
1433 |
1377 |
5.62 |
|
|
|
| 19 |
A |
1290 |
1240 |
21.01 |
|
|
|
| 20 |
A |
1046 |
1005 |
32.86 |
|
|
|
| 21 |
A |
988 |
949 |
118.52 |
|
|
|
| 22 |
A |
958 |
920 |
123.68 |
|
|
|
| 23 |
A |
678 |
652 |
50.52 |
|
|
|
| 24 |
A |
560 |
539 |
38.16 |
|
|
|
| 25 |
A |
388 |
373 |
1.56 |
|
|
|
| 26 |
A |
31 |
30 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18790.7 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 18061.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/LANL2DZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.850 |
| C2 |
0.000 |
1.403 |
-0.323 |
| C3 |
0.000 |
-1.403 |
-0.323 |
| C4 |
0.114 |
2.689 |
0.058 |
| C5 |
-0.114 |
-2.689 |
0.058 |
| H6 |
-0.119 |
1.124 |
-1.368 |
| H7 |
0.119 |
-1.124 |
-1.368 |
| H8 |
0.089 |
3.482 |
-0.685 |
| H9 |
0.232 |
2.988 |
1.096 |
| H10 |
-0.089 |
-3.482 |
-0.685 |
| H11 |
-0.232 |
-2.988 |
1.096 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.8283 | 1.8283 | 2.8053 | 2.8053 | 2.4896 | 2.4896 | 3.8063 | 3.0069 | 3.8063 | 3.0069 |
C2 | 1.8283 | | 2.8057 | 1.3458 | 4.1108 | 1.0887 | 2.7375 | 2.1124 | 2.1399 | 4.8991 | 4.6201 | C3 | 1.8283 | 2.8057 | | 4.1108 | 1.3458 | 2.7375 | 1.0887 | 4.8991 | 4.6201 | 2.1124 | 2.1399 | C4 | 2.8053 | 1.3458 | 4.1108 | | 5.3823 | 2.1297 | 4.0710 | 1.0870 | 1.0871 | 6.2186 | 5.7811 | C5 | 2.8053 | 4.1108 | 1.3458 | 5.3823 | | 4.0710 | 2.1297 | 6.2186 | 5.7811 | 1.0870 | 1.0871 | H6 | 2.4896 | 1.0887 | 2.7375 | 2.1297 | 4.0710 | | 2.2612 | 2.4636 | 3.1096 | 4.6568 | 4.7954 | H7 | 2.4896 | 2.7375 | 1.0887 | 4.0710 | 2.1297 | 2.2612 | | 4.6568 | 4.7954 | 2.4636 | 3.1096 | H8 | 3.8063 | 2.1124 | 4.8991 | 1.0870 | 6.2186 | 2.4636 | 4.6568 | | 1.8540 | 6.9663 | 6.7182 | H9 | 3.0069 | 2.1399 | 4.6201 | 1.0871 | 5.7811 | 3.1096 | 4.7954 | 1.8540 | | 6.7182 | 5.9935 | H10 | 3.8063 | 4.8991 | 2.1124 | 6.2186 | 1.0870 | 4.6568 | 2.4636 | 6.9663 | 6.7182 | | 1.8540 | H11 | 3.0069 | 4.6201 | 2.1399 | 5.7811 | 1.0871 | 4.7954 | 3.1096 | 6.7182 | 5.9935 | 1.8540 | |
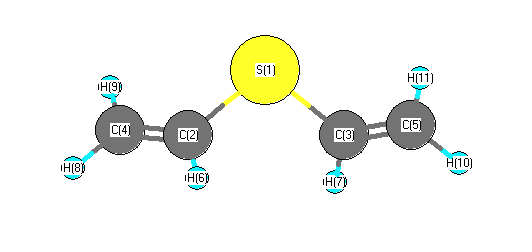 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.494 |
|
S1 |
C2 |
H6 |
114.809 |
| S1 |
C3 |
C5 |
123.494 |
|
S1 |
C3 |
H7 |
114.809 |
| C2 |
S1 |
C3 |
100.224 |
|
C2 |
C4 |
H8 |
120.146 |
| C2 |
C4 |
H9 |
122.825 |
|
C3 |
C5 |
H10 |
120.146 |
| C3 |
C5 |
H11 |
122.825 |
|
C4 |
C2 |
H6 |
121.687 |
| C5 |
C3 |
H7 |
121.687 |
|
H8 |
C4 |
H9 |
117.029 |
| H10 |
C5 |
H11 |
117.029 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.178 |
|
|
|
| 2 |
C |
-0.239 |
|
|
|
| 3 |
C |
-0.239 |
|
|
|
| 4 |
C |
-0.503 |
|
|
|
| 5 |
C |
-0.503 |
|
|
|
| 6 |
H |
0.210 |
|
|
|
| 7 |
H |
0.210 |
|
|
|
| 8 |
H |
0.218 |
|
|
|
| 9 |
H |
0.224 |
|
|
|
| 10 |
H |
0.218 |
|
|
|
| 11 |
H |
0.224 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.460 |
1.460 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.968 |
-0.026 |
0.000 |
| y |
-0.026 |
16.828 |
0.000 |
| z |
0.000 |
0.000 |
7.109 |
<r2> (average value of r
2) Å
2
| <r2> |
190.883 |
| (<r2>)1/2 |
13.816 |
Jump to
S1C1
Energy calculated at B3LYP/LANL2DZ
| | hartrees |
|---|
| Energy at 0K | -166.052172 |
| Energy at 298.15K | -166.057499 |
| HF Energy | -166.052172 |
| Nuclear repulsion energy | 129.363078 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3272 |
3145 |
21.17 |
|
|
|
| 2 |
A |
3271 |
3144 |
10.36 |
|
|
|
| 3 |
A |
3219 |
3094 |
7.52 |
|
|
|
| 4 |
A |
3205 |
3081 |
8.78 |
|
|
|
| 5 |
A |
3165 |
3042 |
1.13 |
|
|
|
| 6 |
A |
3162 |
3039 |
1.56 |
|
|
|
| 7 |
A |
1650 |
1586 |
56.62 |
|
|
|
| 8 |
A |
1643 |
1579 |
40.80 |
|
|
|
| 9 |
A |
1437 |
1381 |
7.42 |
|
|
|
| 10 |
A |
1433 |
1377 |
9.15 |
|
|
|
| 11 |
A |
1313 |
1262 |
3.01 |
|
|
|
| 12 |
A |
1298 |
1248 |
12.65 |
|
|
|
| 13 |
A |
1069 |
1028 |
24.65 |
|
|
|
| 14 |
A |
1048 |
1007 |
11.65 |
|
|
|
| 15 |
A |
1006 |
967 |
33.83 |
|
|
|
| 16 |
A |
995 |
956 |
64.24 |
|
|
|
| 17 |
A |
982 |
944 |
57.61 |
|
|
|
| 18 |
A |
954 |
917 |
80.07 |
|
|
|
| 19 |
A |
690 |
664 |
19.38 |
|
|
|
| 20 |
A |
646 |
621 |
2.17 |
|
|
|
| 21 |
A |
619 |
595 |
25.38 |
|
|
|
| 22 |
A |
603 |
579 |
18.43 |
|
|
|
| 23 |
A |
452 |
434 |
0.88 |
|
|
|
| 24 |
A |
361 |
347 |
0.92 |
|
|
|
| 25 |
A |
205 |
197 |
0.56 |
|
|
|
| 26 |
A |
135 |
130 |
3.79 |
|
|
|
| 27 |
A |
78 |
75 |
1.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18954.8 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 18219.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/LANL2DZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.023 |
-0.995 |
0.000 |
| C2 |
1.125 |
0.424 |
0.345 |
| C3 |
-1.626 |
-0.205 |
-0.003 |
| C4 |
2.349 |
0.548 |
-0.201 |
| C5 |
-1.908 |
1.105 |
-0.137 |
| H6 |
0.744 |
1.121 |
1.089 |
| H7 |
-2.409 |
-0.950 |
0.120 |
| H8 |
3.010 |
1.355 |
0.109 |
| H9 |
2.726 |
-0.141 |
-0.952 |
| H10 |
-1.144 |
1.862 |
-0.291 |
| H11 |
-2.941 |
1.445 |
-0.107 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.8295 | 1.8285 | 2.7990 | 2.8571 | 2.4867 | 2.4351 | 3.8024 | 2.9905 | 3.1007 | 3.8414 |
C2 | 1.8295 | | 2.8431 | 1.3464 | 3.1460 | 1.0882 | 3.7979 | 2.1156 | 2.1367 | 2.7608 | 4.2167 | C3 | 1.8285 | 2.8431 | | 4.0509 | 1.3477 | 2.9270 | 1.0873 | 4.8925 | 4.4550 | 2.1426 | 2.1132 | C4 | 2.7990 | 1.3464 | 4.0509 | | 4.2946 | 2.1374 | 4.9986 | 1.0880 | 1.0869 | 3.7338 | 5.3671 | C5 | 2.8571 | 3.1460 | 1.3477 | 4.2946 | | 2.9223 | 2.1310 | 4.9306 | 4.8681 | 1.0864 | 1.0879 | H6 | 2.4867 | 1.0882 | 2.9270 | 2.1374 | 2.9223 | | 3.8946 | 2.4795 | 3.1123 | 2.4536 | 3.8882 | H7 | 2.4351 | 3.7979 | 1.0873 | 4.9986 | 2.1310 | 3.8946 | | 5.8884 | 5.3075 | 3.1107 | 2.4642 | H8 | 3.8024 | 2.1156 | 4.8925 | 1.0880 | 4.9306 | 2.4795 | 5.8884 | | 1.8564 | 4.2039 | 5.9557 | H9 | 2.9905 | 2.1367 | 4.4550 | 1.0869 | 4.8681 | 3.1123 | 5.3075 | 1.8564 | | 4.4084 | 5.9460 | H10 | 3.1007 | 2.7608 | 2.1426 | 3.7338 | 1.0864 | 2.4536 | 3.1107 | 4.2039 | 4.4084 | | 1.8540 | H11 | 3.8414 | 4.2167 | 2.1132 | 5.3671 | 1.0879 | 3.8882 | 2.4642 | 5.9557 | 5.9460 | 1.8540 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.883 |
|
S1 |
C2 |
H6 |
114.515 |
| S1 |
C3 |
C5 |
127.550 |
|
S1 |
C3 |
H7 |
110.681 |
| C2 |
S1 |
C3 |
102.016 |
|
C2 |
C4 |
H8 |
120.329 |
| C2 |
C4 |
H9 |
122.474 |
|
C3 |
C5 |
H10 |
122.982 |
| C3 |
C5 |
H11 |
119.999 |
|
C4 |
C2 |
H6 |
122.428 |
| C5 |
C3 |
H7 |
121.768 |
|
H8 |
C4 |
H9 |
117.195 |
| H10 |
C5 |
H11 |
117.015 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.190 |
|
|
|
| 2 |
C |
-0.326 |
|
|
|
| 3 |
C |
-0.330 |
|
|
|
| 4 |
C |
-0.455 |
|
|
|
| 5 |
C |
-0.438 |
|
|
|
| 6 |
H |
0.245 |
|
|
|
| 7 |
H |
0.227 |
|
|
|
| 8 |
H |
0.218 |
|
|
|
| 9 |
H |
0.229 |
|
|
|
| 10 |
H |
0.226 |
|
|
|
| 11 |
H |
0.213 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.138 |
1.256 |
0.219 |
1.283 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.566 |
1.762 |
-0.975 |
| y |
1.762 |
-36.712 |
1.329 |
| z |
-0.975 |
1.329 |
-40.517 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.048 |
1.762 |
-0.975 |
| y |
1.762 |
-0.671 |
1.329 |
| z |
-0.975 |
1.329 |
-6.378 |
|
| Polar |
|---|
| 3z2-r2 | -12.755 |
|---|
| x2-y2 | 5.146 |
|---|
| xy | 1.762 |
|---|
| xz | -0.975 |
|---|
| yz | 1.329 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.043 |
-0.117 |
-0.738 |
| y |
-0.117 |
8.755 |
0.295 |
| z |
-0.738 |
0.295 |
4.010 |
<r2> (average value of r
2) Å
2
| <r2> |
163.150 |
| (<r2>)1/2 |
12.773 |
