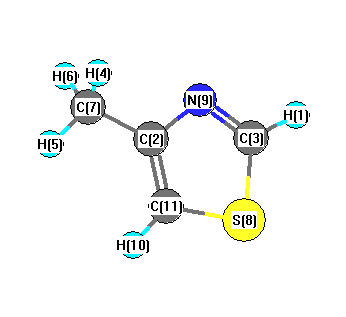
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3310 |
3181 |
2.36 |
|
|
|
| 2 |
A |
3290 |
3162 |
0.07 |
|
|
|
| 3 |
A |
3157 |
3035 |
20.87 |
|
|
|
| 4 |
A |
3131 |
3009 |
19.98 |
|
|
|
| 5 |
A |
3052 |
2933 |
24.19 |
|
|
|
| 6 |
A |
1578 |
1516 |
15.11 |
|
|
|
| 7 |
A |
1509 |
1450 |
26.94 |
|
|
|
| 8 |
A |
1502 |
1444 |
13.44 |
|
|
|
| 9 |
A |
1479 |
1422 |
53.98 |
|
|
|
| 10 |
A |
1448 |
1392 |
19.13 |
|
|
|
| 11 |
A |
1331 |
1279 |
10.29 |
|
|
|
| 12 |
A |
1250 |
1202 |
0.98 |
|
|
|
| 13 |
A |
1175 |
1129 |
3.34 |
|
|
|
| 14 |
A |
1088 |
1046 |
11.16 |
|
|
|
| 15 |
A |
1035 |
995 |
9.76 |
|
|
|
| 16 |
A |
915 |
879 |
11.28 |
|
|
|
| 17 |
A |
834 |
801 |
32.64 |
|
|
|
| 18 |
A |
830 |
798 |
81.86 |
|
|
|
| 19 |
A |
773 |
743 |
1.35 |
|
|
|
| 20 |
A |
773 |
743 |
11.61 |
|
|
|
| 21 |
A |
664 |
638 |
0.27 |
|
|
|
| 22 |
A |
629 |
604 |
12.02 |
|
|
|
| 23 |
A |
542 |
521 |
7.08 |
|
|
|
| 24 |
A |
474 |
455 |
4.74 |
|
|
|
| 25 |
A |
328 |
316 |
3.18 |
|
|
|
| 26 |
A |
232 |
223 |
5.80 |
|
|
|
| 27 |
A |
124 |
119 |
0.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18224.8 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 17517.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.265 |
|
|
|
| 2 |
C |
0.373 |
|
|
|
| 3 |
C |
-0.443 |
|
|
|
| 4 |
H |
0.229 |
|
|
|
| 5 |
H |
0.207 |
|
|
|
| 6 |
H |
0.229 |
|
|
|
| 7 |
C |
-0.690 |
|
|
|
| 8 |
S |
0.243 |
|
|
|
| 9 |
N |
-0.031 |
|
|
|
| 10 |
H |
0.270 |
|
|
|
| 11 |
C |
-0.651 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.389 |
1.185 |
0.000 |
1.247 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.657 |
3.793 |
0.000 |
| y |
3.793 |
-42.368 |
0.000 |
| z |
0.000 |
0.000 |
-44.354 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.704 |
3.793 |
0.000 |
| y |
3.793 |
-1.363 |
0.000 |
| z |
0.000 |
0.000 |
-4.341 |
|
| Polar |
|---|
| 3z2-r2 | -8.682 |
|---|
| x2-y2 | 4.711 |
|---|
| xy | 3.793 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.508 |
-0.604 |
-0.000 |
| y |
-0.604 |
9.855 |
0.002 |
| z |
-0.000 |
0.002 |
4.235 |
<r2> (average value of r
2) Å
2
| <r2> |
150.966 |
| (<r2>)1/2 |
12.287 |