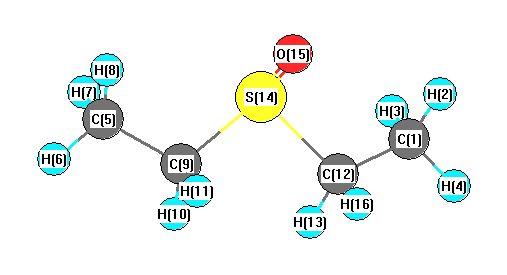
Vibrational Frequencies calculated at B3LYP/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3187 |
3064 |
14.22 |
|
|
|
| 2 |
A |
3175 |
3052 |
21.59 |
|
|
|
| 3 |
A |
3137 |
3015 |
37.72 |
|
|
|
| 4 |
A |
3136 |
3014 |
19.84 |
|
|
|
| 5 |
A |
3132 |
3011 |
6.93 |
|
|
|
| 6 |
A |
3131 |
3009 |
18.40 |
|
|
|
| 7 |
A |
3109 |
2988 |
2.31 |
|
|
|
| 8 |
A |
3098 |
2978 |
20.51 |
|
|
|
| 9 |
A |
3046 |
2927 |
24.52 |
|
|
|
| 10 |
A |
3041 |
2923 |
27.06 |
|
|
|
| 11 |
A |
1533 |
1473 |
12.19 |
|
|
|
| 12 |
A |
1523 |
1464 |
11.54 |
|
|
|
| 13 |
A |
1520 |
1461 |
16.21 |
|
|
|
| 14 |
A |
1513 |
1455 |
13.03 |
|
|
|
| 15 |
A |
1489 |
1431 |
11.55 |
|
|
|
| 16 |
A |
1479 |
1421 |
7.62 |
|
|
|
| 17 |
A |
1446 |
1390 |
20.63 |
|
|
|
| 18 |
A |
1442 |
1386 |
23.82 |
|
|
|
| 19 |
A |
1312 |
1261 |
3.96 |
|
|
|
| 20 |
A |
1293 |
1243 |
10.61 |
|
|
|
| 21 |
A |
1269 |
1220 |
4.06 |
|
|
|
| 22 |
A |
1264 |
1215 |
0.33 |
|
|
|
| 23 |
A |
1100 |
1057 |
9.51 |
|
|
|
| 24 |
A |
1083 |
1041 |
6.56 |
|
|
|
| 25 |
A |
1070 |
1029 |
4.23 |
|
|
|
| 26 |
A |
1043 |
1003 |
2.88 |
|
|
|
| 27 |
A |
1008 |
969 |
9.27 |
|
|
|
| 28 |
A |
1001 |
962 |
10.29 |
|
|
|
| 29 |
A |
806 |
774 |
17.85 |
|
|
|
| 30 |
A |
777 |
747 |
10.19 |
|
|
|
| 31 |
A |
757 |
728 |
19.52 |
|
|
|
| 32 |
A |
596 |
572 |
5.91 |
|
|
|
| 33 |
A |
511 |
491 |
3.48 |
|
|
|
| 34 |
A |
428 |
411 |
3.01 |
|
|
|
| 35 |
A |
330 |
317 |
1.06 |
|
|
|
| 36 |
A |
260 |
250 |
4.42 |
|
|
|
| 37 |
A |
246 |
236 |
6.10 |
|
|
|
| 38 |
A |
200 |
192 |
5.86 |
|
|
|
| 39 |
A |
194 |
187 |
6.54 |
|
|
|
| 40 |
A |
177 |
170 |
2.89 |
|
|
|
| 41 |
A |
111 |
106 |
3.05 |
|
|
|
| 42 |
A |
45 |
44 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30006.3 cm
-1
Scaled (by 0.9612) Zero Point Vibrational Energy (zpe) 28842.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.606 |
|
|
|
| 2 |
H |
0.210 |
|
|
|
| 3 |
H |
0.195 |
|
|
|
| 4 |
H |
0.281 |
|
|
|
| 5 |
C |
-0.610 |
|
|
|
| 6 |
H |
0.217 |
|
|
|
| 7 |
H |
0.223 |
|
|
|
| 8 |
H |
0.207 |
|
|
|
| 9 |
C |
-0.542 |
|
|
|
| 10 |
H |
0.262 |
|
|
|
| 11 |
H |
0.260 |
|
|
|
| 12 |
C |
-0.547 |
|
|
|
| 13 |
H |
0.223 |
|
|
|
| 14 |
S |
0.666 |
|
|
|
| 15 |
O |
-0.684 |
|
|
|
| 16 |
H |
0.246 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.784 |
-4.456 |
0.588 |
5.287 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.655 |
-6.178 |
1.766 |
| y |
-6.178 |
-52.601 |
0.061 |
| z |
1.766 |
0.061 |
-44.147 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.719 |
-6.178 |
1.766 |
| y |
-6.178 |
-9.200 |
0.061 |
| z |
1.766 |
0.061 |
3.481 |
|
| Polar |
|---|
| 3z2-r2 | 6.961 |
|---|
| x2-y2 | 9.946 |
|---|
| xy | -6.178 |
|---|
| xz | 1.766 |
|---|
| yz | 0.061 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.452 |
0.051 |
0.458 |
| y |
0.051 |
10.141 |
-0.302 |
| z |
0.458 |
-0.302 |
7.957 |
<r2> (average value of r
2) Å
2
| <r2> |
228.938 |
| (<r2>)1/2 |
15.131 |