Jump to
S1C2
Energy calculated at HF/SDD
| | hartrees |
|---|
| Energy at 0K | -453.622395 |
| Energy at 298.15K | -453.626052 |
| HF Energy | -453.622395 |
| Nuclear repulsion energy | 57.044280 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3836 |
3454 |
13.54 |
|
|
|
| 2 |
A' |
2632 |
2370 |
46.28 |
|
|
|
| 3 |
A' |
1771 |
1594 |
10.54 |
|
|
|
| 4 |
A' |
1089 |
981 |
7.87 |
|
|
|
| 5 |
A' |
845 |
761 |
43.23 |
|
|
|
| 6 |
A' |
434 |
391 |
360.34 |
|
|
|
| 7 |
A" |
3993 |
3595 |
44.27 |
|
|
|
| 8 |
A" |
1156 |
1041 |
8.63 |
|
|
|
| 9 |
A" |
515 |
464 |
59.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8135.4 cm
-1
Scaled (by 0.9004) Zero Point Vibrational Energy (zpe) 7325.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/SDD
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.032 |
1.114 |
0.000 |
| S2 |
0.032 |
-0.631 |
0.000 |
| H3 |
-1.308 |
-0.828 |
0.000 |
| H4 |
0.290 |
1.564 |
0.848 |
| H5 |
0.290 |
1.564 |
-0.848 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7457 | 2.3596 | 0.9942 | 0.9942 |
S2 | 1.7457 | | 1.3537 | 2.3675 | 2.3675 | H3 | 2.3596 | 1.3537 | | 2.9989 | 2.9989 | H4 | 0.9942 | 2.3675 | 2.9989 | | 1.6970 | H5 | 0.9942 | 2.3675 | 2.9989 | 1.6970 | |
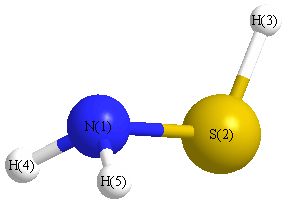 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
98.369 |
|
S2 |
N1 |
H4 |
116.874 |
| S2 |
N1 |
H5 |
116.874 |
|
H4 |
N1 |
H5 |
117.178 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-1.006 |
|
|
|
| 2 |
S |
0.185 |
|
|
|
| 3 |
H |
0.088 |
|
|
|
| 4 |
H |
0.366 |
|
|
|
| 5 |
H |
0.366 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.059 |
0.822 |
0.000 |
0.824 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.493 |
3.511 |
0.000 |
| y |
3.511 |
-18.596 |
0.000 |
| z |
0.000 |
0.000 |
-19.844 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.273 |
3.511 |
0.000 |
| y |
3.511 |
1.572 |
0.000 |
| z |
0.000 |
0.000 |
-0.300 |
|
| Polar |
|---|
| 3z2-r2 | -0.599 |
|---|
| x2-y2 | -1.897 |
|---|
| xy | 3.511 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.741 |
0.608 |
0.000 |
| y |
0.608 |
4.205 |
0.000 |
| z |
0.000 |
0.000 |
1.925 |
<r2> (average value of r
2) Å
2
| <r2> |
36.257 |
| (<r2>)1/2 |
6.021 |
Jump to
S1C1
Energy calculated at HF/SDD
| | hartrees |
|---|
| Energy at 0K | -453.622395 |
| Energy at 298.15K | -453.626051 |
| HF Energy | -453.622395 |
| Nuclear repulsion energy | 57.039783 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3837 |
3454 |
13.53 |
|
|
|
| 2 |
A' |
2633 |
2370 |
46.18 |
|
|
|
| 3 |
A' |
1770 |
1594 |
10.53 |
|
|
|
| 4 |
A' |
1089 |
981 |
7.86 |
|
|
|
| 5 |
A' |
845 |
761 |
43.26 |
|
|
|
| 6 |
A' |
433 |
390 |
360.14 |
|
|
|
| 7 |
A" |
3993 |
3596 |
44.27 |
|
|
|
| 8 |
A" |
1156 |
1041 |
8.62 |
|
|
|
| 9 |
A" |
515 |
464 |
59.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8134.9 cm
-1
Scaled (by 0.9004) Zero Point Vibrational Energy (zpe) 7324.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/SDD
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.032 |
1.115 |
0.000 |
| S2 |
0.032 |
-0.631 |
0.000 |
| H3 |
-1.308 |
-0.828 |
0.000 |
| H4 |
0.290 |
1.564 |
0.848 |
| H5 |
0.290 |
1.564 |
-0.848 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7459 | 2.3594 | 0.9942 | 0.9942 |
S2 | 1.7459 | | 1.3537 | 2.3676 | 2.3676 | H3 | 2.3594 | 1.3537 | | 2.9986 | 2.9986 | H4 | 0.9942 | 2.3676 | 2.9986 | | 1.6969 | H5 | 0.9942 | 2.3676 | 2.9986 | 1.6969 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
98.341 |
|
S2 |
N1 |
H4 |
116.869 |
| S2 |
N1 |
H5 |
116.869 |
|
H4 |
N1 |
H5 |
117.179 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-1.006 |
|
|
|
| 2 |
S |
0.185 |
|
|
|
| 3 |
H |
0.088 |
|
|
|
| 4 |
H |
0.366 |
|
|
|
| 5 |
H |
0.366 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.059 |
0.822 |
0.000 |
0.824 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.492 |
3.512 |
0.000 |
| y |
3.512 |
-18.597 |
0.000 |
| z |
0.000 |
0.000 |
-19.844 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.271 |
3.512 |
0.000 |
| y |
3.512 |
1.571 |
0.000 |
| z |
0.000 |
0.000 |
-0.300 |
|
| Polar |
|---|
| 3z2-r2 | -0.600 |
|---|
| x2-y2 | -1.895 |
|---|
| xy | 3.512 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.741 |
0.607 |
0.000 |
| y |
0.607 |
4.206 |
0.000 |
| z |
0.000 |
0.000 |
1.925 |
<r2> (average value of r
2) Å
2
| <r2> |
36.261 |
| (<r2>)1/2 |
6.022 |
