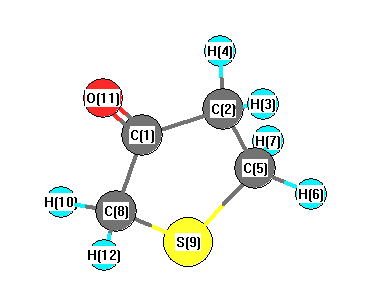
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3165 |
3033 |
4.93 |
|
|
|
| 2 |
A |
3159 |
3028 |
0.20 |
|
|
|
| 3 |
A |
3144 |
3013 |
6.92 |
|
|
|
| 4 |
A |
3087 |
2959 |
24.00 |
|
|
|
| 5 |
A |
3074 |
2946 |
15.20 |
|
|
|
| 6 |
A |
3073 |
2945 |
2.91 |
|
|
|
| 7 |
A |
1833 |
1756 |
247.91 |
|
|
|
| 8 |
A |
1474 |
1413 |
2.53 |
|
|
|
| 9 |
A |
1426 |
1367 |
5.97 |
|
|
|
| 10 |
A |
1423 |
1364 |
14.59 |
|
|
|
| 11 |
A |
1299 |
1244 |
13.94 |
|
|
|
| 12 |
A |
1284 |
1230 |
15.11 |
|
|
|
| 13 |
A |
1224 |
1173 |
22.30 |
|
|
|
| 14 |
A |
1213 |
1163 |
15.89 |
|
|
|
| 15 |
A |
1164 |
1115 |
18.71 |
|
|
|
| 16 |
A |
1154 |
1106 |
11.68 |
|
|
|
| 17 |
A |
1087 |
1041 |
1.85 |
|
|
|
| 18 |
A |
1011 |
969 |
8.61 |
|
|
|
| 19 |
A |
967 |
927 |
0.44 |
|
|
|
| 20 |
A |
867 |
831 |
5.17 |
|
|
|
| 21 |
A |
831 |
796 |
2.45 |
|
|
|
| 22 |
A |
800 |
767 |
0.82 |
|
|
|
| 23 |
A |
737 |
706 |
5.87 |
|
|
|
| 24 |
A |
687 |
659 |
0.92 |
|
|
|
| 25 |
A |
561 |
537 |
4.77 |
|
|
|
| 26 |
A |
491 |
470 |
5.13 |
|
|
|
| 27 |
A |
449 |
431 |
6.93 |
|
|
|
| 28 |
A |
428 |
410 |
2.91 |
|
|
|
| 29 |
A |
186 |
178 |
1.79 |
|
|
|
| 30 |
A |
68 |
65 |
11.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20682.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 19820.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.145 |
|
|
|
| 2 |
C |
0.372 |
|
|
|
| 3 |
H |
-0.135 |
|
|
|
| 4 |
H |
-0.074 |
|
|
|
| 5 |
C |
0.248 |
|
|
|
| 6 |
H |
-0.172 |
|
|
|
| 7 |
H |
-0.125 |
|
|
|
| 8 |
C |
-0.019 |
|
|
|
| 9 |
S |
0.408 |
|
|
|
| 10 |
H |
-0.026 |
|
|
|
| 11 |
O |
-0.624 |
|
|
|
| 12 |
H |
0.001 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.368 |
1.368 |
0.457 |
1.988 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-54.503 |
1.018 |
0.238 |
| y |
1.018 |
-38.493 |
-0.049 |
| z |
0.238 |
-0.049 |
-42.183 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-14.165 |
1.018 |
0.238 |
| y |
1.018 |
9.850 |
-0.049 |
| z |
0.238 |
-0.049 |
4.314 |
|
| Polar |
|---|
| 3z2-r2 | 8.629 |
|---|
| x2-y2 | -16.010 |
|---|
| xy | 1.018 |
|---|
| xz | 0.238 |
|---|
| yz | -0.049 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.666 |
0.033 |
-0.106 |
| y |
0.033 |
10.676 |
-0.021 |
| z |
-0.106 |
-0.021 |
8.050 |
<r2> (average value of r
2) Å
2
| <r2> |
185.362 |
| (<r2>)1/2 |
13.615 |