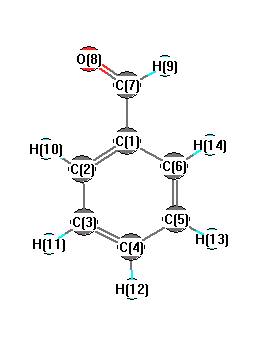
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3233 |
3098 |
8.43 |
244.64 |
0.10 |
0.18 |
| 2 |
A' |
3227 |
3092 |
9.20 |
82.31 |
0.14 |
0.25 |
| 3 |
A' |
3217 |
3082 |
9.87 |
80.20 |
0.75 |
0.86 |
| 4 |
A' |
3205 |
3072 |
1.05 |
81.08 |
0.69 |
0.82 |
| 5 |
A' |
3195 |
3062 |
3.30 |
44.88 |
0.40 |
0.57 |
| 6 |
A' |
2930 |
2808 |
106.30 |
139.98 |
0.28 |
0.44 |
| 7 |
A' |
1797 |
1722 |
289.14 |
113.11 |
0.35 |
0.52 |
| 8 |
A' |
1670 |
1601 |
31.09 |
93.11 |
0.56 |
0.72 |
| 9 |
A' |
1653 |
1584 |
12.31 |
11.15 |
0.56 |
0.72 |
| 10 |
A' |
1523 |
1460 |
1.44 |
1.14 |
0.50 |
0.67 |
| 11 |
A' |
1485 |
1423 |
12.79 |
2.03 |
0.31 |
0.48 |
| 12 |
A' |
1413 |
1354 |
7.46 |
2.36 |
0.53 |
0.70 |
| 13 |
A' |
1389 |
1331 |
7.55 |
0.80 |
0.75 |
0.86 |
| 14 |
A' |
1326 |
1270 |
15.19 |
1.37 |
0.30 |
0.47 |
| 15 |
A' |
1237 |
1185 |
61.44 |
31.85 |
0.18 |
0.31 |
| 16 |
A' |
1184 |
1134 |
16.39 |
8.11 |
0.13 |
0.23 |
| 17 |
A' |
1178 |
1129 |
1.70 |
6.24 |
0.71 |
0.83 |
| 18 |
A' |
1104 |
1058 |
5.76 |
1.11 |
0.08 |
0.15 |
| 19 |
A' |
1052 |
1008 |
2.78 |
22.50 |
0.04 |
0.07 |
| 20 |
A' |
1014 |
972 |
0.16 |
28.88 |
0.04 |
0.07 |
| 21 |
A' |
844 |
809 |
31.57 |
13.55 |
0.06 |
0.12 |
| 22 |
A' |
659 |
632 |
23.60 |
2.83 |
0.29 |
0.45 |
| 23 |
A' |
623 |
597 |
0.55 |
4.50 |
0.75 |
0.85 |
| 24 |
A' |
443 |
424 |
0.22 |
5.95 |
0.21 |
0.34 |
| 25 |
A' |
216 |
207 |
8.34 |
0.60 |
0.44 |
0.61 |
| 26 |
A" |
1040 |
996 |
1.21 |
1.38 |
0.75 |
0.86 |
| 27 |
A" |
1012 |
970 |
0.01 |
0.04 |
0.75 |
0.86 |
| 28 |
A" |
1000 |
958 |
0.00 |
0.05 |
0.75 |
0.86 |
| 29 |
A" |
946 |
906 |
1.33 |
0.55 |
0.75 |
0.86 |
| 30 |
A" |
866 |
830 |
0.00 |
0.38 |
0.75 |
0.86 |
| 31 |
A" |
766 |
734 |
42.68 |
0.37 |
0.75 |
0.86 |
| 32 |
A" |
707 |
678 |
35.71 |
0.02 |
0.75 |
0.86 |
| 33 |
A" |
470 |
450 |
7.44 |
0.14 |
0.75 |
0.86 |
| 34 |
A" |
417 |
400 |
0.11 |
0.00 |
0.75 |
0.86 |
| 35 |
A" |
240 |
230 |
7.67 |
0.81 |
0.75 |
0.86 |
| 36 |
A" |
121 |
116 |
4.73 |
1.20 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24199.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 23190.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.763 |
|
|
|
| 2 |
C |
0.381 |
|
|
|
| 3 |
C |
0.447 |
|
|
|
| 4 |
C |
0.277 |
|
|
|
| 5 |
C |
0.346 |
|
|
|
| 6 |
C |
0.305 |
|
|
|
| 7 |
C |
0.654 |
|
|
|
| 8 |
O |
-0.604 |
|
|
|
| 9 |
H |
-0.382 |
|
|
|
| 10 |
H |
-0.741 |
|
|
|
| 11 |
H |
-0.572 |
|
|
|
| 12 |
H |
-0.558 |
|
|
|
| 13 |
H |
-0.576 |
|
|
|
| 14 |
H |
-0.741 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.348 |
-2.571 |
0.000 |
3.482 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.560 |
6.481 |
0.000 |
| y |
6.481 |
-47.159 |
0.000 |
| z |
0.000 |
0.000 |
-49.206 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.623 |
6.481 |
0.000 |
| y |
6.481 |
-0.277 |
0.000 |
| z |
0.000 |
0.000 |
-3.346 |
|
| Polar |
|---|
| 3z2-r2 | -6.692 |
|---|
| x2-y2 | 2.600 |
|---|
| xy | 6.481 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.131 |
-1.227 |
0.000 |
| y |
-1.227 |
16.806 |
0.000 |
| z |
0.000 |
0.000 |
7.380 |
<r2> (average value of r
2) Å
2
| <r2> |
261.935 |
| (<r2>)1/2 |
16.184 |