return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C14H14 (Bibenzyl)

using model chemistry: B97D3/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at B97D3/aug-cc-pVDZ
 hartrees
Energy at 0K-541.636086
Energy at 298.15K 
HF Energy-541.636086
Nuclear repulsion energy810.244273
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3144 3144 24.98      
2 A 3133 3133 3.14      
3 A 3124 3124 26.71      
4 A 3115 3115 0.31      
5 A 3104 3104 19.15      
6 A 3031 3031 23.26      
7 A 2988 2988 62.24      
8 A 1611 1611 2.99      
9 A 1588 1588 0.12      
10 A 1484 1484 6.90      
11 A 1452 1452 0.44      
12 A 1441 1441 2.68      
13 A 1355 1355 0.27      
14 A 1321 1321 0.56      
15 A 1288 1288 0.36      
16 A 1211 1211 1.16      
17 A 1178 1178 2.14      
18 A 1175 1175 2.33      
19 A 1153 1153 0.02      
20 A 1086 1086 3.71      
21 A 1030 1030 2.93      
22 A 994 994 0.04      
23 A 986 986 0.24      
24 A 956 956 0.14      
25 A 938 938 0.04      
26 A 932 932 0.15      
27 A 880 880 0.17      
28 A 820 820 0.15      
29 A 762 762 0.77      
30 A 734 734 1.61      
31 A 688 688 6.58      
32 A 620 620 0.02      
33 A 529 529 2.09      
34 A 483 483 0.56      
35 A 401 401 0.01      
36 A 373 373 0.10      
37 A 230 230 0.31      
38 A 116 116 0.02      
39 A 54 54 0.00      
40 A 30 30 0.00      
41 B 3144 3144 11.64      
42 B 3133 3133 87.53      
43 B 3124 3124 3.53      
44 B 3115 3115 2.32      
45 B 3104 3104 2.14      
46 B 3044 3044 36.06      
47 B 2983 2983 20.41      
48 B 1609 1609 6.75      
49 B 1588 1588 0.51      
50 B 1486 1486 7.55      
51 B 1443 1443 6.24      
52 B 1442 1442 6.65      
53 B 1355 1355 0.64      
54 B 1328 1328 5.23      
55 B 1315 1315 0.17      
56 B 1194 1194 2.36      
57 B 1175 1175 0.27      
58 B 1153 1153 0.00      
59 B 1139 1139 1.13      
60 B 1071 1071 8.80      
61 B 1028 1028 4.13      
62 B 993 993 0.21      
63 B 957 957 0.10      
64 B 939 939 0.47      
65 B 923 923 5.37      
66 B 870 870 0.86      
67 B 820 820 0.16      
68 B 768 768 2.87      
69 B 741 741 33.95      
70 B 687 687 85.80      
71 B 620 620 0.29      
72 B 579 579 17.25      
73 B 503 503 13.63      
74 B 400 400 0.07      
75 B 360 360 0.21      
76 B 323 323 0.40      
77 B 118 118 1.14      
78 B 35 35 0.13      

Unscaled Zero Point Vibrational Energy (zpe) 51071.4 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 51071.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B97D3/aug-cc-pVDZ
ABC
0.04169 0.01649 0.01384

See section I.F.4 to change rotational constant units
Geometric Data calculated at B97D3/aug-cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.682 -1.429 -1.527
C2 1.682 1.429 -1.527
C3 1.526 0.638 -0.379
C4 0.718 3.319 -0.351
C5 -1.276 -2.772 -1.518
C6 1.276 2.772 -1.518
C7 -1.526 -0.638 -0.379
C8 -0.718 -3.319 -0.351
C9 -0.967 -1.173 0.796
C10 0.967 1.173 0.796
C11 0.566 2.523 0.794
C12 -0.566 -2.523 0.794
C13 -0.718 -0.300 2.005
C14 0.718 0.300 2.005
H15 -2.115 -0.993 -2.433
H16 2.115 0.993 -2.433
H17 -1.397 -3.391 -2.412
H18 1.397 3.391 -2.412
H19 -1.824 0.414 -0.399
H20 1.824 -0.414 -0.399
H21 0.404 4.367 -0.332
H22 -0.404 -4.367 -0.332
H23 0.128 2.955 1.700
H24 -0.128 -2.955 1.700
H25 1.446 -0.527 2.027
H26 -1.446 0.527 2.027
H27 -0.853 -0.886 2.929
H28 0.853 0.886 2.929

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 C10 C11 C12 C13 C14 H15 H16 H17 H18 H19 H20 H21 H22 H23 H24 H25 H26 H27 H28
C14.41313.98495.44761.40325.13761.40242.42562.44404.37975.10412.79803.83064.60651.09444.59382.17115.78672.16523.82026.27423.41955.73643.89354.81944.06324.56495.6250
C24.41311.40242.42565.13761.40323.98495.44764.37972.44402.79805.10414.60653.83064.59381.09445.78672.17113.82022.16523.41956.27423.89355.73644.06324.81945.62504.5649
C33.98491.40242.79994.55842.43163.30854.54883.29821.40732.41883.96793.40582.53954.48772.16565.37663.42443.35831.09393.89445.36453.41294.46882.67443.82584.35083.3853
C45.44762.42562.79996.51401.40414.54886.79064.93262.44561.40266.08944.55063.82875.56353.41947.33032.17123.86023.89371.09457.76702.16526.65454.57964.25815.55974.0858
C51.40325.13764.55846.51406.10342.43161.40412.82945.09426.06402.43114.33905.08172.16905.14961.09426.77673.42104.05277.42922.17016.71763.42165.00164.84514.84866.1394
C65.13761.40322.43161.40416.10344.55846.51405.09422.82942.43116.06405.08174.33905.14962.16906.77671.09424.05273.42102.17017.42923.42166.71764.84515.00166.13944.8486
C71.40243.98493.30854.54882.43164.55842.79991.40733.29823.96792.41882.53953.40582.16564.48773.42445.37661.09393.35835.36453.89444.46883.41293.82582.67443.38534.3508
C82.42565.44764.54886.79061.40416.51402.79992.44564.93266.08941.40263.82874.55063.41945.56352.17127.33033.89373.86027.76701.09456.65452.16524.25814.57964.08585.5597
C92.44404.37973.29824.93262.82945.09421.40732.44563.04074.00131.40791.51132.54383.43164.96183.92366.05882.16443.13015.81783.43384.36562.16712.78492.15282.15523.4791
C104.37972.44401.40732.44565.09422.82943.29824.93263.04071.40794.00132.54381.51134.96183.43166.05883.92363.13012.16443.43385.81782.16714.36562.15282.78493.47912.1552
C115.10412.79802.41881.40266.06402.43113.96796.08944.00131.40795.17103.32942.53545.47383.89237.00713.42383.40353.41092.16707.04831.09555.59553.40523.09074.26572.7054
C122.79805.10413.96796.08942.43116.06402.41881.40261.40794.00135.17102.53543.32943.89235.47383.42387.00713.41093.40357.04832.16705.59551.09553.09073.40522.70544.2657
C133.83064.60653.40584.55064.33905.08172.53953.82871.51132.54383.32942.53541.55674.70335.42135.43306.13222.74123.50105.33904.70083.37732.73652.17611.10221.10302.1752
C144.60653.83062.53953.82875.08174.33903.40584.55062.54381.51132.53543.32941.55675.42134.70336.13225.43303.50102.74124.70085.33902.73653.37731.10222.17612.17521.1030
H151.09444.59384.48775.56352.16905.14962.16563.41943.43164.96185.47383.89234.70335.42134.67402.50275.61752.48994.47126.28414.32676.14024.98785.72614.75875.50946.4103
H164.59381.09442.16563.41945.14962.16904.48775.56354.96183.43163.89235.47385.42134.70334.67405.61752.50274.47122.48994.32676.28414.98786.14024.75875.72616.41035.5094
H172.17115.78675.37667.33031.09426.77673.42442.17123.92366.05887.00713.42385.43306.13222.50275.61757.33424.32564.82578.23092.50287.71394.32575.99915.92065.92427.2031
H185.78672.17113.42442.17126.77671.09425.37667.33036.05883.92363.42387.00716.13225.43305.61752.50277.33424.82574.32562.50288.23094.32577.71395.92065.99917.20315.9242
H192.16523.82023.35833.86023.42104.05271.09393.89372.16443.13013.40353.41092.74123.50102.48994.47124.32564.82573.74194.53814.98823.83104.31734.17972.45823.70354.2977
H203.82022.16521.09393.89374.05273.42103.35833.86023.13012.16443.41093.40353.50102.74124.47122.48994.82574.32563.74194.98824.53814.31733.83102.45824.17974.29773.7035
H216.27423.41953.89441.09457.42922.17015.36457.76705.81783.43382.16707.04835.33904.70086.28414.32678.23092.50284.53814.98828.77122.49037.61755.53194.87196.31014.7910
H223.41956.27425.36457.76702.17017.42923.89441.09453.43385.81787.04832.16704.70085.33904.32676.28412.50288.23094.98824.53818.77127.61752.49034.87195.53194.79106.3101
H235.73643.89353.41292.16526.71763.42164.46886.65454.36562.16711.09555.59553.37732.73656.14024.98787.71394.32573.83104.31732.49037.61755.91573.73742.91214.15092.5129
H243.89355.73644.46886.65453.42166.71763.41292.16522.16714.36565.59551.09552.73653.37734.98786.14024.32577.71394.31733.83107.61752.49035.91572.91213.73742.51294.1509
H254.81944.06322.67444.57965.00164.84513.82584.25812.78492.15283.40523.09072.17611.10225.72614.75875.99915.92064.17972.45825.53194.87193.73742.91213.07832.49571.7787
H264.06324.81943.82584.25814.84515.00162.67444.57962.15282.78493.09073.40521.10222.17614.75875.72615.92065.99912.45824.17974.87195.53192.91213.73743.07831.77872.4957
H274.56495.62504.35085.55974.84866.13943.38534.08582.15523.47914.26572.70541.10302.17525.50946.41035.92427.20313.70354.29776.31014.79104.15092.51292.49571.77872.4603
H285.62504.56493.38534.08586.13944.84864.35085.55973.47912.15522.70544.26572.17521.10306.41035.50947.20315.92424.29773.70354.79106.31012.51294.15091.77872.49572.4603

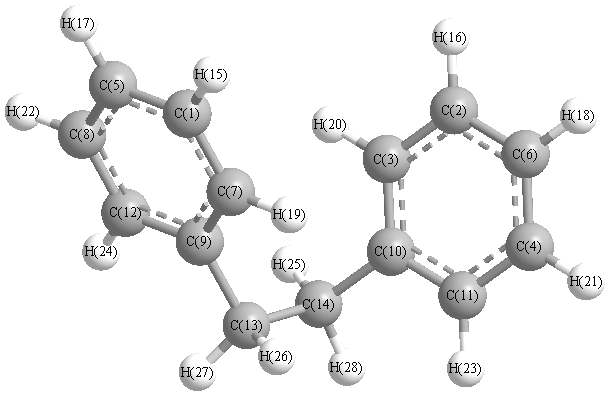
picture of Bibenzyl state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 C8 119.543 C1 C5 H17 120.261
C1 C7 C9 120.878 C1 C7 H19 119.796
C2 C3 C10 120.878 C2 C3 H20 119.796
C2 C6 C4 119.543 C2 C6 H18 120.261
C3 C2 C6 120.149 C3 C2 H16 119.799
C3 C10 C11 118.449 C3 C10 C14 120.904
C4 C6 H18 120.197 C4 C11 C10 120.951
C4 C11 H23 119.658 C5 C1 C7 120.149
C5 C1 H15 120.051 C5 C8 C12 120.031
C5 C8 H22 120.070 C6 C2 H16 120.051
C6 C4 C11 120.031 C6 C4 H21 120.070
C7 C1 H15 119.799 C7 C9 C12 118.449
C7 C9 C13 120.904 C8 C5 H17 120.197
C8 C12 C9 120.951 C8 C12 H24 119.658
C9 C7 H19 119.315 C9 C12 H24 119.391
C9 C13 C14 112.013 C9 C13 H26 109.928
C9 C13 H27 110.075 C10 C3 H20 119.315
C10 C11 H23 119.391 C10 C14 C13 112.013
C10 C14 H25 109.928 C10 C14 H28 110.075
C11 C4 H21 119.898 C11 C10 C14 120.529
C12 C8 H22 119.898 C12 C9 C13 120.529
C13 C14 H25 108.647 C13 C14 H28 108.529
C14 C13 H26 108.647 C14 C13 H27 108.529
H25 C14 H28 107.527 H26 C13 H27 107.527
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.070      
2 C 0.070      
3 C 1.079      
4 C 0.250      
5 C 0.807      
6 C 0.807      
7 C 1.079      
8 C 0.250      
9 C 1.367      
10 C 1.367      
11 C 0.875      
12 C 0.875      
13 C 0.639      
14 C 0.639      
15 H -0.755      
16 H -0.755      
17 H -0.714      
18 H -0.714      
19 H -1.044      
20 H -1.044      
21 H -0.727      
22 H -0.727      
23 H -0.980      
24 H -0.980      
25 H -0.440      
26 H -0.440      
27 H -0.427      
28 H -0.427      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.000 0.000 0.343 0.343
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -88.862 -4.312 0.000
y -4.312 -79.906 0.000
z 0.000 0.000 -77.724
Traceless
 xyz
x -10.047 -4.312 0.000
y -4.312 3.387 0.000
z 0.000 0.000 6.659
Polar
3z2-r213.319
x2-y2-8.956
xy-4.312
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 18.272 0.262 0.000
y 0.262 29.102 0.000
z 0.000 0.000 26.419


<r2> (average value of r2) Å2
<r2> 793.832
(<r2>)1/2 28.175